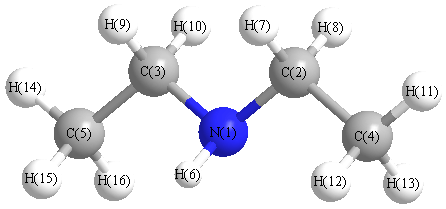
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3740 |
3390 |
0.10 |
|
|
|
| 2 |
A' |
3231 |
2928 |
80.91 |
|
|
|
| 3 |
A' |
3212 |
2912 |
113.98 |
|
|
|
| 4 |
A' |
3174 |
2877 |
46.14 |
|
|
|
| 5 |
A' |
3156 |
2861 |
23.08 |
|
|
|
| 6 |
A' |
3075 |
2787 |
182.07 |
|
|
|
| 7 |
A' |
1651 |
1496 |
1.08 |
|
|
|
| 8 |
A' |
1615 |
1464 |
2.98 |
|
|
|
| 9 |
A' |
1607 |
1456 |
11.08 |
|
|
|
| 10 |
A' |
1561 |
1415 |
2.16 |
|
|
|
| 11 |
A' |
1520 |
1378 |
0.48 |
|
|
|
| 12 |
A' |
1429 |
1295 |
5.84 |
|
|
|
| 13 |
A' |
1329 |
1205 |
2.47 |
|
|
|
| 14 |
A' |
1269 |
1150 |
18.53 |
|
|
|
| 15 |
A' |
1123 |
1018 |
6.50 |
|
|
|
| 16 |
A' |
961 |
871 |
2.68 |
|
|
|
| 17 |
A' |
889 |
805 |
3.44 |
|
|
|
| 18 |
A' |
816 |
740 |
79.36 |
|
|
|
| 19 |
A' |
450 |
408 |
0.45 |
|
|
|
| 20 |
A' |
276 |
250 |
1.22 |
|
|
|
| 21 |
A' |
199 |
181 |
0.87 |
|
|
|
| 22 |
A' |
119 |
108 |
1.50 |
|
|
|
| 23 |
A" |
3230 |
2928 |
11.63 |
|
|
|
| 24 |
A" |
3211 |
2910 |
35.19 |
|
|
|
| 25 |
A" |
3167 |
2871 |
2.07 |
|
|
|
| 26 |
A" |
3155 |
2860 |
39.07 |
|
|
|
| 27 |
A" |
3066 |
2779 |
7.08 |
|
|
|
| 28 |
A" |
1644 |
1490 |
41.68 |
|
|
|
| 29 |
A" |
1637 |
1484 |
0.64 |
|
|
|
| 30 |
A" |
1612 |
1461 |
3.63 |
|
|
|
| 31 |
A" |
1601 |
1452 |
8.77 |
|
|
|
| 32 |
A" |
1535 |
1392 |
9.21 |
|
|
|
| 33 |
A" |
1481 |
1342 |
24.72 |
|
|
|
| 34 |
A" |
1392 |
1262 |
2.26 |
|
|
|
| 35 |
A" |
1249 |
1132 |
58.04 |
|
|
|
| 36 |
A" |
1209 |
1096 |
7.13 |
|
|
|
| 37 |
A" |
1146 |
1038 |
0.04 |
|
|
|
| 38 |
A" |
1000 |
906 |
0.72 |
|
|
|
| 39 |
A" |
865 |
784 |
0.43 |
|
|
|
| 40 |
A" |
457 |
414 |
0.23 |
|
|
|
| 41 |
A" |
276 |
250 |
0.55 |
|
|
|
| 42 |
A" |
113 |
102 |
0.87 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34723.2 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 31473.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.310 |
|
|
|
| 2 |
C |
0.067 |
|
|
|
| 3 |
C |
0.067 |
|
|
|
| 4 |
C |
-0.167 |
|
|
|
| 5 |
C |
-0.167 |
|
|
|
| 6 |
H |
0.109 |
|
|
|
| 7 |
H |
0.018 |
|
|
|
| 8 |
H |
0.035 |
|
|
|
| 9 |
H |
0.018 |
|
|
|
| 10 |
H |
0.035 |
|
|
|
| 11 |
H |
0.049 |
|
|
|
| 12 |
H |
0.036 |
|
|
|
| 13 |
H |
0.062 |
|
|
|
| 14 |
H |
0.049 |
|
|
|
| 15 |
H |
0.036 |
|
|
|
| 16 |
H |
0.062 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.826 |
0.411 |
0.000 |
0.922 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.980 |
1.637 |
0.000 |
| y |
1.637 |
-34.592 |
0.000 |
| z |
0.000 |
0.000 |
-33.680 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.156 |
1.637 |
0.000 |
| y |
1.637 |
-0.762 |
0.000 |
| z |
0.000 |
0.000 |
0.606 |
|
| Polar |
|---|
| 3z2-r2 | 1.212 |
|---|
| x2-y2 | 0.612 |
|---|
| xy | 1.637 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.418 |
0.015 |
0.000 |
| y |
0.015 |
7.890 |
0.000 |
| z |
0.000 |
0.000 |
9.786 |
<r2> (average value of r
2) Å
2
| <r2> |
186.305 |
| (<r2>)1/2 |
13.649 |