Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -229.860223 |
| Energy at 298.15K | -229.866223 |
| HF Energy | -229.860223 |
| Nuclear repulsion energy | 160.698994 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3403 |
3084 |
10.05 |
|
|
|
| 2 |
A1 |
3331 |
3019 |
14.94 |
|
|
|
| 3 |
A1 |
3307 |
2998 |
5.05 |
|
|
|
| 4 |
A1 |
1879 |
1703 |
5.98 |
|
|
|
| 5 |
A1 |
1564 |
1418 |
0.70 |
|
|
|
| 6 |
A1 |
1471 |
1334 |
2.71 |
|
|
|
| 7 |
A1 |
1319 |
1195 |
14.22 |
|
|
|
| 8 |
A1 |
941 |
853 |
5.05 |
|
|
|
| 9 |
A1 |
558 |
506 |
0.52 |
|
|
|
| 10 |
A1 |
249 |
226 |
0.01 |
|
|
|
| 11 |
A2 |
1080 |
979 |
0.00 |
|
|
|
| 12 |
A2 |
996 |
902 |
0.00 |
|
|
|
| 13 |
A2 |
781 |
708 |
0.00 |
|
|
|
| 14 |
A2 |
87 |
79 |
0.00 |
|
|
|
| 15 |
B1 |
1092 |
990 |
33.85 |
|
|
|
| 16 |
B1 |
1000 |
906 |
152.68 |
|
|
|
| 17 |
B1 |
770 |
698 |
10.02 |
|
|
|
| 18 |
B1 |
66 |
60 |
1.50 |
|
|
|
| 19 |
B2 |
3403 |
3084 |
3.93 |
|
|
|
| 20 |
B2 |
3320 |
3009 |
1.17 |
|
|
|
| 21 |
B2 |
3303 |
2993 |
2.43 |
|
|
|
| 22 |
B2 |
1827 |
1656 |
569.96 |
|
|
|
| 23 |
B2 |
1549 |
1404 |
97.31 |
|
|
|
| 24 |
B2 |
1443 |
1308 |
24.21 |
|
|
|
| 25 |
B2 |
1317 |
1194 |
599.89 |
|
|
|
| 26 |
B2 |
1127 |
1022 |
10.73 |
|
|
|
| 27 |
B2 |
529 |
480 |
9.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20854.3 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 18902.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.345 |
| C2 |
0.000 |
1.174 |
-0.386 |
| C3 |
0.000 |
-1.174 |
-0.386 |
| C4 |
0.000 |
2.350 |
0.235 |
| C5 |
0.000 |
-2.350 |
0.235 |
| H6 |
0.000 |
1.107 |
-1.433 |
| H7 |
0.000 |
-1.107 |
-1.433 |
| H8 |
0.000 |
3.234 |
-0.336 |
| H9 |
0.000 |
2.404 |
1.291 |
| H10 |
0.000 |
-3.234 |
-0.336 |
| H11 |
0.000 |
-2.404 |
1.291 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3830 | 1.3830 | 2.3530 | 2.3530 | 2.0943 | 2.0943 | 3.3048 | 2.5835 | 3.3048 | 2.5835 |
C2 | 1.3830 | | 2.3485 | 1.3299 | 3.5789 | 1.0497 | 2.5099 | 2.0601 | 2.0791 | 4.4083 | 3.9516 | C3 | 1.3830 | 2.3485 | | 3.5789 | 1.3299 | 2.5099 | 1.0497 | 4.4083 | 3.9516 | 2.0601 | 2.0791 | C4 | 2.3530 | 1.3299 | 3.5789 | | 4.7008 | 2.0809 | 3.8385 | 1.0522 | 1.0568 | 5.6133 | 4.8703 | C5 | 2.3530 | 3.5789 | 1.3299 | 4.7008 | | 3.8385 | 2.0809 | 5.6133 | 4.8703 | 1.0522 | 1.0568 | H6 | 2.0943 | 1.0497 | 2.5099 | 2.0809 | 3.8385 | | 2.2132 | 2.3932 | 3.0171 | 4.4768 | 4.4435 | H7 | 2.0943 | 2.5099 | 1.0497 | 3.8385 | 2.0809 | 2.2132 | | 4.4768 | 4.4435 | 2.3932 | 3.0171 | H8 | 3.3048 | 2.0601 | 4.4083 | 1.0522 | 5.6133 | 2.3932 | 4.4768 | | 1.8265 | 6.4675 | 5.8681 | H9 | 2.5835 | 2.0791 | 3.9516 | 1.0568 | 4.8703 | 3.0171 | 4.4435 | 1.8265 | | 5.8681 | 4.8083 | H10 | 3.3048 | 4.4083 | 2.0601 | 5.6133 | 1.0522 | 4.4768 | 2.3932 | 6.4675 | 5.8681 | | 1.8265 | H11 | 2.5835 | 3.9516 | 2.0791 | 4.8703 | 1.0568 | 4.4435 | 3.0171 | 5.8681 | 4.8083 | 1.8265 | |
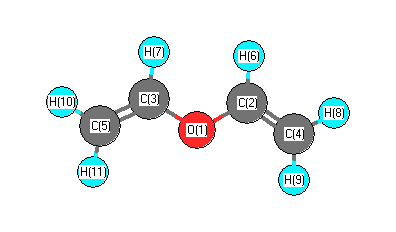 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
120.287 |
|
O1 |
C2 |
H6 |
118.192 |
| O1 |
C3 |
C5 |
120.287 |
|
O1 |
C3 |
H7 |
118.192 |
| C2 |
O1 |
C3 |
116.225 |
|
C2 |
C4 |
H8 |
119.264 |
| C2 |
C4 |
H9 |
120.742 |
|
C3 |
C5 |
H10 |
119.264 |
| C3 |
C5 |
H11 |
120.742 |
|
C4 |
C2 |
H6 |
121.521 |
| C5 |
C3 |
H7 |
121.521 |
|
H8 |
C4 |
H9 |
119.994 |
| H10 |
C5 |
H11 |
119.994 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.324 |
|
|
|
| 2 |
C |
0.150 |
|
|
|
| 3 |
C |
0.150 |
|
|
|
| 4 |
C |
-0.201 |
|
|
|
| 5 |
C |
-0.201 |
|
|
|
| 6 |
H |
0.063 |
|
|
|
| 7 |
H |
0.063 |
|
|
|
| 8 |
H |
0.077 |
|
|
|
| 9 |
H |
0.074 |
|
|
|
| 10 |
H |
0.077 |
|
|
|
| 11 |
H |
0.074 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.417 |
1.417 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.543 |
0.000 |
0.000 |
| y |
0.000 |
-28.098 |
0.000 |
| z |
0.000 |
0.000 |
-27.676 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.656 |
0.000 |
0.000 |
| y |
0.000 |
3.011 |
0.000 |
| z |
0.000 |
0.000 |
3.645 |
|
| Polar |
|---|
| 3z2-r2 | 7.290 |
|---|
| x2-y2 | -6.445 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.065 |
0.000 |
0.000 |
| y |
0.000 |
12.125 |
0.000 |
| z |
0.000 |
0.000 |
6.651 |
<r2> (average value of r
2) Å
2
| <r2> |
145.551 |
| (<r2>)1/2 |
12.064 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -229.860035 |
| Energy at 298.15K | -229.866368 |
| HF Energy | -229.860035 |
| Nuclear repulsion energy | 163.264328 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3401 |
3083 |
15.22 |
|
|
|
| 2 |
A |
3399 |
3081 |
5.20 |
|
|
|
| 3 |
A |
3359 |
3045 |
9.09 |
|
|
|
| 4 |
A |
3333 |
3021 |
6.48 |
|
|
|
| 5 |
A |
3313 |
3003 |
1.45 |
|
|
|
| 6 |
A |
3304 |
2995 |
1.18 |
|
|
|
| 7 |
A |
1853 |
1680 |
107.31 |
|
|
|
| 8 |
A |
1826 |
1655 |
255.71 |
|
|
|
| 9 |
A |
1567 |
1420 |
1.02 |
|
|
|
| 10 |
A |
1545 |
1401 |
31.27 |
|
|
|
| 11 |
A |
1476 |
1338 |
86.41 |
|
|
|
| 12 |
A |
1440 |
1305 |
4.34 |
|
|
|
| 13 |
A |
1357 |
1230 |
363.28 |
|
|
|
| 14 |
A |
1234 |
1119 |
15.83 |
|
|
|
| 15 |
A |
1120 |
1015 |
12.21 |
|
|
|
| 16 |
A |
1110 |
1006 |
12.11 |
|
|
|
| 17 |
A |
1102 |
999 |
69.23 |
|
|
|
| 18 |
A |
1022 |
927 |
66.16 |
|
|
|
| 19 |
A |
992 |
900 |
77.84 |
|
|
|
| 20 |
A |
955 |
866 |
16.18 |
|
|
|
| 21 |
A |
810 |
734 |
9.98 |
|
|
|
| 22 |
A |
789 |
715 |
2.45 |
|
|
|
| 23 |
A |
635 |
575 |
3.29 |
|
|
|
| 24 |
A |
495 |
448 |
3.77 |
|
|
|
| 25 |
A |
298 |
270 |
1.74 |
|
|
|
| 26 |
A |
196 |
178 |
8.95 |
|
|
|
| 27 |
A |
92 |
83 |
0.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21010.0 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 19043.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.038 |
-0.786 |
0.063 |
| C2 |
1.273 |
-0.480 |
-0.009 |
| C3 |
-0.934 |
0.194 |
0.332 |
| C4 |
1.809 |
0.711 |
-0.149 |
| C5 |
-2.119 |
0.222 |
-0.224 |
| H6 |
1.868 |
-1.370 |
0.047 |
| H7 |
-0.612 |
0.909 |
1.066 |
| H8 |
2.876 |
0.794 |
-0.190 |
| H9 |
1.232 |
1.609 |
-0.238 |
| H10 |
-2.825 |
0.973 |
0.066 |
| H11 |
-2.411 |
-0.504 |
-0.957 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3474 | 1.3551 | 2.3868 | 2.3300 | 1.9927 | 2.0515 | 3.3241 | 2.7279 | 3.2958 | 2.5989 |
C2 | 1.3474 | | 2.3322 | 1.3140 | 3.4700 | 1.0715 | 2.5760 | 2.0561 | 2.1026 | 4.3480 | 3.8041 | C3 | 1.3551 | 2.3322 | | 2.8322 | 1.3092 | 3.2210 | 1.0738 | 3.8917 | 2.6492 | 2.0623 | 2.0815 | C4 | 2.3868 | 1.3140 | 2.8322 | | 3.9590 | 2.0907 | 2.7157 | 1.0708 | 1.0715 | 4.6461 | 4.4657 | C5 | 2.3300 | 3.4700 | 1.3092 | 3.9590 | | 4.3009 | 2.0993 | 5.0275 | 3.6269 | 1.0710 | 1.0724 | H6 | 1.9927 | 1.0715 | 3.2210 | 2.0907 | 4.3009 | | 3.5183 | 2.3989 | 3.0594 | 5.2447 | 4.4795 | H7 | 2.0515 | 2.5760 | 1.0738 | 2.7157 | 2.0993 | 3.5183 | | 3.7087 | 2.3645 | 2.4291 | 3.0541 | H8 | 3.3241 | 2.0561 | 3.8917 | 1.0708 | 5.0275 | 2.3989 | 3.7087 | | 1.8354 | 5.7090 | 5.4980 | H9 | 2.7279 | 2.1026 | 2.6492 | 1.0715 | 3.6269 | 3.0594 | 2.3645 | 1.8354 | | 4.1174 | 4.2730 | H10 | 3.2958 | 4.3480 | 2.0623 | 4.6461 | 1.0710 | 5.2447 | 2.4291 | 5.7090 | 4.1174 | | 1.8440 | H11 | 2.5989 | 3.8041 | 2.0815 | 4.4657 | 1.0724 | 4.4795 | 3.0541 | 5.4980 | 4.2730 | 1.8440 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
127.480 |
|
O1 |
C2 |
H6 |
110.420 |
| O1 |
C3 |
C5 |
121.969 |
|
O1 |
C3 |
H7 |
114.763 |
| C2 |
O1 |
C3 |
119.302 |
|
C2 |
C4 |
H8 |
118.770 |
| C2 |
C4 |
H9 |
123.308 |
|
C3 |
C5 |
H10 |
119.757 |
| C3 |
C5 |
H11 |
121.530 |
|
C4 |
C2 |
H6 |
122.092 |
| C5 |
C3 |
H7 |
123.203 |
|
H8 |
C4 |
H9 |
117.916 |
| H10 |
C5 |
H11 |
118.705 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.357 |
|
|
|
| 2 |
C |
0.149 |
|
|
|
| 3 |
C |
0.135 |
|
|
|
| 4 |
C |
-0.194 |
|
|
|
| 5 |
C |
-0.181 |
|
|
|
| 6 |
H |
0.082 |
|
|
|
| 7 |
H |
0.081 |
|
|
|
| 8 |
H |
0.069 |
|
|
|
| 9 |
H |
0.062 |
|
|
|
| 10 |
H |
0.077 |
|
|
|
| 11 |
H |
0.077 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.343 |
0.663 |
0.273 |
0.794 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.610 |
-1.825 |
0.530 |
| y |
-1.825 |
-29.902 |
1.183 |
| z |
0.530 |
1.183 |
-32.759 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.721 |
-1.825 |
0.530 |
| y |
-1.825 |
0.282 |
1.183 |
| z |
0.530 |
1.183 |
-4.003 |
|
| Polar |
|---|
| 3z2-r2 | -8.005 |
|---|
| x2-y2 | 2.292 |
|---|
| xy | -1.825 |
|---|
| xz | 0.530 |
|---|
| yz | 1.183 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.245 |
0.589 |
0.572 |
| y |
0.589 |
7.033 |
0.002 |
| z |
0.572 |
0.002 |
5.552 |
<r2> (average value of r
2) Å
2
| <r2> |
128.606 |
| (<r2>)1/2 |
11.340 |
