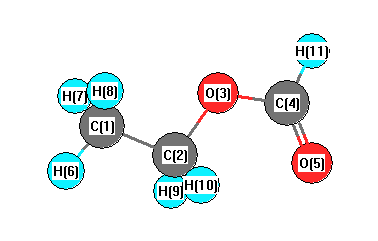
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3237 |
2934 |
42.90 |
58.54 |
0.60 |
0.75 |
| 2 |
A' |
3224 |
2922 |
61.95 |
122.18 |
0.21 |
0.34 |
| 3 |
A' |
3210 |
2909 |
13.72 |
109.40 |
0.15 |
0.26 |
| 4 |
A' |
3172 |
2875 |
20.90 |
139.10 |
0.01 |
0.01 |
| 5 |
A' |
1967 |
1783 |
432.36 |
6.70 |
0.24 |
0.39 |
| 6 |
A' |
1647 |
1493 |
7.72 |
2.07 |
0.70 |
0.82 |
| 7 |
A' |
1616 |
1465 |
2.73 |
8.05 |
0.74 |
0.85 |
| 8 |
A' |
1562 |
1415 |
15.20 |
1.91 |
0.44 |
0.61 |
| 9 |
A' |
1538 |
1394 |
4.76 |
3.34 |
0.34 |
0.50 |
| 10 |
A' |
1521 |
1379 |
0.20 |
0.16 |
0.50 |
0.66 |
| 11 |
A' |
1362 |
1234 |
558.59 |
2.01 |
0.14 |
0.24 |
| 12 |
A' |
1228 |
1113 |
16.39 |
5.73 |
0.14 |
0.25 |
| 13 |
A' |
1107 |
1004 |
20.33 |
4.15 |
0.69 |
0.82 |
| 14 |
A' |
938 |
851 |
19.74 |
6.94 |
0.25 |
0.40 |
| 15 |
A' |
856 |
776 |
2.51 |
3.83 |
0.37 |
0.54 |
| 16 |
A' |
409 |
371 |
6.06 |
1.63 |
0.29 |
0.46 |
| 17 |
A' |
243 |
220 |
9.44 |
0.39 |
0.33 |
0.49 |
| 18 |
A" |
3262 |
2957 |
56.29 |
4.29 |
0.75 |
0.86 |
| 19 |
A" |
3232 |
2929 |
3.80 |
96.61 |
0.75 |
0.86 |
| 20 |
A" |
1602 |
1452 |
5.32 |
7.15 |
0.75 |
0.86 |
| 21 |
A" |
1415 |
1283 |
2.16 |
4.89 |
0.75 |
0.86 |
| 22 |
A" |
1288 |
1167 |
4.60 |
0.19 |
0.75 |
0.86 |
| 23 |
A" |
1190 |
1078 |
1.89 |
1.31 |
0.75 |
0.86 |
| 24 |
A" |
874 |
792 |
0.12 |
0.07 |
0.75 |
0.86 |
| 25 |
A" |
349 |
317 |
26.13 |
1.05 |
0.75 |
0.86 |
| 26 |
A" |
255 |
231 |
4.59 |
0.58 |
0.75 |
0.86 |
| 27 |
A" |
75 |
68 |
1.02 |
0.13 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 21188.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 19205.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.150 |
|
|
|
| 2 |
C |
0.144 |
|
|
|
| 3 |
O |
-0.365 |
|
|
|
| 4 |
C |
0.448 |
|
|
|
| 5 |
O |
-0.438 |
|
|
|
| 6 |
H |
0.052 |
|
|
|
| 7 |
H |
0.060 |
|
|
|
| 8 |
H |
0.060 |
|
|
|
| 9 |
H |
0.060 |
|
|
|
| 10 |
H |
0.060 |
|
|
|
| 11 |
H |
0.068 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.177 |
0.733 |
0.000 |
2.297 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.072 |
5.241 |
0.000 |
| y |
5.241 |
-28.958 |
0.000 |
| z |
0.000 |
0.000 |
-29.229 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.978 |
5.241 |
0.000 |
| y |
5.241 |
2.692 |
0.000 |
| z |
0.000 |
0.000 |
2.286 |
|
| Polar |
|---|
| 3z2-r2 | 4.571 |
|---|
| x2-y2 | -5.114 |
|---|
| xy | 5.241 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.062 |
0.132 |
0.000 |
| y |
0.132 |
5.940 |
0.000 |
| z |
0.000 |
0.000 |
4.828 |
<r2> (average value of r
2) Å
2
| <r2> |
134.241 |
| (<r2>)1/2 |
11.586 |