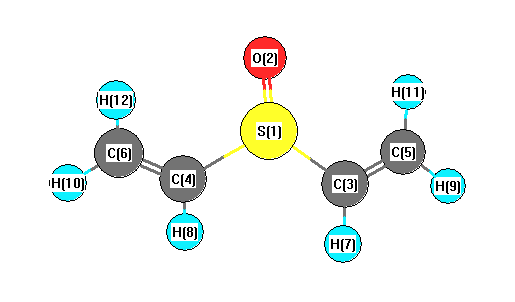
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3389 |
3072 |
0.48 |
|
|
|
| 2 |
A' |
3328 |
3016 |
7.25 |
|
|
|
| 3 |
A' |
3294 |
2986 |
2.57 |
|
|
|
| 4 |
A' |
1810 |
1641 |
0.27 |
|
|
|
| 5 |
A' |
1533 |
1389 |
14.14 |
|
|
|
| 6 |
A' |
1400 |
1269 |
0.19 |
|
|
|
| 7 |
A' |
1222 |
1108 |
120.86 |
|
|
|
| 8 |
A' |
1117 |
1013 |
53.88 |
|
|
|
| 9 |
A' |
1106 |
1002 |
7.79 |
|
|
|
| 10 |
A' |
1095 |
992 |
24.01 |
|
|
|
| 11 |
A' |
788 |
714 |
67.34 |
|
|
|
| 12 |
A' |
705 |
639 |
6.46 |
|
|
|
| 13 |
A' |
536 |
486 |
1.53 |
|
|
|
| 14 |
A' |
331 |
300 |
2.36 |
|
|
|
| 15 |
A' |
224 |
203 |
2.98 |
|
|
|
| 16 |
A' |
101 |
91 |
1.05 |
|
|
|
| 17 |
A" |
3389 |
3072 |
1.42 |
|
|
|
| 18 |
A" |
3322 |
3011 |
5.59 |
|
|
|
| 19 |
A" |
3293 |
2985 |
6.39 |
|
|
|
| 20 |
A" |
1800 |
1631 |
0.53 |
|
|
|
| 21 |
A" |
1528 |
1385 |
7.50 |
|
|
|
| 22 |
A" |
1383 |
1253 |
8.79 |
|
|
|
| 23 |
A" |
1114 |
1010 |
26.11 |
|
|
|
| 24 |
A" |
1094 |
992 |
0.27 |
|
|
|
| 25 |
A" |
1081 |
980 |
26.88 |
|
|
|
| 26 |
A" |
726 |
658 |
13.30 |
|
|
|
| 27 |
A" |
622 |
563 |
3.61 |
|
|
|
| 28 |
A" |
536 |
486 |
20.99 |
|
|
|
| 29 |
A" |
280 |
254 |
5.77 |
|
|
|
| 30 |
A" |
164 |
148 |
0.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21154.2 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 19174.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.718 |
|
|
|
| 2 |
O |
-0.682 |
|
|
|
| 3 |
C |
-0.149 |
|
|
|
| 4 |
C |
-0.149 |
|
|
|
| 5 |
C |
-0.143 |
|
|
|
| 6 |
C |
-0.143 |
|
|
|
| 7 |
H |
0.075 |
|
|
|
| 8 |
H |
0.075 |
|
|
|
| 9 |
H |
0.085 |
|
|
|
| 10 |
H |
0.085 |
|
|
|
| 11 |
H |
0.114 |
|
|
|
| 12 |
H |
0.114 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.247 |
-1.764 |
0.000 |
3.695 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.336 |
-0.017 |
0.000 |
| y |
-0.017 |
-43.012 |
0.000 |
| z |
0.000 |
0.000 |
-36.622 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.519 |
-0.017 |
0.000 |
| y |
-0.017 |
-1.533 |
0.000 |
| z |
0.000 |
0.000 |
8.052 |
|
| Polar |
|---|
| 3z2-r2 | 16.104 |
|---|
| x2-y2 | -3.324 |
|---|
| xy | -0.017 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.864 |
0.457 |
0.000 |
| y |
0.457 |
9.049 |
0.000 |
| z |
0.000 |
0.000 |
13.147 |
<r2> (average value of r
2) Å
2
| <r2> |
193.505 |
| (<r2>)1/2 |
13.911 |