Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -153.702089 |
| Energy at 298.15K | -153.705460 |
| HF Energy | -153.702089 |
| Nuclear repulsion energy | 99.897123 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3404 |
3085 |
0.00 |
|
|
|
| 2 |
Ag |
1751 |
1587 |
0.00 |
|
|
|
| 3 |
Ag |
1206 |
1093 |
0.00 |
|
|
|
| 4 |
Ag |
1025 |
929 |
0.00 |
|
|
|
| 5 |
Au |
1014 |
920 |
0.00 |
|
|
|
| 6 |
Au |
599 |
543 |
0.00 |
|
|
|
| 7 |
B1g |
1015 |
920 |
0.00 |
|
|
|
| 8 |
B1u |
3393 |
3076 |
14.79 |
|
|
|
| 9 |
B1u |
1773 |
1607 |
10.71 |
|
|
|
| 10 |
B1u |
1148 |
1041 |
0.09 |
|
|
|
| 11 |
B2g |
704 |
638 |
0.00 |
|
|
|
| 12 |
B2u |
3370 |
3055 |
9.02 |
|
|
|
| 13 |
B2u |
1393 |
1263 |
41.58 |
|
|
|
| 14 |
B2u |
801 |
726 |
14.94 |
|
|
|
| 15 |
B3g |
3354 |
3040 |
0.00 |
|
|
|
| 16 |
B3g |
1298 |
1177 |
0.00 |
|
|
|
| 17 |
B3g |
922 |
836 |
0.00 |
|
|
|
| 18 |
B3u |
676 |
613 |
170.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14423.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 13073.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.657 |
0.781 |
| C2 |
0.000 |
0.657 |
-0.781 |
| C3 |
0.000 |
-0.657 |
0.781 |
| C4 |
0.000 |
-0.657 |
-0.781 |
| H5 |
0.000 |
1.416 |
1.535 |
| H6 |
0.000 |
1.416 |
-1.535 |
| H7 |
0.000 |
-1.416 |
1.535 |
| H8 |
0.000 |
-1.416 |
-1.535 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5627 | 1.3147 | 2.0421 | 1.0698 | 2.4377 | 2.2066 | 3.1091 |
C2 | 1.5627 | | 2.0421 | 1.3147 | 2.4377 | 1.0698 | 3.1091 | 2.2066 | C3 | 1.3147 | 2.0421 | | 1.5627 | 2.2066 | 3.1091 | 1.0698 | 2.4377 | C4 | 2.0421 | 1.3147 | 1.5627 | | 3.1091 | 2.2066 | 2.4377 | 1.0698 | H5 | 1.0698 | 2.4377 | 2.2066 | 3.1091 | | 3.0703 | 2.8329 | 4.1776 | H6 | 2.4377 | 1.0698 | 3.1091 | 2.2066 | 3.0703 | | 4.1776 | 2.8329 | H7 | 2.2066 | 3.1091 | 1.0698 | 2.4377 | 2.8329 | 4.1776 | | 3.0703 | H8 | 3.1091 | 2.2066 | 2.4377 | 1.0698 | 4.1776 | 2.8329 | 3.0703 | |
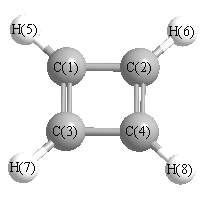 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
90.000 |
|
C1 |
C2 |
H6 |
134.800 |
| C1 |
C3 |
C4 |
90.000 |
|
C1 |
C3 |
H7 |
135.200 |
| C2 |
C1 |
C3 |
90.000 |
|
C2 |
C1 |
H5 |
134.800 |
| C2 |
C4 |
C3 |
90.000 |
|
C3 |
C1 |
H5 |
135.200 |
| C4 |
C2 |
H6 |
135.200 |
|
C4 |
C3 |
H7 |
134.800 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.131 |
|
|
|
| 2 |
C |
-0.131 |
|
|
|
| 3 |
C |
-0.131 |
|
|
|
| 4 |
C |
-0.131 |
|
|
|
| 5 |
H |
0.131 |
|
|
|
| 6 |
H |
0.131 |
|
|
|
| 7 |
H |
0.131 |
|
|
|
| 8 |
H |
0.131 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.291 |
0.000 |
0.000 |
| y |
0.000 |
-20.626 |
0.000 |
| z |
0.000 |
0.000 |
-22.487 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.734 |
0.000 |
0.000 |
| y |
0.000 |
4.263 |
0.000 |
| z |
0.000 |
0.000 |
1.471 |
|
| Polar |
|---|
| 3z2-r2 | 2.943 |
|---|
| x2-y2 | -6.665 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.139 |
0.000 |
0.000 |
| y |
0.000 |
7.542 |
0.000 |
| z |
0.000 |
0.000 |
6.054 |
<r2> (average value of r
2) Å
2
| <r2> |
57.145 |
| (<r2>)1/2 |
7.559 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -153.650816 |
| Energy at 298.15K | |
| HF Energy | -153.650816 |
| Nuclear repulsion energy | 100.523616 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3477 |
3152 |
0.00 |
|
|
|
| 2 |
Ag |
3308 |
2998 |
0.00 |
|
|
|
| 3 |
Ag |
1368 |
1240 |
0.00 |
|
|
|
| 4 |
Ag |
1035 |
938 |
0.00 |
|
|
|
| 5 |
B1g |
1219 |
1105 |
0.00 |
|
|
|
| 6 |
B1u |
3467 |
3143 |
17.85 |
|
|
|
| 7 |
B1u |
1568 |
1422 |
116.34 |
|
|
|
| 8 |
B1u |
1027 |
931 |
225.22 |
|
|
|
| 9 |
B2g |
440i |
399i |
0.00 |
|
|
|
| 10 |
B2u |
3314 |
3004 |
58.69 |
|
|
|
| 11 |
B2u |
1400 |
1269 |
184.83 |
|
|
|
| 12 |
B2u |
1036 |
939 |
25.01 |
|
|
|
| 13 |
B3g |
7636 |
6921 |
0.00 |
|
|
|
| 14 |
B3g |
1327 |
1203 |
0.00 |
|
|
|
| 15 |
B3g |
1139 |
1033 |
0.00 |
|
|
|
| 16 |
B3u |
1067 |
967 |
4.82 |
|
|
|
| 17 |
B3u |
604 |
547 |
131.02 |
|
|
|
| 18 |
B3u |
377i |
342i |
199.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16587.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 15034.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.063 |
| C2 |
0.000 |
0.947 |
0.000 |
| C3 |
0.000 |
-0.947 |
0.000 |
| C4 |
0.000 |
0.000 |
-1.063 |
| H5 |
0.000 |
0.000 |
2.129 |
| H6 |
0.000 |
2.020 |
0.000 |
| H7 |
0.000 |
-2.020 |
0.000 |
| H8 |
0.000 |
0.000 |
-2.129 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4242 | 1.4242 | 2.1268 | 1.0652 | 2.2831 | 2.2831 | 3.1920 |
C2 | 1.4242 | | 1.8947 | 1.4242 | 2.3299 | 1.0730 | 2.9677 | 2.3299 | C3 | 1.4242 | 1.8947 | | 1.4242 | 2.3299 | 2.9677 | 1.0730 | 2.3299 | C4 | 2.1268 | 1.4242 | 1.4242 | | 3.1920 | 2.2831 | 2.2831 | 1.0652 | H5 | 1.0652 | 2.3299 | 2.3299 | 3.1920 | | 2.9347 | 2.9347 | 4.2572 | H6 | 2.2831 | 1.0730 | 2.9677 | 2.2831 | 2.9347 | | 4.0407 | 2.9347 | H7 | 2.2831 | 2.9677 | 1.0730 | 2.2831 | 2.9347 | 4.0407 | | 2.9347 | H8 | 3.1920 | 2.3299 | 2.3299 | 1.0652 | 4.2572 | 2.9347 | 2.9347 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
96.608 |
|
C1 |
C2 |
H6 |
131.696 |
| C1 |
C3 |
C4 |
96.608 |
|
C1 |
C3 |
H7 |
131.696 |
| C2 |
C1 |
C3 |
83.392 |
|
C2 |
C1 |
H5 |
138.304 |
| C2 |
C4 |
C3 |
83.392 |
|
C3 |
C1 |
H5 |
138.304 |
| C4 |
C2 |
H6 |
131.696 |
|
C4 |
C3 |
H7 |
131.696 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.549 |
|
|
|
| 2 |
C |
0.278 |
|
|
|
| 3 |
C |
0.278 |
|
|
|
| 4 |
C |
-0.549 |
|
|
|
| 5 |
H |
0.137 |
|
|
|
| 6 |
H |
0.133 |
|
|
|
| 7 |
H |
0.133 |
|
|
|
| 8 |
H |
0.137 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.004 |
0.000 |
0.000 |
| y |
0.000 |
-17.847 |
0.000 |
| z |
0.000 |
0.000 |
-26.062 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.049 |
0.000 |
0.000 |
| y |
0.000 |
9.186 |
0.000 |
| z |
0.000 |
0.000 |
-3.136 |
|
| Polar |
|---|
| 3z2-r2 | -6.273 |
|---|
| x2-y2 | -10.156 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.540 |
0.000 |
0.000 |
| y |
0.000 |
6.579 |
0.000 |
| z |
0.000 |
0.000 |
8.021 |
<r2> (average value of r
2) Å
2
| <r2> |
56.506 |
| (<r2>)1/2 |
7.517 |
