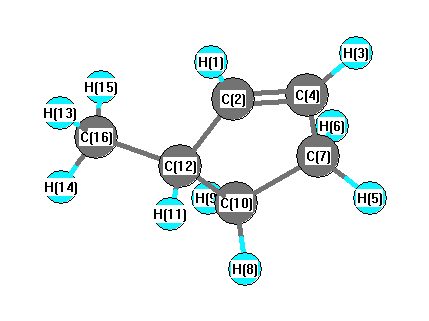
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3339 |
3026 |
25.89 |
|
|
|
| 2 |
A |
3311 |
3001 |
11.66 |
|
|
|
| 3 |
A |
3218 |
2917 |
97.54 |
|
|
|
| 4 |
A |
3209 |
2909 |
16.77 |
|
|
|
| 5 |
A |
3207 |
2907 |
54.23 |
|
|
|
| 6 |
A |
3191 |
2892 |
54.80 |
|
|
|
| 7 |
A |
3168 |
2872 |
30.36 |
|
|
|
| 8 |
A |
3150 |
2855 |
27.94 |
|
|
|
| 9 |
A |
3146 |
2851 |
58.73 |
|
|
|
| 10 |
A |
3116 |
2825 |
33.89 |
|
|
|
| 11 |
A |
1818 |
1648 |
2.62 |
|
|
|
| 12 |
A |
1628 |
1476 |
1.22 |
|
|
|
| 13 |
A |
1615 |
1464 |
4.14 |
|
|
|
| 14 |
A |
1613 |
1462 |
4.34 |
|
|
|
| 15 |
A |
1606 |
1456 |
0.74 |
|
|
|
| 16 |
A |
1534 |
1390 |
2.45 |
|
|
|
| 17 |
A |
1518 |
1376 |
7.82 |
|
|
|
| 18 |
A |
1465 |
1328 |
0.51 |
|
|
|
| 19 |
A |
1456 |
1320 |
0.76 |
|
|
|
| 20 |
A |
1432 |
1298 |
4.96 |
|
|
|
| 21 |
A |
1419 |
1286 |
0.97 |
|
|
|
| 22 |
A |
1340 |
1215 |
0.77 |
|
|
|
| 23 |
A |
1305 |
1183 |
0.75 |
|
|
|
| 24 |
A |
1234 |
1119 |
1.21 |
|
|
|
| 25 |
A |
1214 |
1101 |
3.48 |
|
|
|
| 26 |
A |
1201 |
1088 |
9.76 |
|
|
|
| 27 |
A |
1143 |
1036 |
1.85 |
|
|
|
| 28 |
A |
1093 |
991 |
0.30 |
|
|
|
| 29 |
A |
1067 |
967 |
1.14 |
|
|
|
| 30 |
A |
1042 |
944 |
1.58 |
|
|
|
| 31 |
A |
1007 |
912 |
0.95 |
|
|
|
| 32 |
A |
965 |
874 |
3.75 |
|
|
|
| 33 |
A |
917 |
831 |
2.79 |
|
|
|
| 34 |
A |
883 |
800 |
1.75 |
|
|
|
| 35 |
A |
815 |
739 |
38.04 |
|
|
|
| 36 |
A |
810 |
735 |
4.32 |
|
|
|
| 37 |
A |
630 |
571 |
4.41 |
|
|
|
| 38 |
A |
534 |
484 |
1.78 |
|
|
|
| 39 |
A |
366 |
332 |
0.06 |
|
|
|
| 40 |
A |
308 |
279 |
0.07 |
|
|
|
| 41 |
A |
258 |
234 |
0.03 |
|
|
|
| 42 |
A |
123 |
111 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33705.5 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 30550.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.078 |
|
|
|
| 2 |
C |
-0.198 |
|
|
|
| 3 |
H |
0.079 |
|
|
|
| 4 |
C |
-0.139 |
|
|
|
| 5 |
H |
0.042 |
|
|
|
| 6 |
H |
0.047 |
|
|
|
| 7 |
C |
-0.002 |
|
|
|
| 8 |
H |
0.050 |
|
|
|
| 9 |
H |
0.042 |
|
|
|
| 10 |
C |
-0.150 |
|
|
|
| 11 |
H |
0.016 |
|
|
|
| 12 |
C |
0.178 |
|
|
|
| 13 |
H |
0.056 |
|
|
|
| 14 |
H |
0.060 |
|
|
|
| 15 |
H |
0.049 |
|
|
|
| 16 |
C |
-0.206 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.006 |
-0.184 |
0.160 |
0.244 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.071 |
-0.293 |
-0.719 |
| y |
-0.293 |
-38.204 |
-0.116 |
| z |
-0.719 |
-0.116 |
-39.606 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.834 |
-0.293 |
-0.719 |
| y |
-0.293 |
0.635 |
-0.116 |
| z |
-0.719 |
-0.116 |
-1.469 |
|
| Polar |
|---|
| 3z2-r2 | -2.937 |
|---|
| x2-y2 | 0.133 |
|---|
| xy | -0.293 |
|---|
| xz | -0.719 |
|---|
| yz | -0.116 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.708 |
-0.358 |
-0.358 |
| y |
-0.358 |
9.482 |
-0.014 |
| z |
-0.358 |
-0.014 |
7.962 |
<r2> (average value of r
2) Å
2
| <r2> |
163.794 |
| (<r2>)1/2 |
12.798 |