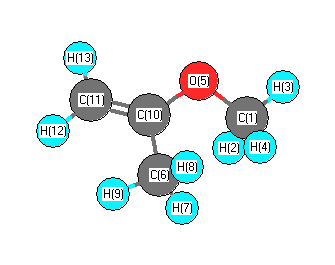
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3394 |
3076 |
10.68 |
|
|
|
| 2 |
A |
3307 |
2997 |
1.28 |
|
|
|
| 3 |
A |
3265 |
2960 |
20.19 |
|
|
|
| 4 |
A |
3262 |
2957 |
42.22 |
|
|
|
| 5 |
A |
3216 |
2915 |
16.53 |
|
|
|
| 6 |
A |
3211 |
2911 |
66.78 |
|
|
|
| 7 |
A |
3168 |
2871 |
31.40 |
|
|
|
| 8 |
A |
3154 |
2859 |
47.37 |
|
|
|
| 9 |
A |
1855 |
1681 |
156.76 |
|
|
|
| 10 |
A |
1632 |
1479 |
5.00 |
|
|
|
| 11 |
A |
1625 |
1473 |
9.08 |
|
|
|
| 12 |
A |
1614 |
1463 |
6.38 |
|
|
|
| 13 |
A |
1605 |
1454 |
4.90 |
|
|
|
| 14 |
A |
1597 |
1448 |
7.08 |
|
|
|
| 15 |
A |
1554 |
1409 |
7.68 |
|
|
|
| 16 |
A |
1527 |
1384 |
43.23 |
|
|
|
| 17 |
A |
1419 |
1286 |
186.80 |
|
|
|
| 18 |
A |
1315 |
1192 |
1.39 |
|
|
|
| 19 |
A |
1282 |
1162 |
8.40 |
|
|
|
| 20 |
A |
1234 |
1118 |
149.44 |
|
|
|
| 21 |
A |
1163 |
1055 |
3.89 |
|
|
|
| 22 |
A |
1109 |
1005 |
3.84 |
|
|
|
| 23 |
A |
1014 |
919 |
17.76 |
|
|
|
| 24 |
A |
1000 |
906 |
56.64 |
|
|
|
| 25 |
A |
872 |
790 |
9.09 |
|
|
|
| 26 |
A |
820 |
743 |
4.55 |
|
|
|
| 27 |
A |
596 |
540 |
7.09 |
|
|
|
| 28 |
A |
516 |
468 |
6.24 |
|
|
|
| 29 |
A |
452 |
410 |
1.23 |
|
|
|
| 30 |
A |
323 |
293 |
1.77 |
|
|
|
| 31 |
A |
246 |
223 |
1.72 |
|
|
|
| 32 |
A |
189 |
171 |
1.36 |
|
|
|
| 33 |
A |
75 |
68 |
7.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26305.6 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 23843.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.014 |
|
|
|
| 2 |
H |
0.061 |
|
|
|
| 3 |
H |
0.083 |
|
|
|
| 4 |
H |
0.056 |
|
|
|
| 5 |
O |
-0.398 |
|
|
|
| 6 |
C |
-0.146 |
|
|
|
| 7 |
H |
0.063 |
|
|
|
| 8 |
H |
0.073 |
|
|
|
| 9 |
H |
0.074 |
|
|
|
| 10 |
C |
0.196 |
|
|
|
| 11 |
C |
-0.209 |
|
|
|
| 12 |
H |
0.062 |
|
|
|
| 13 |
H |
0.072 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.673 |
1.456 |
0.799 |
1.792 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.625 |
-0.033 |
0.544 |
| y |
-0.033 |
-31.452 |
-0.655 |
| z |
0.544 |
-0.655 |
-33.872 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.037 |
-0.033 |
0.544 |
| y |
-0.033 |
-0.203 |
-0.655 |
| z |
0.544 |
-0.655 |
-3.834 |
|
| Polar |
|---|
| 3z2-r2 | -7.667 |
|---|
| x2-y2 | 2.827 |
|---|
| xy | -0.033 |
|---|
| xz | 0.544 |
|---|
| yz | -0.655 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.018 |
1.130 |
-0.206 |
| y |
1.130 |
7.995 |
-0.223 |
| z |
-0.206 |
-0.223 |
5.946 |
<r2> (average value of r
2) Å
2
| <r2> |
126.995 |
| (<r2>)1/2 |
11.269 |