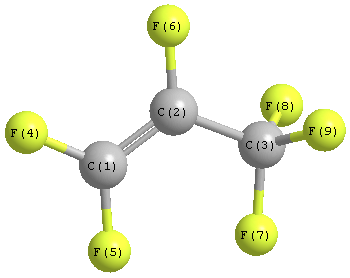
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2022 |
1833 |
223.69 |
35.72 |
0.24 |
0.39 |
| 2 |
A' |
1550 |
1405 |
296.25 |
1.89 |
0.18 |
0.31 |
| 3 |
A' |
1484 |
1345 |
217.93 |
1.95 |
0.12 |
0.22 |
| 4 |
A' |
1384 |
1254 |
185.40 |
1.42 |
0.54 |
0.71 |
| 5 |
A' |
1336 |
1211 |
236.37 |
0.61 |
0.40 |
0.57 |
| 6 |
A' |
1148 |
1041 |
340.84 |
0.62 |
0.58 |
0.73 |
| 7 |
A' |
848 |
769 |
20.29 |
7.47 |
0.02 |
0.04 |
| 8 |
A' |
725 |
657 |
18.68 |
2.03 |
0.25 |
0.40 |
| 9 |
A' |
664 |
602 |
1.05 |
0.98 |
0.70 |
0.83 |
| 10 |
A' |
566 |
513 |
4.14 |
0.55 |
0.58 |
0.73 |
| 11 |
A' |
406 |
368 |
0.13 |
1.59 |
0.73 |
0.84 |
| 12 |
A' |
394 |
357 |
1.86 |
2.59 |
0.27 |
0.43 |
| 13 |
A' |
281 |
255 |
1.49 |
0.03 |
0.18 |
0.30 |
| 14 |
A' |
187 |
170 |
1.69 |
0.09 |
0.46 |
0.63 |
| 15 |
A" |
1346 |
1220 |
342.59 |
2.84 |
0.75 |
0.86 |
| 16 |
A" |
780 |
707 |
5.64 |
8.39 |
0.75 |
0.86 |
| 17 |
A" |
647 |
587 |
0.39 |
2.75 |
0.75 |
0.86 |
| 18 |
A" |
520 |
472 |
4.44 |
0.32 |
0.75 |
0.86 |
| 19 |
A" |
278 |
252 |
1.21 |
0.32 |
0.75 |
0.86 |
| 20 |
A" |
146 |
132 |
0.42 |
0.46 |
0.75 |
0.86 |
| 21 |
A" |
47 |
42 |
0.01 |
0.22 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8379.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 7595.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.509 |
|
|
|
| 2 |
C |
0.103 |
|
|
|
| 3 |
C |
0.714 |
|
|
|
| 4 |
F |
-0.204 |
|
|
|
| 5 |
F |
-0.201 |
|
|
|
| 6 |
F |
-0.241 |
|
|
|
| 7 |
F |
-0.226 |
|
|
|
| 8 |
F |
-0.227 |
|
|
|
| 9 |
F |
-0.227 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.290 |
1.066 |
0.000 |
1.104 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.926 |
0.129 |
0.000 |
| y |
0.129 |
-47.668 |
0.000 |
| z |
0.000 |
0.000 |
-45.414 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.385 |
0.129 |
0.000 |
| y |
0.129 |
-0.499 |
0.000 |
| z |
0.000 |
0.000 |
2.884 |
|
| Polar |
|---|
| 3z2-r2 | 5.767 |
|---|
| x2-y2 | -1.258 |
|---|
| xy | 0.129 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.008 |
0.419 |
0.000 |
| y |
0.419 |
6.546 |
0.000 |
| z |
0.000 |
0.000 |
4.053 |
<r2> (average value of r
2) Å
2
| <r2> |
287.997 |
| (<r2>)1/2 |
16.970 |