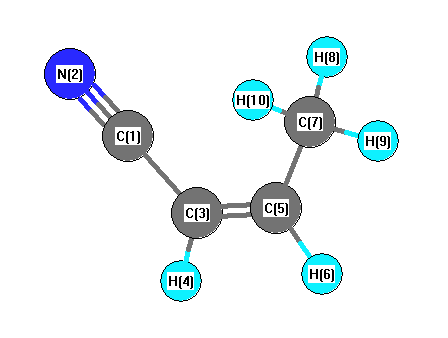
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3353 |
3039 |
3.85 |
|
|
|
| 2 |
A' |
3308 |
2998 |
10.64 |
|
|
|
| 3 |
A' |
3265 |
2959 |
5.84 |
|
|
|
| 4 |
A' |
3166 |
2870 |
15.86 |
|
|
|
| 5 |
A' |
2574 |
2333 |
42.50 |
|
|
|
| 6 |
A' |
1837 |
1665 |
25.34 |
|
|
|
| 7 |
A' |
1601 |
1451 |
15.57 |
|
|
|
| 8 |
A' |
1554 |
1409 |
1.38 |
|
|
|
| 9 |
A' |
1517 |
1375 |
2.56 |
|
|
|
| 10 |
A' |
1358 |
1231 |
1.69 |
|
|
|
| 11 |
A' |
1215 |
1101 |
0.21 |
|
|
|
| 12 |
A' |
1017 |
922 |
12.85 |
|
|
|
| 13 |
A' |
944 |
856 |
0.74 |
|
|
|
| 14 |
A' |
716 |
649 |
1.61 |
|
|
|
| 15 |
A' |
441 |
399 |
0.54 |
|
|
|
| 16 |
A' |
178 |
161 |
5.74 |
|
|
|
| 17 |
A" |
3213 |
2912 |
17.25 |
|
|
|
| 18 |
A" |
1600 |
1450 |
7.74 |
|
|
|
| 19 |
A" |
1188 |
1077 |
1.05 |
|
|
|
| 20 |
A" |
1092 |
989 |
2.68 |
|
|
|
| 21 |
A" |
830 |
753 |
46.42 |
|
|
|
| 22 |
A" |
585 |
530 |
1.35 |
|
|
|
| 23 |
A" |
323 |
293 |
2.82 |
|
|
|
| 24 |
A" |
165 |
150 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18518.6 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 16785.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.111 |
|
|
|
| 2 |
N |
-0.135 |
|
|
|
| 3 |
C |
-0.023 |
|
|
|
| 4 |
H |
0.100 |
|
|
|
| 5 |
C |
-0.003 |
|
|
|
| 6 |
H |
0.095 |
|
|
|
| 7 |
C |
-0.144 |
|
|
|
| 8 |
H |
0.080 |
|
|
|
| 9 |
H |
0.071 |
|
|
|
| 10 |
H |
0.071 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.403 |
1.259 |
0.000 |
4.580 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.116 |
-2.619 |
0.000 |
| y |
-2.619 |
-27.643 |
0.000 |
| z |
0.000 |
0.000 |
-31.327 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.631 |
-2.619 |
0.000 |
| y |
-2.619 |
6.078 |
0.000 |
| z |
0.000 |
0.000 |
0.553 |
|
| Polar |
|---|
| 3z2-r2 | 1.105 |
|---|
| x2-y2 | -8.473 |
|---|
| xy | -2.619 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.295 |
-0.229 |
0.000 |
| y |
-0.229 |
6.756 |
0.000 |
| z |
0.000 |
0.000 |
5.007 |
<r2> (average value of r
2) Å
2
| <r2> |
125.496 |
| (<r2>)1/2 |
11.202 |