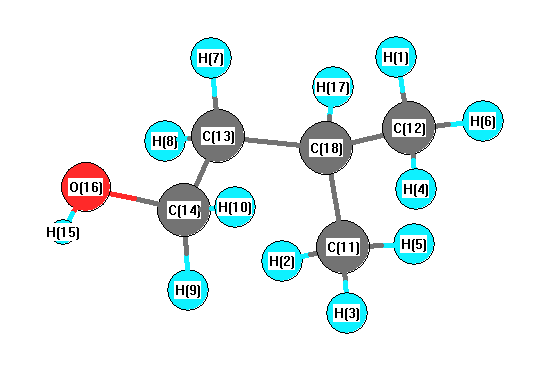
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
4184 |
3793 |
58.00 |
|
|
|
| 2 |
A' |
3217 |
2916 |
122.61 |
|
|
|
| 3 |
A' |
3213 |
2912 |
79.98 |
|
|
|
| 4 |
A' |
3168 |
2872 |
15.97 |
|
|
|
| 5 |
A' |
3159 |
2863 |
71.07 |
|
|
|
| 6 |
A' |
3146 |
2852 |
16.57 |
|
|
|
| 7 |
A' |
3137 |
2844 |
18.59 |
|
|
|
| 8 |
A' |
1657 |
1502 |
2.36 |
|
|
|
| 9 |
A' |
1628 |
1476 |
9.88 |
|
|
|
| 10 |
A' |
1627 |
1475 |
5.63 |
|
|
|
| 11 |
A' |
1615 |
1464 |
1.84 |
|
|
|
| 12 |
A' |
1592 |
1443 |
4.81 |
|
|
|
| 13 |
A' |
1548 |
1403 |
2.87 |
|
|
|
| 14 |
A' |
1497 |
1356 |
1.96 |
|
|
|
| 15 |
A' |
1464 |
1327 |
23.82 |
|
|
|
| 16 |
A' |
1342 |
1217 |
56.70 |
|
|
|
| 17 |
A' |
1302 |
1181 |
1.79 |
|
|
|
| 18 |
A' |
1210 |
1097 |
2.27 |
|
|
|
| 19 |
A' |
1168 |
1059 |
115.77 |
|
|
|
| 20 |
A' |
1068 |
968 |
2.67 |
|
|
|
| 21 |
A' |
1018 |
923 |
9.31 |
|
|
|
| 22 |
A' |
841 |
762 |
4.88 |
|
|
|
| 23 |
A' |
596 |
541 |
1.76 |
|
|
|
| 24 |
A' |
435 |
394 |
6.30 |
|
|
|
| 25 |
A' |
349 |
317 |
5.10 |
|
|
|
| 26 |
A' |
273 |
247 |
0.02 |
|
|
|
| 27 |
A' |
226 |
205 |
1.98 |
|
|
|
| 28 |
A" |
3211 |
2910 |
64.75 |
|
|
|
| 29 |
A" |
3209 |
2908 |
9.96 |
|
|
|
| 30 |
A" |
3201 |
2901 |
46.03 |
|
|
|
| 31 |
A" |
3167 |
2871 |
37.96 |
|
|
|
| 32 |
A" |
3151 |
2856 |
22.73 |
|
|
|
| 33 |
A" |
1616 |
1465 |
0.94 |
|
|
|
| 34 |
A" |
1604 |
1454 |
0.40 |
|
|
|
| 35 |
A" |
1531 |
1388 |
6.66 |
|
|
|
| 36 |
A" |
1488 |
1348 |
1.11 |
|
|
|
| 37 |
A" |
1436 |
1301 |
0.00 |
|
|
|
| 38 |
A" |
1358 |
1231 |
0.37 |
|
|
|
| 39 |
A" |
1275 |
1156 |
6.57 |
|
|
|
| 40 |
A" |
1078 |
977 |
0.12 |
|
|
|
| 41 |
A" |
1035 |
938 |
0.00 |
|
|
|
| 42 |
A" |
1000 |
906 |
0.28 |
|
|
|
| 43 |
A" |
850 |
771 |
0.00 |
|
|
|
| 44 |
A" |
393 |
357 |
0.02 |
|
|
|
| 45 |
A" |
285 |
258 |
116.95 |
|
|
|
| 46 |
A" |
254 |
230 |
3.69 |
|
|
|
| 47 |
A" |
115 |
104 |
0.01 |
|
|
|
| 48 |
A" |
77 |
70 |
10.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38506.0 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 34901.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.063 |
|
|
|
| 2 |
H |
0.063 |
|
|
|
| 3 |
H |
0.045 |
|
|
|
| 4 |
H |
0.045 |
|
|
|
| 5 |
H |
0.061 |
|
|
|
| 6 |
H |
0.061 |
|
|
|
| 7 |
H |
0.049 |
|
|
|
| 8 |
H |
0.049 |
|
|
|
| 9 |
H |
0.037 |
|
|
|
| 10 |
H |
0.037 |
|
|
|
| 11 |
C |
-0.225 |
|
|
|
| 12 |
C |
-0.225 |
|
|
|
| 13 |
C |
-0.063 |
|
|
|
| 14 |
C |
0.102 |
|
|
|
| 15 |
H |
0.193 |
|
|
|
| 16 |
O |
-0.412 |
|
|
|
| 17 |
H |
0.009 |
|
|
|
| 18 |
C |
0.111 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.316 |
-1.462 |
0.000 |
1.496 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.993 |
2.875 |
0.000 |
| y |
2.875 |
-44.439 |
0.000 |
| z |
0.000 |
0.000 |
-40.634 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.544 |
2.875 |
0.000 |
| y |
2.875 |
-6.626 |
0.000 |
| z |
0.000 |
0.000 |
-0.918 |
|
| Polar |
|---|
| 3z2-r2 | -1.835 |
|---|
| x2-y2 | 9.446 |
|---|
| xy | 2.875 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.621 |
-0.718 |
0.000 |
| y |
-0.718 |
9.009 |
0.000 |
| z |
0.000 |
0.000 |
8.941 |
<r2> (average value of r
2) Å
2
| <r2> |
223.135 |
| (<r2>)1/2 |
14.938 |