Jump to
S1C2
Energy calculated at B2PLYP=FULLultrafine/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -722.624006 |
| Energy at 298.15K | |
| HF Energy | -722.410895 |
| Nuclear repulsion energy | 91.151494 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
284 |
284 |
0.00 |
284.59 |
0.11 |
0.20 |
| 2 |
Σu |
433 |
433 |
112.32 |
0.00 |
0.00 |
0.00 |
| 3 |
Πu |
43i |
43i |
58.93 |
0.00 |
0.00 |
0.00 |
| 3 |
Πu |
43i |
43i |
58.93 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 315.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 315.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/Def2TZVPP
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.000 |
| Na2 |
0.000 |
0.000 |
2.395 |
| Na3 |
0.000 |
0.000 |
-2.395 |
Atom - Atom Distances (Å)
| |
S1 |
Na2 |
Na3 |
| S1 | | 2.3948 | 2.3948 |
Na2 | 2.3948 | | 4.7895 | Na3 | 2.3948 | 4.7895 | |
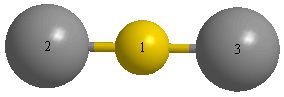 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Na2 |
S1 |
Na3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULLultrafine/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -722.626190 |
| Energy at 298.15K | |
| HF Energy | -722.411648 |
| Nuclear repulsion energy | 92.200489 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
313 |
313 |
12.39 |
387.80 |
0.08 |
0.15 |
| 2 |
A1 |
59 |
59 |
29.61 |
12.65 |
0.74 |
0.85 |
| 3 |
B2 |
389 |
389 |
79.84 |
18.09 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 380.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 380.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/Def2TZVPP
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.717 |
| Na2 |
0.000 |
2.084 |
-0.521 |
| Na3 |
0.000 |
-2.084 |
-0.521 |
Atom - Atom Distances (Å)
| |
S1 |
Na2 |
Na3 |
| S1 | | 2.4242 | 2.4242 |
Na2 | 2.4242 | | 4.1680 | Na3 | 2.4242 | 4.1680 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Na2 |
S1 |
Na3 |
118.558 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
