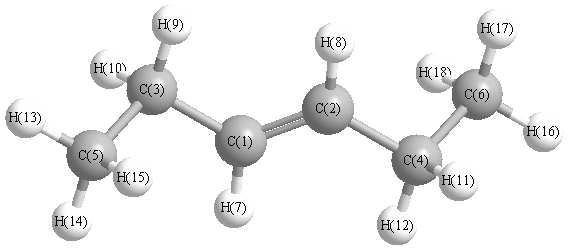
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3255 |
2950 |
0.00 |
|
|
|
| 2 |
Ag |
3220 |
2919 |
0.00 |
|
|
|
| 3 |
Ag |
3214 |
2914 |
0.00 |
|
|
|
| 4 |
Ag |
3186 |
2888 |
0.00 |
|
|
|
| 5 |
Ag |
3156 |
2861 |
0.00 |
|
|
|
| 6 |
Ag |
3145 |
2851 |
0.00 |
|
|
|
| 7 |
Ag |
1877 |
1701 |
0.00 |
|
|
|
| 8 |
Ag |
1621 |
1470 |
0.00 |
|
|
|
| 9 |
Ag |
1612 |
1461 |
0.00 |
|
|
|
| 10 |
Ag |
1604 |
1453 |
0.00 |
|
|
|
| 11 |
Ag |
1533 |
1389 |
0.00 |
|
|
|
| 12 |
Ag |
1469 |
1331 |
0.00 |
|
|
|
| 13 |
Ag |
1441 |
1306 |
0.00 |
|
|
|
| 14 |
Ag |
1390 |
1260 |
0.00 |
|
|
|
| 15 |
Ag |
1280 |
1160 |
0.00 |
|
|
|
| 16 |
Ag |
1190 |
1078 |
0.00 |
|
|
|
| 17 |
Ag |
1071 |
971 |
0.00 |
|
|
|
| 18 |
Ag |
972 |
881 |
0.00 |
|
|
|
| 19 |
Ag |
929 |
842 |
0.00 |
|
|
|
| 20 |
Ag |
811 |
735 |
0.00 |
|
|
|
| 21 |
Ag |
500 |
453 |
0.00 |
|
|
|
| 22 |
Ag |
343 |
311 |
0.00 |
|
|
|
| 23 |
Ag |
253 |
230 |
0.00 |
|
|
|
| 24 |
Ag |
142 |
129 |
0.00 |
|
|
|
| 25 |
Au |
3261 |
2956 |
76.32 |
|
|
|
| 26 |
Au |
3220 |
2919 |
114.38 |
|
|
|
| 27 |
Au |
3214 |
2913 |
119.41 |
|
|
|
| 28 |
Au |
3185 |
2887 |
9.20 |
|
|
|
| 29 |
Au |
3155 |
2860 |
80.65 |
|
|
|
| 30 |
Au |
3145 |
2850 |
77.21 |
|
|
|
| 31 |
Au |
1622 |
1470 |
9.26 |
|
|
|
| 32 |
Au |
1612 |
1462 |
11.11 |
|
|
|
| 33 |
Au |
1603 |
1453 |
5.19 |
|
|
|
| 34 |
Au |
1533 |
1390 |
4.22 |
|
|
|
| 35 |
Au |
1506 |
1365 |
7.30 |
|
|
|
| 36 |
Au |
1434 |
1300 |
13.94 |
|
|
|
| 37 |
Au |
1370 |
1242 |
0.59 |
|
|
|
| 38 |
Au |
1195 |
1084 |
7.85 |
|
|
|
| 39 |
Au |
1172 |
1062 |
2.97 |
|
|
|
| 40 |
Au |
1115 |
1011 |
7.35 |
|
|
|
| 41 |
Au |
1101 |
998 |
37.22 |
|
|
|
| 42 |
Au |
960 |
870 |
4.13 |
|
|
|
| 43 |
Au |
851 |
772 |
3.44 |
|
|
|
| 44 |
Au |
520 |
471 |
1.80 |
|
|
|
| 45 |
Au |
347 |
314 |
1.00 |
|
|
|
| 46 |
Au |
207 |
187 |
0.28 |
|
|
|
| 47 |
Au |
132 |
120 |
0.84 |
|
|
|
| 48 |
Au |
63 |
57 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38367.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 34776.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.106 |
|
|
|
| 2 |
C |
-0.106 |
|
|
|
| 3 |
C |
-0.000 |
|
|
|
| 4 |
C |
-0.000 |
|
|
|
| 5 |
C |
-0.167 |
|
|
|
| 6 |
C |
-0.167 |
|
|
|
| 7 |
H |
0.043 |
|
|
|
| 8 |
H |
0.043 |
|
|
|
| 9 |
H |
0.037 |
|
|
|
| 10 |
H |
0.037 |
|
|
|
| 11 |
H |
0.037 |
|
|
|
| 12 |
H |
0.037 |
|
|
|
| 13 |
H |
0.057 |
|
|
|
| 14 |
H |
0.053 |
|
|
|
| 15 |
H |
0.046 |
|
|
|
| 16 |
H |
0.057 |
|
|
|
| 17 |
H |
0.053 |
|
|
|
| 18 |
H |
0.046 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.148 |
-0.034 |
0.120 |
| y |
-0.034 |
-42.297 |
-0.650 |
| z |
0.120 |
-0.650 |
-39.041 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.521 |
-0.034 |
0.120 |
| y |
-0.034 |
-3.202 |
-0.650 |
| z |
0.120 |
-0.650 |
1.682 |
|
| Polar |
|---|
| 3z2-r2 | 3.363 |
|---|
| x2-y2 | 3.149 |
|---|
| xy | -0.034 |
|---|
| xz | 0.120 |
|---|
| yz | -0.650 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.927 |
-0.352 |
-1.581 |
| y |
-0.352 |
9.049 |
0.987 |
| z |
-1.581 |
0.987 |
13.294 |
<r2> (average value of r
2) Å
2
| <r2> |
288.940 |
| (<r2>)1/2 |
16.998 |