Jump to
S1C2
Energy calculated at mPW1PW91/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -166.541196 |
| Energy at 298.15K | |
| HF Energy | -166.541196 |
| Nuclear repulsion energy | 49.044099 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
4057 |
4057 |
53.68 |
|
|
|
| 2 |
A1 |
747 |
747 |
6.11 |
|
|
|
| 3 |
A1 |
583 |
583 |
175.53 |
|
|
|
| 4 |
A1 |
295 |
295 |
9.62 |
|
|
|
| 5 |
A2 |
202i |
202i |
0.00 |
|
|
|
| 6 |
B1 |
345 |
345 |
63.38 |
|
|
|
| 7 |
B2 |
4055 |
4055 |
172.97 |
|
|
|
| 8 |
B2 |
1537 |
1537 |
408.54 |
|
|
|
| 9 |
B2 |
476 |
476 |
270.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5946.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 5946.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/Def2TZVPP
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
-0.000 |
| O2 |
0.000 |
1.423 |
0.076 |
| O3 |
0.000 |
-1.423 |
0.076 |
| H4 |
0.000 |
2.082 |
-0.606 |
| H5 |
0.000 |
-2.082 |
-0.606 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.4251 | 1.4251 | 2.1684 | 2.1684 |
O2 | 1.4251 | | 2.8462 | 0.9482 | 3.5709 | O3 | 1.4251 | 2.8462 | | 3.5709 | 0.9482 | H4 | 2.1684 | 0.9482 | 3.5709 | | 4.1642 | H5 | 2.1684 | 3.5709 | 0.9482 | 4.1642 | |
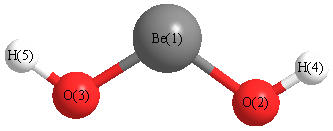 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
130.978 |
|
Be1 |
O3 |
H5 |
130.978 |
| O2 |
Be1 |
O3 |
173.895 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
0.321 |
|
|
|
| 2 |
O |
-0.313 |
|
|
|
| 3 |
O |
-0.313 |
|
|
|
| 4 |
H |
0.153 |
|
|
|
| 5 |
H |
0.153 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.342 |
2.342 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.780 |
0.000 |
0.000 |
| y |
0.000 |
-14.601 |
0.000 |
| z |
0.000 |
0.000 |
-14.574 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.192 |
0.000 |
0.000 |
| y |
0.000 |
1.076 |
0.000 |
| z |
0.000 |
0.000 |
1.116 |
|
| Polar |
|---|
| 3z2-r2 | 2.233 |
|---|
| x2-y2 | -2.178 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.563 |
0.000 |
0.000 |
| y |
0.000 |
3.857 |
0.000 |
| z |
0.000 |
0.000 |
2.677 |
<r2> (average value of r
2) Å
2
| <r2> |
51.467 |
| (<r2>)1/2 |
7.174 |
Jump to
S1C1
Energy calculated at mPW1PW91/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -166.541887 |
| Energy at 298.15K | -166.543342 |
| HF Energy | -166.541887 |
| Nuclear repulsion energy | 49.028855 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4043 |
4043 |
25.07 |
|
|
|
| 2 |
A |
742 |
742 |
1.78 |
|
|
|
| 3 |
A |
573 |
573 |
91.00 |
|
|
|
| 4 |
A |
314 |
314 |
37.79 |
|
|
|
| 5 |
A |
195 |
195 |
127.67 |
|
|
|
| 6 |
B |
4041 |
4041 |
187.42 |
|
|
|
| 7 |
B |
1538 |
1538 |
399.57 |
|
|
|
| 8 |
B |
569 |
569 |
337.08 |
|
|
|
| 9 |
B |
306 |
306 |
71.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6160.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 6160.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/Def2TZVPP
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
-0.001 |
| O2 |
0.000 |
1.425 |
-0.052 |
| O3 |
0.000 |
-1.425 |
-0.052 |
| H4 |
0.548 |
2.039 |
0.421 |
| H5 |
-0.548 |
-2.039 |
0.421 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.4263 | 1.4263 | 2.1531 | 2.1531 |
O2 | 1.4263 | | 2.8509 | 0.9491 | 3.5393 | O3 | 1.4263 | 2.8509 | | 3.5393 | 0.9491 | H4 | 2.1531 | 0.9491 | 3.5393 | | 4.2228 | H5 | 2.1531 | 3.5393 | 0.9491 | 4.2228 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
128.917 |
|
Be1 |
O3 |
H5 |
128.917 |
| O2 |
Be1 |
O3 |
175.892 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
0.332 |
|
|
|
| 2 |
O |
-0.322 |
|
|
|
| 3 |
O |
-0.322 |
|
|
|
| 4 |
H |
0.156 |
|
|
|
| 5 |
H |
0.156 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.629 |
1.629 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.130 |
4.390 |
0.000 |
| y |
4.390 |
-15.381 |
0.000 |
| z |
0.000 |
0.000 |
-15.733 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.427 |
4.390 |
0.000 |
| y |
4.390 |
0.050 |
0.000 |
| z |
0.000 |
0.000 |
-0.477 |
|
| Polar |
|---|
| 3z2-r2 | -0.954 |
|---|
| x2-y2 | 0.251 |
|---|
| xy | 4.390 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.636 |
0.113 |
0.000 |
| y |
0.113 |
3.846 |
0.000 |
| z |
0.000 |
0.000 |
2.621 |
<r2> (average value of r
2) Å
2
| <r2> |
51.451 |
| (<r2>)1/2 |
7.173 |
