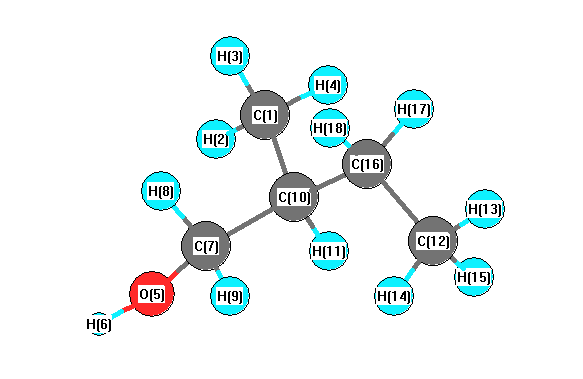
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4187 |
3795 |
61.87 |
|
|
|
| 2 |
A |
3249 |
2945 |
29.52 |
|
|
|
| 3 |
A |
3225 |
2923 |
82.80 |
|
|
|
| 4 |
A |
3212 |
2911 |
77.61 |
|
|
|
| 5 |
A |
3206 |
2906 |
58.86 |
|
|
|
| 6 |
A |
3183 |
2885 |
70.17 |
|
|
|
| 7 |
A |
3174 |
2877 |
31.67 |
|
|
|
| 8 |
A |
3158 |
2863 |
12.15 |
|
|
|
| 9 |
A |
3156 |
2860 |
83.45 |
|
|
|
| 10 |
A |
3147 |
2852 |
11.38 |
|
|
|
| 11 |
A |
3138 |
2844 |
7.91 |
|
|
|
| 12 |
A |
3122 |
2829 |
36.70 |
|
|
|
| 13 |
A |
1650 |
1495 |
2.90 |
|
|
|
| 14 |
A |
1628 |
1476 |
6.28 |
|
|
|
| 15 |
A |
1623 |
1471 |
8.52 |
|
|
|
| 16 |
A |
1618 |
1466 |
3.86 |
|
|
|
| 17 |
A |
1614 |
1463 |
3.34 |
|
|
|
| 18 |
A |
1605 |
1455 |
1.62 |
|
|
|
| 19 |
A |
1587 |
1439 |
3.85 |
|
|
|
| 20 |
A |
1541 |
1397 |
1.46 |
|
|
|
| 21 |
A |
1535 |
1391 |
4.72 |
|
|
|
| 22 |
A |
1517 |
1375 |
0.67 |
|
|
|
| 23 |
A |
1485 |
1346 |
4.80 |
|
|
|
| 24 |
A |
1446 |
1311 |
1.63 |
|
|
|
| 25 |
A |
1413 |
1280 |
10.07 |
|
|
|
| 26 |
A |
1382 |
1253 |
5.26 |
|
|
|
| 27 |
A |
1347 |
1221 |
51.67 |
|
|
|
| 28 |
A |
1277 |
1157 |
0.35 |
|
|
|
| 29 |
A |
1276 |
1156 |
8.34 |
|
|
|
| 30 |
A |
1208 |
1095 |
2.97 |
|
|
|
| 31 |
A |
1171 |
1062 |
108.54 |
|
|
|
| 32 |
A |
1105 |
1001 |
0.59 |
|
|
|
| 33 |
A |
1100 |
997 |
7.49 |
|
|
|
| 34 |
A |
1049 |
951 |
3.40 |
|
|
|
| 35 |
A |
1001 |
908 |
1.10 |
|
|
|
| 36 |
A |
968 |
878 |
0.48 |
|
|
|
| 37 |
A |
880 |
798 |
3.46 |
|
|
|
| 38 |
A |
815 |
739 |
4.19 |
|
|
|
| 39 |
A |
527 |
478 |
8.01 |
|
|
|
| 40 |
A |
475 |
430 |
1.42 |
|
|
|
| 41 |
A |
413 |
375 |
1.35 |
|
|
|
| 42 |
A |
298 |
270 |
4.18 |
|
|
|
| 43 |
A |
283 |
256 |
26.56 |
|
|
|
| 44 |
A |
256 |
232 |
89.16 |
|
|
|
| 45 |
A |
240 |
217 |
4.15 |
|
|
|
| 46 |
A |
232 |
211 |
1.21 |
|
|
|
| 47 |
A |
122 |
110 |
4.33 |
|
|
|
| 48 |
A |
69 |
63 |
4.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38455.2 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 34855.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.219 |
|
|
|
| 2 |
H |
0.087 |
|
|
|
| 3 |
H |
0.041 |
|
|
|
| 4 |
H |
0.054 |
|
|
|
| 5 |
O |
-0.414 |
|
|
|
| 6 |
H |
0.191 |
|
|
|
| 7 |
C |
0.086 |
|
|
|
| 8 |
H |
0.034 |
|
|
|
| 9 |
H |
0.046 |
|
|
|
| 10 |
C |
0.130 |
|
|
|
| 11 |
H |
-0.000 |
|
|
|
| 12 |
C |
-0.193 |
|
|
|
| 13 |
H |
0.062 |
|
|
|
| 14 |
H |
0.043 |
|
|
|
| 15 |
H |
0.051 |
|
|
|
| 16 |
C |
-0.076 |
|
|
|
| 17 |
H |
0.047 |
|
|
|
| 18 |
H |
0.029 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.510 |
-0.985 |
1.032 |
1.515 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.739 |
3.211 |
-2.569 |
| y |
3.211 |
-37.906 |
-1.619 |
| z |
-2.569 |
-1.619 |
-40.244 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.665 |
3.211 |
-2.569 |
| y |
3.211 |
2.585 |
-1.619 |
| z |
-2.569 |
-1.619 |
-0.921 |
|
| Polar |
|---|
| 3z2-r2 | -1.841 |
|---|
| x2-y2 | -2.833 |
|---|
| xy | 3.211 |
|---|
| xz | -2.569 |
|---|
| yz | -1.619 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.051 |
-0.156 |
-0.017 |
| y |
-0.156 |
9.342 |
-0.045 |
| z |
-0.017 |
-0.045 |
8.291 |
<r2> (average value of r
2) Å
2
| <r2> |
215.300 |
| (<r2>)1/2 |
14.673 |