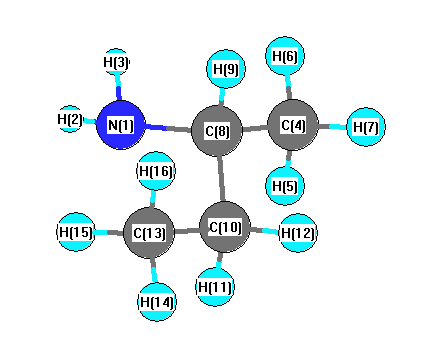
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3813 |
3456 |
1.12 |
|
|
|
| 2 |
A |
3737 |
3387 |
0.10 |
|
|
|
| 3 |
A |
3228 |
2926 |
82.21 |
|
|
|
| 4 |
A |
3219 |
2918 |
49.95 |
|
|
|
| 5 |
A |
3211 |
2911 |
78.84 |
|
|
|
| 6 |
A |
3209 |
2909 |
26.78 |
|
|
|
| 7 |
A |
3184 |
2886 |
8.25 |
|
|
|
| 8 |
A |
3159 |
2864 |
24.45 |
|
|
|
| 9 |
A |
3154 |
2858 |
51.70 |
|
|
|
| 10 |
A |
3151 |
2856 |
26.68 |
|
|
|
| 11 |
A |
3068 |
2781 |
70.14 |
|
|
|
| 12 |
A |
1797 |
1629 |
45.04 |
|
|
|
| 13 |
A |
1626 |
1474 |
6.94 |
|
|
|
| 14 |
A |
1617 |
1465 |
0.10 |
|
|
|
| 15 |
A |
1616 |
1465 |
7.25 |
|
|
|
| 16 |
A |
1610 |
1459 |
3.02 |
|
|
|
| 17 |
A |
1600 |
1451 |
0.21 |
|
|
|
| 18 |
A |
1555 |
1410 |
4.67 |
|
|
|
| 19 |
A |
1536 |
1392 |
8.27 |
|
|
|
| 20 |
A |
1532 |
1388 |
6.63 |
|
|
|
| 21 |
A |
1508 |
1367 |
14.18 |
|
|
|
| 22 |
A |
1455 |
1319 |
3.92 |
|
|
|
| 23 |
A |
1406 |
1274 |
1.70 |
|
|
|
| 24 |
A |
1356 |
1229 |
2.03 |
|
|
|
| 25 |
A |
1287 |
1167 |
2.70 |
|
|
|
| 26 |
A |
1263 |
1145 |
14.63 |
|
|
|
| 27 |
A |
1134 |
1028 |
5.11 |
|
|
|
| 28 |
A |
1093 |
991 |
3.94 |
|
|
|
| 29 |
A |
1091 |
989 |
2.26 |
|
|
|
| 30 |
A |
1059 |
960 |
3.45 |
|
|
|
| 31 |
A |
996 |
903 |
4.13 |
|
|
|
| 32 |
A |
931 |
844 |
116.03 |
|
|
|
| 33 |
A |
865 |
784 |
12.14 |
|
|
|
| 34 |
A |
825 |
748 |
0.61 |
|
|
|
| 35 |
A |
514 |
466 |
7.44 |
|
|
|
| 36 |
A |
484 |
439 |
5.22 |
|
|
|
| 37 |
A |
401 |
364 |
0.78 |
|
|
|
| 38 |
A |
309 |
280 |
30.96 |
|
|
|
| 39 |
A |
278 |
252 |
0.03 |
|
|
|
| 40 |
A |
248 |
225 |
0.63 |
|
|
|
| 41 |
A |
241 |
219 |
17.34 |
|
|
|
| 42 |
A |
117 |
106 |
1.44 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34742.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 31491.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.431 |
|
|
|
| 2 |
H |
0.132 |
|
|
|
| 3 |
H |
0.132 |
|
|
|
| 4 |
C |
-0.204 |
|
|
|
| 5 |
H |
0.062 |
|
|
|
| 6 |
H |
0.045 |
|
|
|
| 7 |
H |
0.047 |
|
|
|
| 8 |
C |
0.224 |
|
|
|
| 9 |
H |
0.021 |
|
|
|
| 10 |
C |
-0.052 |
|
|
|
| 11 |
H |
0.039 |
|
|
|
| 12 |
H |
0.027 |
|
|
|
| 13 |
C |
-0.205 |
|
|
|
| 14 |
H |
0.062 |
|
|
|
| 15 |
H |
0.049 |
|
|
|
| 16 |
H |
0.050 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.028 |
0.095 |
1.209 |
1.213 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.605 |
0.412 |
1.071 |
| y |
0.412 |
-32.736 |
1.948 |
| z |
1.071 |
1.948 |
-35.813 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.670 |
0.412 |
1.071 |
| y |
0.412 |
1.973 |
1.948 |
| z |
1.071 |
1.948 |
-2.642 |
|
| Polar |
|---|
| 3z2-r2 | -5.284 |
|---|
| x2-y2 | -0.869 |
|---|
| xy | 0.412 |
|---|
| xz | 1.071 |
|---|
| yz | 1.948 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.124 |
-0.065 |
-0.005 |
| y |
-0.065 |
8.165 |
-0.003 |
| z |
-0.005 |
-0.003 |
7.371 |
<r2> (average value of r
2) Å
2
| <r2> |
150.578 |
| (<r2>)1/2 |
12.271 |