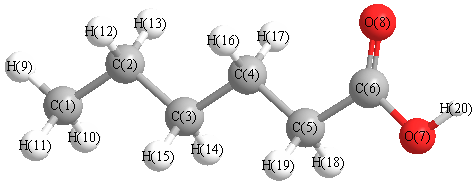
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4112 |
3727 |
121.29 |
|
|
|
| 2 |
A |
3224 |
2922 |
99.83 |
|
|
|
| 3 |
A |
3214 |
2913 |
62.47 |
|
|
|
| 4 |
A |
3210 |
2910 |
59.50 |
|
|
|
| 5 |
A |
3197 |
2898 |
0.02 |
|
|
|
| 6 |
A |
3181 |
2883 |
48.53 |
|
|
|
| 7 |
A |
3175 |
2877 |
1.76 |
|
|
|
| 8 |
A |
3174 |
2877 |
15.96 |
|
|
|
| 9 |
A |
3154 |
2858 |
32.40 |
|
|
|
| 10 |
A |
3150 |
2855 |
4.59 |
|
|
|
| 11 |
A |
3148 |
2853 |
49.42 |
|
|
|
| 12 |
A |
3133 |
2840 |
16.74 |
|
|
|
| 13 |
A |
1991 |
1804 |
383.69 |
|
|
|
| 14 |
A |
1632 |
1479 |
7.75 |
|
|
|
| 15 |
A |
1621 |
1469 |
3.36 |
|
|
|
| 16 |
A |
1615 |
1463 |
6.27 |
|
|
|
| 17 |
A |
1610 |
1459 |
0.40 |
|
|
|
| 18 |
A |
1607 |
1457 |
0.44 |
|
|
|
| 19 |
A |
1585 |
1437 |
17.19 |
|
|
|
| 20 |
A |
1565 |
1419 |
49.86 |
|
|
|
| 21 |
A |
1540 |
1396 |
4.07 |
|
|
|
| 22 |
A |
1533 |
1390 |
15.44 |
|
|
|
| 23 |
A |
1482 |
1343 |
42.66 |
|
|
|
| 24 |
A |
1443 |
1308 |
0.99 |
|
|
|
| 25 |
A |
1440 |
1305 |
0.03 |
|
|
|
| 26 |
A |
1424 |
1291 |
2.72 |
|
|
|
| 27 |
A |
1400 |
1269 |
0.08 |
|
|
|
| 28 |
A |
1375 |
1246 |
15.27 |
|
|
|
| 29 |
A |
1332 |
1208 |
0.29 |
|
|
|
| 30 |
A |
1276 |
1157 |
295.81 |
|
|
|
| 31 |
A |
1240 |
1124 |
2.42 |
|
|
|
| 32 |
A |
1215 |
1102 |
28.33 |
|
|
|
| 33 |
A |
1130 |
1025 |
1.00 |
|
|
|
| 34 |
A |
1123 |
1018 |
7.29 |
|
|
|
| 35 |
A |
1082 |
980 |
4.87 |
|
|
|
| 36 |
A |
1052 |
954 |
1.20 |
|
|
|
| 37 |
A |
981 |
889 |
2.60 |
|
|
|
| 38 |
A |
930 |
843 |
1.42 |
|
|
|
| 39 |
A |
925 |
838 |
7.25 |
|
|
|
| 40 |
A |
825 |
748 |
4.88 |
|
|
|
| 41 |
A |
782 |
709 |
6.86 |
|
|
|
| 42 |
A |
696 |
631 |
39.69 |
|
|
|
| 43 |
A |
690 |
626 |
92.81 |
|
|
|
| 44 |
A |
560 |
507 |
36.58 |
|
|
|
| 45 |
A |
546 |
495 |
20.22 |
|
|
|
| 46 |
A |
433 |
393 |
1.61 |
|
|
|
| 47 |
A |
324 |
294 |
1.49 |
|
|
|
| 48 |
A |
263 |
239 |
0.00 |
|
|
|
| 49 |
A |
241 |
219 |
2.55 |
|
|
|
| 50 |
A |
148 |
134 |
0.04 |
|
|
|
| 51 |
A |
122 |
111 |
0.10 |
|
|
|
| 52 |
A |
105 |
95 |
1.15 |
|
|
|
| 53 |
A |
64 |
58 |
0.16 |
|
|
|
| 54 |
A |
44 |
40 |
0.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41031.1 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 37190.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.189 |
|
|
|
| 2 |
C |
-0.023 |
|
|
|
| 3 |
C |
-0.058 |
|
|
|
| 4 |
C |
-0.026 |
|
|
|
| 5 |
C |
-0.102 |
|
|
|
| 6 |
C |
0.482 |
|
|
|
| 7 |
O |
-0.377 |
|
|
|
| 8 |
O |
-0.450 |
|
|
|
| 9 |
H |
0.062 |
|
|
|
| 10 |
H |
0.047 |
|
|
|
| 11 |
H |
0.047 |
|
|
|
| 12 |
H |
0.031 |
|
|
|
| 13 |
H |
0.031 |
|
|
|
| 14 |
H |
0.031 |
|
|
|
| 15 |
H |
0.031 |
|
|
|
| 16 |
H |
0.045 |
|
|
|
| 17 |
H |
0.045 |
|
|
|
| 18 |
H |
0.069 |
|
|
|
| 19 |
H |
0.069 |
|
|
|
| 20 |
H |
0.234 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.445 |
-1.645 |
0.000 |
1.704 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.078 |
3.688 |
-0.001 |
| y |
3.688 |
-55.953 |
0.001 |
| z |
-0.001 |
0.001 |
-49.008 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.402 |
3.688 |
-0.001 |
| y |
3.688 |
-7.909 |
0.001 |
| z |
-0.001 |
0.001 |
2.507 |
|
| Polar |
|---|
| 3z2-r2 | 5.015 |
|---|
| x2-y2 | 8.874 |
|---|
| xy | 3.688 |
|---|
| xz | -0.001 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.211 |
-0.019 |
0.000 |
| y |
-0.019 |
10.544 |
-0.000 |
| z |
0.000 |
-0.000 |
8.953 |
<r2> (average value of r
2) Å
2
| <r2> |
507.677 |
| (<r2>)1/2 |
22.532 |