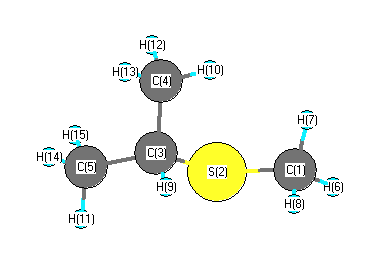
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3263 |
2957 |
16.41 |
|
|
|
| 2 |
A |
3253 |
2949 |
17.91 |
|
|
|
| 3 |
A |
3239 |
2936 |
51.17 |
|
|
|
| 4 |
A |
3235 |
2932 |
26.27 |
|
|
|
| 5 |
A |
3222 |
2920 |
69.10 |
|
|
|
| 6 |
A |
3212 |
2912 |
5.33 |
|
|
|
| 7 |
A |
3178 |
2880 |
32.07 |
|
|
|
| 8 |
A |
3175 |
2878 |
10.04 |
|
|
|
| 9 |
A |
3164 |
2867 |
43.14 |
|
|
|
| 10 |
A |
3156 |
2861 |
26.42 |
|
|
|
| 11 |
A |
1626 |
1473 |
7.24 |
|
|
|
| 12 |
A |
1617 |
1466 |
4.52 |
|
|
|
| 13 |
A |
1608 |
1458 |
7.93 |
|
|
|
| 14 |
A |
1606 |
1456 |
2.19 |
|
|
|
| 15 |
A |
1601 |
1451 |
3.18 |
|
|
|
| 16 |
A |
1588 |
1439 |
8.00 |
|
|
|
| 17 |
A |
1545 |
1400 |
3.15 |
|
|
|
| 18 |
A |
1529 |
1386 |
3.85 |
|
|
|
| 19 |
A |
1483 |
1344 |
3.41 |
|
|
|
| 20 |
A |
1463 |
1326 |
1.32 |
|
|
|
| 21 |
A |
1399 |
1268 |
32.39 |
|
|
|
| 22 |
A |
1283 |
1163 |
9.17 |
|
|
|
| 23 |
A |
1213 |
1100 |
1.05 |
|
|
|
| 24 |
A |
1168 |
1058 |
2.50 |
|
|
|
| 25 |
A |
1058 |
959 |
4.77 |
|
|
|
| 26 |
A |
1049 |
950 |
1.68 |
|
|
|
| 27 |
A |
1035 |
938 |
0.11 |
|
|
|
| 28 |
A |
1008 |
914 |
0.68 |
|
|
|
| 29 |
A |
948 |
859 |
1.21 |
|
|
|
| 30 |
A |
785 |
711 |
1.50 |
|
|
|
| 31 |
A |
691 |
626 |
2.54 |
|
|
|
| 32 |
A |
454 |
411 |
0.89 |
|
|
|
| 33 |
A |
372 |
337 |
0.56 |
|
|
|
| 34 |
A |
355 |
322 |
0.35 |
|
|
|
| 35 |
A |
271 |
246 |
0.08 |
|
|
|
| 36 |
A |
241 |
218 |
0.04 |
|
|
|
| 37 |
A |
220 |
200 |
0.41 |
|
|
|
| 38 |
A |
180 |
163 |
0.56 |
|
|
|
| 39 |
A |
69 |
63 |
2.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30778.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 27898.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.222 |
|
|
|
| 2 |
S |
-0.096 |
|
|
|
| 3 |
C |
0.075 |
|
|
|
| 4 |
C |
-0.194 |
|
|
|
| 5 |
C |
-0.191 |
|
|
|
| 6 |
H |
0.093 |
|
|
|
| 7 |
H |
0.075 |
|
|
|
| 8 |
H |
0.074 |
|
|
|
| 9 |
H |
0.018 |
|
|
|
| 10 |
H |
0.055 |
|
|
|
| 11 |
H |
0.077 |
|
|
|
| 12 |
H |
0.060 |
|
|
|
| 13 |
H |
0.055 |
|
|
|
| 14 |
H |
0.055 |
|
|
|
| 15 |
H |
0.066 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.263 |
1.395 |
1.031 |
1.755 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.620 |
1.411 |
0.504 |
| y |
1.411 |
-42.665 |
-0.045 |
| z |
0.504 |
-0.045 |
-41.770 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.597 |
1.411 |
0.504 |
| y |
1.411 |
-2.969 |
-0.045 |
| z |
0.504 |
-0.045 |
-1.627 |
|
| Polar |
|---|
| 3z2-r2 | -3.255 |
|---|
| x2-y2 | 5.044 |
|---|
| xy | 1.411 |
|---|
| xz | 0.504 |
|---|
| yz | -0.045 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.708 |
0.109 |
-0.005 |
| y |
0.109 |
9.429 |
0.287 |
| z |
-0.005 |
0.287 |
8.274 |
<r2> (average value of r
2) Å
2
| <r2> |
183.024 |
| (<r2>)1/2 |
13.529 |