Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -234.252271 |
| Energy at 298.15K | |
| HF Energy | -234.252271 |
| Nuclear repulsion energy | 252.510115 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3237 |
2934 |
148.86 |
|
|
|
| 2 |
A' |
3221 |
2919 |
60.18 |
|
|
|
| 3 |
A' |
3206 |
2906 |
40.37 |
|
|
|
| 4 |
A' |
3191 |
2893 |
15.28 |
|
|
|
| 5 |
A' |
3178 |
2881 |
61.02 |
|
|
|
| 6 |
A' |
3155 |
2859 |
46.28 |
|
|
|
| 7 |
A' |
1627 |
1475 |
0.48 |
|
|
|
| 8 |
A' |
1622 |
1470 |
7.71 |
|
|
|
| 9 |
A' |
1620 |
1468 |
6.45 |
|
|
|
| 10 |
A' |
1539 |
1395 |
4.30 |
|
|
|
| 11 |
A' |
1485 |
1346 |
2.45 |
|
|
|
| 12 |
A' |
1414 |
1282 |
0.18 |
|
|
|
| 13 |
A' |
1367 |
1239 |
1.10 |
|
|
|
| 14 |
A' |
1291 |
1170 |
0.49 |
|
|
|
| 15 |
A' |
1193 |
1081 |
0.91 |
|
|
|
| 16 |
A' |
1104 |
1001 |
0.68 |
|
|
|
| 17 |
A' |
1051 |
953 |
0.74 |
|
|
|
| 18 |
A' |
1016 |
921 |
0.88 |
|
|
|
| 19 |
A' |
969 |
878 |
0.09 |
|
|
|
| 20 |
A' |
811 |
735 |
0.41 |
|
|
|
| 21 |
A' |
788 |
714 |
0.83 |
|
|
|
| 22 |
A' |
384 |
348 |
0.08 |
|
|
|
| 23 |
A' |
298 |
270 |
0.02 |
|
|
|
| 24 |
A' |
251 |
227 |
0.01 |
|
|
|
| 25 |
A" |
3213 |
2912 |
18.92 |
|
|
|
| 26 |
A" |
3206 |
2906 |
5.00 |
|
|
|
| 27 |
A" |
3201 |
2901 |
67.99 |
|
|
|
| 28 |
A" |
3175 |
2877 |
63.35 |
|
|
|
| 29 |
A" |
3159 |
2863 |
9.09 |
|
|
|
| 30 |
A" |
3149 |
2854 |
15.75 |
|
|
|
| 31 |
A" |
1609 |
1459 |
3.05 |
|
|
|
| 32 |
A" |
1606 |
1456 |
2.15 |
|
|
|
| 33 |
A" |
1598 |
1449 |
2.42 |
|
|
|
| 34 |
A" |
1535 |
1391 |
2.48 |
|
|
|
| 35 |
A" |
1485 |
1346 |
1.99 |
|
|
|
| 36 |
A" |
1422 |
1289 |
0.54 |
|
|
|
| 37 |
A" |
1387 |
1257 |
0.02 |
|
|
|
| 38 |
A" |
1344 |
1218 |
0.25 |
|
|
|
| 39 |
A" |
1283 |
1163 |
0.02 |
|
|
|
| 40 |
A" |
1208 |
1095 |
1.55 |
|
|
|
| 41 |
A" |
1095 |
992 |
0.88 |
|
|
|
| 42 |
A" |
967 |
876 |
0.09 |
|
|
|
| 43 |
A" |
924 |
837 |
0.16 |
|
|
|
| 44 |
A" |
728 |
660 |
0.57 |
|
|
|
| 45 |
A" |
503 |
456 |
0.22 |
|
|
|
| 46 |
A" |
362 |
328 |
0.00 |
|
|
|
| 47 |
A" |
255 |
232 |
0.01 |
|
|
|
| 48 |
A" |
100i |
90i |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38665.0 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 35045.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.505 |
-0.041 |
0.786 |
| C2 |
0.505 |
-0.041 |
-0.786 |
| C3 |
-0.210 |
1.334 |
0.772 |
| C4 |
-0.210 |
1.334 |
-0.772 |
| C5 |
-0.210 |
-1.155 |
1.534 |
| C6 |
-0.210 |
-1.155 |
-1.534 |
| H7 |
1.514 |
0.036 |
1.175 |
| H8 |
1.514 |
0.036 |
-1.175 |
| H9 |
-1.192 |
1.322 |
1.228 |
| H10 |
-1.192 |
1.322 |
-1.228 |
| H11 |
0.357 |
2.142 |
1.215 |
| H12 |
0.357 |
2.142 |
-1.215 |
| H13 |
-1.255 |
-1.224 |
1.261 |
| H14 |
-1.255 |
-1.224 |
-1.261 |
| H15 |
0.240 |
-2.125 |
1.353 |
| H16 |
-0.174 |
-0.978 |
2.604 |
| H17 |
0.240 |
-2.125 |
-1.353 |
| H18 |
-0.174 |
-0.978 |
-2.604 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5713 | 1.5501 | 2.1977 | 1.5203 | 2.6705 | 1.0842 | 2.2063 | 2.2211 | 2.9653 | 2.2298 | 2.9647 | 2.1733 | 2.9471 | 2.1757 | 2.1548 | 2.9977 | 3.5812 |
C2 | 1.5713 | | 2.1977 | 1.5501 | 2.6705 | 1.5203 | 2.2063 | 1.0842 | 2.9653 | 2.2211 | 2.9647 | 2.2298 | 2.9471 | 2.1733 | 2.9977 | 3.5812 | 2.1757 | 2.1548 | C3 | 1.5501 | 2.1977 | | 1.5447 | 2.6024 | 3.3928 | 2.1952 | 2.9066 | 1.0825 | 2.2284 | 1.0819 | 2.2188 | 2.8061 | 3.4306 | 3.5357 | 2.9495 | 4.0844 | 4.0918 | C4 | 2.1977 | 1.5501 | 1.5447 | | 3.3928 | 2.6024 | 2.9066 | 2.1952 | 2.2284 | 1.0825 | 2.2188 | 1.0819 | 3.4306 | 2.8061 | 4.0844 | 4.0918 | 3.5357 | 2.9495 | C5 | 1.5203 | 2.6705 | 2.6024 | 3.3928 | | 3.0677 | 2.1261 | 3.4248 | 2.6813 | 3.8372 | 3.3602 | 4.3294 | 1.0821 | 2.9843 | 1.0844 | 1.0848 | 3.0788 | 4.1413 | C6 | 2.6705 | 1.5203 | 3.3928 | 2.6024 | 3.0677 | | 3.4248 | 2.1261 | 3.8372 | 2.6813 | 4.3294 | 3.3602 | 2.9843 | 1.0821 | 3.0788 | 4.1413 | 1.0844 | 1.0848 | H7 | 1.0842 | 2.2063 | 2.1952 | 2.9066 | 2.1261 | 3.4248 | | 2.3499 | 2.9963 | 3.8405 | 2.4028 | 3.3886 | 3.0437 | 3.8973 | 2.5151 | 2.4327 | 3.5618 | 4.2608 | H8 | 2.2063 | 1.0842 | 2.9066 | 2.1952 | 3.4248 | 2.1261 | 2.3499 | | 3.8405 | 2.9963 | 3.3886 | 2.4028 | 3.8973 | 3.0437 | 3.5618 | 4.2608 | 2.5151 | 2.4327 | H9 | 2.2211 | 2.9653 | 1.0825 | 2.2284 | 2.6813 | 3.8372 | 2.9963 | 3.8405 | | 2.4561 | 1.7530 | 3.0067 | 2.5468 | 3.5608 | 3.7339 | 2.8665 | 4.5376 | 4.5833 | H10 | 2.9653 | 2.2211 | 2.2284 | 1.0825 | 3.8372 | 2.6813 | 3.8405 | 2.9963 | 2.4561 | | 3.0067 | 1.7530 | 3.5608 | 2.5468 | 4.5376 | 4.5833 | 3.7339 | 2.8665 | H11 | 2.2298 | 2.9647 | 1.0819 | 2.2188 | 3.3602 | 4.3294 | 2.4028 | 3.3886 | 1.7530 | 3.0067 | | 2.4294 | 3.7326 | 4.4786 | 4.2704 | 3.4560 | 4.9812 | 4.9593 | H12 | 2.9647 | 2.2298 | 2.2188 | 1.0819 | 4.3294 | 3.3602 | 3.3886 | 2.4028 | 3.0067 | 1.7530 | 2.4294 | | 4.4786 | 3.7326 | 4.9812 | 4.9593 | 4.2704 | 3.4560 | H13 | 2.1733 | 2.9471 | 2.8061 | 3.4306 | 1.0821 | 2.9843 | 3.0437 | 3.8973 | 2.5468 | 3.5608 | 3.7326 | 4.4786 | | 2.5215 | 1.7473 | 1.7416 | 3.1429 | 4.0203 | H14 | 2.9471 | 2.1733 | 3.4306 | 2.8061 | 2.9843 | 1.0821 | 3.8973 | 3.0437 | 3.5608 | 2.5468 | 4.4786 | 3.7326 | 2.5215 | | 3.1429 | 4.0203 | 1.7473 | 1.7416 | H15 | 2.1757 | 2.9977 | 3.5357 | 4.0844 | 1.0844 | 3.0788 | 2.5151 | 3.5618 | 3.7339 | 4.5376 | 4.2704 | 4.9812 | 1.7473 | 3.1429 | | 1.7461 | 2.7065 | 4.1403 | H16 | 2.1548 | 3.5812 | 2.9495 | 4.0918 | 1.0848 | 4.1413 | 2.4327 | 4.2608 | 2.8665 | 4.5833 | 3.4560 | 4.9593 | 1.7416 | 4.0203 | 1.7461 | | 4.1403 | 5.2071 | H17 | 2.9977 | 2.1757 | 4.0844 | 3.5357 | 3.0788 | 1.0844 | 3.5618 | 2.5151 | 4.5376 | 3.7339 | 4.9812 | 4.2704 | 3.1429 | 1.7473 | 2.7065 | 4.1403 | | 1.7461 | H18 | 3.5812 | 2.1548 | 4.0918 | 2.9495 | 4.1413 | 1.0848 | 4.2608 | 2.4327 | 4.5833 | 2.8665 | 4.9593 | 3.4560 | 4.0203 | 1.7416 | 4.1403 | 5.2071 | 1.7461 | |
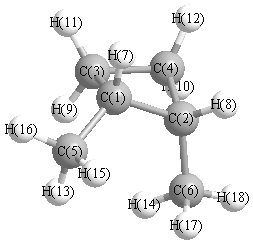 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
89.508 |
|
C1 |
C2 |
C6 |
119.482 |
| C1 |
C2 |
H8 |
111.044 |
|
C1 |
C3 |
C4 |
90.492 |
| C1 |
C3 |
H9 |
113.883 |
|
C1 |
C3 |
H11 |
114.649 |
| C1 |
C5 |
H13 |
112.161 |
|
C1 |
C5 |
H15 |
112.210 |
| C1 |
C5 |
H16 |
110.495 |
|
C2 |
C1 |
C3 |
89.508 |
| C2 |
C1 |
C5 |
119.482 |
|
C2 |
C1 |
H7 |
111.044 |
| C2 |
C4 |
C3 |
90.492 |
|
C2 |
C4 |
H10 |
113.883 |
| C2 |
C4 |
H12 |
114.649 |
|
C2 |
C6 |
H14 |
112.161 |
| C2 |
C6 |
H17 |
112.210 |
|
C2 |
C6 |
H18 |
110.495 |
| C3 |
C1 |
C5 |
115.898 |
|
C3 |
C1 |
H7 |
111.659 |
| C3 |
C4 |
H10 |
114.897 |
|
C3 |
C4 |
H12 |
114.136 |
| C4 |
C2 |
C6 |
115.898 |
|
C4 |
C2 |
H8 |
111.659 |
| C4 |
C3 |
H9 |
114.897 |
|
C4 |
C3 |
H11 |
114.136 |
| C5 |
C1 |
H7 |
108.272 |
|
C6 |
C2 |
H8 |
108.272 |
| H9 |
C3 |
H11 |
108.177 |
|
H10 |
C4 |
H12 |
108.177 |
| H13 |
C5 |
H15 |
107.513 |
|
H13 |
C5 |
H16 |
106.973 |
| H14 |
C6 |
H17 |
107.513 |
|
H14 |
C6 |
H18 |
106.973 |
| H15 |
C5 |
H16 |
107.212 |
|
H17 |
C6 |
H18 |
107.212 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.082 |
|
|
|
| 2 |
C |
0.082 |
|
|
|
| 3 |
C |
-0.159 |
|
|
|
| 4 |
C |
-0.159 |
|
|
|
| 5 |
C |
-0.239 |
|
|
|
| 6 |
C |
-0.239 |
|
|
|
| 7 |
H |
0.041 |
|
|
|
| 8 |
H |
0.041 |
|
|
|
| 9 |
H |
0.053 |
|
|
|
| 10 |
H |
0.053 |
|
|
|
| 11 |
H |
0.055 |
|
|
|
| 12 |
H |
0.055 |
|
|
|
| 13 |
H |
0.045 |
|
|
|
| 14 |
H |
0.045 |
|
|
|
| 15 |
H |
0.058 |
|
|
|
| 16 |
H |
0.065 |
|
|
|
| 17 |
H |
0.058 |
|
|
|
| 18 |
H |
0.065 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.170 |
-0.015 |
0.000 |
0.171 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.680 |
0.169 |
0.000 |
| y |
0.169 |
-41.064 |
0.000 |
| z |
0.000 |
0.000 |
-40.859 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.282 |
0.169 |
0.000 |
| y |
0.169 |
-0.795 |
0.000 |
| z |
0.000 |
0.000 |
-0.487 |
|
| Polar |
|---|
| 3z2-r2 | -0.975 |
|---|
| x2-y2 | 1.385 |
|---|
| xy | 0.169 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.820 |
-0.080 |
0.000 |
| y |
-0.080 |
10.238 |
0.000 |
| z |
0.000 |
0.000 |
10.383 |
<r2> (average value of r
2) Å
2
| <r2> |
175.590 |
| (<r2>)1/2 |
13.251 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -234.254223 |
| Energy at 298.15K | -234.268080 |
| HF Energy | -234.254223 |
| Nuclear repulsion energy | 253.555593 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3234 |
2931 |
137.09 |
|
|
|
| 2 |
A |
3219 |
2918 |
26.94 |
|
|
|
| 3 |
A |
3213 |
2913 |
83.23 |
|
|
|
| 4 |
A |
3210 |
2910 |
7.85 |
|
|
|
| 5 |
A |
3207 |
2907 |
57.23 |
|
|
|
| 6 |
A |
3199 |
2900 |
45.25 |
|
|
|
| 7 |
A |
3189 |
2891 |
19.63 |
|
|
|
| 8 |
A |
3176 |
2879 |
65.79 |
|
|
|
| 9 |
A |
3172 |
2875 |
41.94 |
|
|
|
| 10 |
A |
3157 |
2861 |
15.07 |
|
|
|
| 11 |
A |
3153 |
2858 |
43.98 |
|
|
|
| 12 |
A |
3149 |
2854 |
22.31 |
|
|
|
| 13 |
A |
1629 |
1477 |
1.60 |
|
|
|
| 14 |
A |
1622 |
1470 |
1.40 |
|
|
|
| 15 |
A |
1620 |
1468 |
9.10 |
|
|
|
| 16 |
A |
1610 |
1459 |
4.68 |
|
|
|
| 17 |
A |
1608 |
1457 |
3.66 |
|
|
|
| 18 |
A |
1603 |
1453 |
1.60 |
|
|
|
| 19 |
A |
1539 |
1395 |
4.08 |
|
|
|
| 20 |
A |
1538 |
1394 |
4.21 |
|
|
|
| 21 |
A |
1497 |
1357 |
2.19 |
|
|
|
| 22 |
A |
1487 |
1348 |
2.75 |
|
|
|
| 23 |
A |
1420 |
1287 |
0.14 |
|
|
|
| 24 |
A |
1407 |
1275 |
0.20 |
|
|
|
| 25 |
A |
1383 |
1253 |
0.07 |
|
|
|
| 26 |
A |
1371 |
1243 |
0.71 |
|
|
|
| 27 |
A |
1325 |
1201 |
0.58 |
|
|
|
| 28 |
A |
1288 |
1167 |
0.06 |
|
|
|
| 29 |
A |
1279 |
1159 |
0.84 |
|
|
|
| 30 |
A |
1206 |
1093 |
1.21 |
|
|
|
| 31 |
A |
1195 |
1083 |
1.34 |
|
|
|
| 32 |
A |
1118 |
1013 |
0.96 |
|
|
|
| 33 |
A |
1085 |
983 |
0.55 |
|
|
|
| 34 |
A |
1039 |
942 |
1.00 |
|
|
|
| 35 |
A |
1015 |
920 |
0.70 |
|
|
|
| 36 |
A |
971 |
880 |
0.21 |
|
|
|
| 37 |
A |
960 |
870 |
0.25 |
|
|
|
| 38 |
A |
948 |
860 |
0.27 |
|
|
|
| 39 |
A |
834 |
756 |
0.55 |
|
|
|
| 40 |
A |
802 |
727 |
0.31 |
|
|
|
| 41 |
A |
686 |
622 |
1.79 |
|
|
|
| 42 |
A |
522 |
473 |
0.22 |
|
|
|
| 43 |
A |
379 |
344 |
0.09 |
|
|
|
| 44 |
A |
364 |
330 |
0.04 |
|
|
|
| 45 |
A |
286 |
259 |
0.01 |
|
|
|
| 46 |
A |
261 |
237 |
0.02 |
|
|
|
| 47 |
A |
239 |
217 |
0.02 |
|
|
|
| 48 |
A |
140 |
127 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38776.6 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 35147.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.737 |
0.241 |
0.527 |
| C2 |
-0.777 |
-0.162 |
0.475 |
| C3 |
1.102 |
-1.127 |
-0.102 |
| C4 |
-0.396 |
-1.433 |
-0.321 |
| C5 |
1.209 |
1.454 |
-0.260 |
| C6 |
-1.806 |
0.765 |
-0.151 |
| H7 |
1.091 |
0.341 |
1.547 |
| H8 |
-1.124 |
-0.422 |
1.469 |
| H9 |
1.700 |
-1.068 |
-1.003 |
| H10 |
-0.701 |
-1.395 |
-1.360 |
| H11 |
1.595 |
-1.811 |
0.576 |
| H12 |
-0.747 |
-2.370 |
0.091 |
| H13 |
0.940 |
1.392 |
-1.307 |
| H14 |
-1.548 |
1.041 |
-1.164 |
| H15 |
0.799 |
2.377 |
0.131 |
| H16 |
2.290 |
1.534 |
-0.217 |
| H17 |
-1.927 |
1.680 |
0.418 |
| H18 |
-2.777 |
0.282 |
-0.191 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5676 | 1.5492 | 2.1922 | 1.5210 | 2.6842 | 1.0839 | 2.1884 | 2.2314 | 2.8819 | 2.2242 | 3.0342 | 2.1755 | 2.9535 | 2.1738 | 2.1531 | 3.0299 | 3.5871 |
C2 | 1.5676 | | 2.1898 | 1.5476 | 2.6638 | 1.5208 | 2.2117 | 1.0840 | 3.0234 | 2.2123 | 2.8903 | 2.2406 | 2.9230 | 2.1753 | 3.0085 | 3.5722 | 2.1728 | 2.1546 | C3 | 1.5492 | 2.1898 | | 1.5447 | 2.5879 | 3.4704 | 2.2075 | 2.8139 | 1.0830 | 2.2149 | 1.0815 | 2.2361 | 2.7979 | 3.5852 | 3.5253 | 2.9164 | 4.1625 | 4.1282 | C4 | 2.1922 | 1.5476 | 1.5447 | | 3.3035 | 2.6175 | 2.9748 | 2.1810 | 2.2343 | 1.0830 | 2.2163 | 1.0817 | 3.2773 | 2.8566 | 4.0189 | 4.0034 | 3.5474 | 2.9374 | C5 | 1.5210 | 2.6638 | 2.5879 | 3.3035 | | 3.0948 | 2.1251 | 3.4568 | 2.6738 | 3.6019 | 3.3919 | 4.3088 | 1.0829 | 2.9303 | 1.0837 | 1.0847 | 3.2160 | 4.1550 | C6 | 2.6842 | 1.5208 | 3.4704 | 2.6175 | 3.0948 | | 3.3852 | 2.1217 | 4.0474 | 2.7113 | 4.3284 | 3.3177 | 3.0452 | 1.0816 | 3.0766 | 4.1681 | 1.0845 | 1.0849 | H7 | 1.0839 | 2.2117 | 2.2075 | 2.9748 | 2.1251 | 3.3852 | | 2.3440 | 2.9754 | 3.8308 | 2.4138 | 3.5842 | 3.0452 | 3.8477 | 2.4970 | 2.4429 | 3.4894 | 4.2413 | H8 | 2.1884 | 1.0840 | 2.8139 | 2.1810 | 3.4568 | 2.1217 | 2.3440 | | 3.8079 | 3.0213 | 3.1806 | 2.4151 | 3.9064 | 3.0426 | 3.6501 | 4.2800 | 2.4841 | 2.4470 | H9 | 2.2314 | 3.0234 | 1.0830 | 2.2343 | 2.6738 | 4.0474 | 2.9754 | 3.8079 | | 2.4498 | 1.7479 | 2.9801 | 2.5928 | 3.8761 | 3.7370 | 2.7808 | 4.7671 | 4.7462 | H10 | 2.8819 | 2.2123 | 2.2149 | 1.0830 | 3.6019 | 2.7113 | 3.8308 | 3.0213 | 2.4498 | | 3.0318 | 1.7480 | 3.2353 | 2.5869 | 4.3250 | 4.3396 | 3.7582 | 2.9136 | H11 | 2.2242 | 2.8903 | 1.0815 | 2.2163 | 3.3919 | 4.3284 | 2.4138 | 3.1806 | 1.7479 | 3.0318 | | 2.4563 | 3.7732 | 4.5870 | 4.2863 | 3.5069 | 4.9615 | 4.9075 | H12 | 3.0342 | 2.2406 | 2.2361 | 1.0817 | 4.3088 | 3.3177 | 3.5842 | 2.4151 | 2.9801 | 1.7480 | 2.4563 | | 4.3536 | 3.7215 | 4.9924 | 4.9552 | 4.2309 | 3.3513 | H13 | 2.1755 | 2.9230 | 2.7979 | 3.2773 | 1.0829 | 3.0452 | 3.0452 | 3.9064 | 2.5928 | 3.2353 | 3.7732 | 4.3536 | | 2.5166 | 1.7491 | 1.7410 | 3.3583 | 4.0366 | H14 | 2.9535 | 2.1753 | 3.5852 | 2.8566 | 2.9303 | 1.0816 | 3.8477 | 3.0426 | 3.8761 | 2.5869 | 4.5870 | 3.7215 | 2.5166 | | 2.9951 | 3.9834 | 1.7483 | 1.7418 | H15 | 2.1738 | 3.0085 | 3.5253 | 4.0189 | 1.0837 | 3.0766 | 2.4970 | 3.6501 | 3.7370 | 4.3250 | 4.2863 | 4.9924 | 1.7491 | 2.9951 | | 1.7477 | 2.8277 | 4.1569 | H16 | 2.1531 | 3.5722 | 2.9164 | 4.0034 | 1.0847 | 4.1681 | 2.4429 | 4.2800 | 2.7808 | 4.3396 | 3.5069 | 4.9552 | 1.7410 | 3.9834 | 1.7477 | | 4.2663 | 5.2190 | H17 | 3.0299 | 2.1728 | 4.1625 | 3.5474 | 3.2160 | 1.0845 | 3.4894 | 2.4841 | 4.7671 | 3.7582 | 4.9615 | 4.2309 | 3.3583 | 1.7483 | 2.8277 | 4.2663 | | 1.7464 | H18 | 3.5871 | 2.1546 | 4.1282 | 2.9374 | 4.1550 | 1.0849 | 4.2413 | 2.4470 | 4.7462 | 2.9136 | 4.9075 | 3.3513 | 4.0366 | 1.7418 | 4.1569 | 5.2190 | 1.7464 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
89.447 |
|
C1 |
C2 |
C6 |
120.701 |
| C1 |
C2 |
H8 |
109.890 |
|
C1 |
C3 |
C4 |
90.234 |
| C1 |
C3 |
H9 |
114.784 |
|
C1 |
C3 |
H11 |
114.275 |
| C1 |
C5 |
H13 |
112.238 |
|
C1 |
C5 |
H15 |
112.049 |
| C1 |
C5 |
H16 |
110.320 |
|
C2 |
C1 |
C3 |
89.264 |
| C2 |
C1 |
C5 |
119.175 |
|
C2 |
C1 |
H7 |
111.745 |
| C2 |
C4 |
C3 |
90.167 |
|
C2 |
C4 |
H10 |
113.311 |
| C2 |
C4 |
H12 |
115.766 |
|
C2 |
C6 |
H14 |
112.318 |
| C2 |
C6 |
H17 |
111.931 |
|
C2 |
C6 |
H18 |
110.444 |
| C3 |
C1 |
C5 |
114.895 |
|
C3 |
C1 |
H7 |
112.741 |
| C3 |
C4 |
H10 |
113.738 |
|
C3 |
C4 |
H12 |
115.603 |
| C4 |
C2 |
C6 |
117.089 |
|
C4 |
C2 |
H8 |
110.717 |
| C4 |
C3 |
H9 |
115.366 |
|
C4 |
C3 |
H11 |
113.947 |
| C5 |
C1 |
H7 |
108.157 |
|
C6 |
C2 |
H8 |
107.909 |
| H9 |
C3 |
H11 |
107.717 |
|
H10 |
C4 |
H12 |
107.711 |
| H13 |
C5 |
H15 |
107.672 |
|
H13 |
C5 |
H16 |
106.880 |
| H14 |
C6 |
H17 |
107.637 |
|
H14 |
C6 |
H18 |
107.020 |
| H15 |
C5 |
H16 |
107.420 |
|
H17 |
C6 |
H18 |
107.225 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.093 |
|
|
|
| 2 |
C |
0.064 |
|
|
|
| 3 |
C |
-0.117 |
|
|
|
| 4 |
C |
-0.145 |
|
|
|
| 5 |
C |
-0.254 |
|
|
|
| 6 |
C |
-0.225 |
|
|
|
| 7 |
H |
0.028 |
|
|
|
| 8 |
H |
0.038 |
|
|
|
| 9 |
H |
0.042 |
|
|
|
| 10 |
H |
0.040 |
|
|
|
| 11 |
H |
0.050 |
|
|
|
| 12 |
H |
0.044 |
|
|
|
| 13 |
H |
0.050 |
|
|
|
| 14 |
H |
0.044 |
|
|
|
| 15 |
H |
0.059 |
|
|
|
| 16 |
H |
0.065 |
|
|
|
| 17 |
H |
0.059 |
|
|
|
| 18 |
H |
0.064 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.024 |
0.006 |
0.149 |
0.151 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.008 |
0.252 |
-0.125 |
| y |
0.252 |
-40.749 |
-0.245 |
| z |
-0.125 |
-0.245 |
-39.858 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.705 |
0.252 |
-0.125 |
| y |
0.252 |
-0.316 |
-0.245 |
| z |
-0.125 |
-0.245 |
1.021 |
|
| Polar |
|---|
| 3z2-r2 | 2.042 |
|---|
| x2-y2 | -0.259 |
|---|
| xy | 0.252 |
|---|
| xz | -0.125 |
|---|
| yz | -0.245 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.456 |
-0.015 |
0.005 |
| y |
-0.015 |
10.153 |
0.083 |
| z |
0.005 |
0.083 |
8.925 |
<r2> (average value of r
2) Å
2
| <r2> |
173.777 |
| (<r2>)1/2 |
13.182 |
