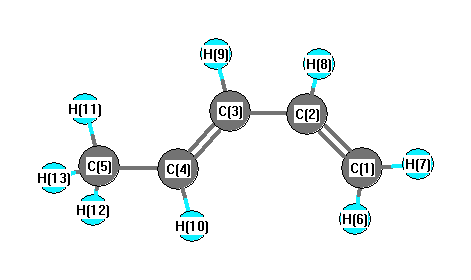
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3367 |
3052 |
17.08 |
|
|
|
| 2 |
A |
3292 |
2984 |
6.70 |
|
|
|
| 3 |
A |
3290 |
2982 |
31.80 |
|
|
|
| 4 |
A |
3280 |
2973 |
19.65 |
|
|
|
| 5 |
A |
3269 |
2963 |
13.17 |
|
|
|
| 6 |
A |
3230 |
2928 |
22.04 |
|
|
|
| 7 |
A |
3195 |
2896 |
31.22 |
|
|
|
| 8 |
A |
3153 |
2858 |
43.34 |
|
|
|
| 9 |
A |
1862 |
1688 |
8.35 |
|
|
|
| 10 |
A |
1827 |
1656 |
13.55 |
|
|
|
| 11 |
A |
1608 |
1457 |
11.05 |
|
|
|
| 12 |
A |
1597 |
1447 |
6.41 |
|
|
|
| 13 |
A |
1569 |
1422 |
4.94 |
|
|
|
| 14 |
A |
1538 |
1394 |
2.28 |
|
|
|
| 15 |
A |
1452 |
1316 |
0.40 |
|
|
|
| 16 |
A |
1436 |
1302 |
1.18 |
|
|
|
| 17 |
A |
1419 |
1286 |
1.76 |
|
|
|
| 18 |
A |
1218 |
1104 |
1.43 |
|
|
|
| 19 |
A |
1176 |
1066 |
4.78 |
|
|
|
| 20 |
A |
1162 |
1053 |
0.07 |
|
|
|
| 21 |
A |
1130 |
1024 |
19.17 |
|
|
|
| 22 |
A |
1097 |
994 |
42.46 |
|
|
|
| 23 |
A |
1069 |
969 |
37.80 |
|
|
|
| 24 |
A |
1055 |
956 |
13.58 |
|
|
|
| 25 |
A |
966 |
876 |
5.28 |
|
|
|
| 26 |
A |
915 |
829 |
0.47 |
|
|
|
| 27 |
A |
720 |
653 |
9.14 |
|
|
|
| 28 |
A |
609 |
552 |
5.22 |
|
|
|
| 29 |
A |
416 |
377 |
2.10 |
|
|
|
| 30 |
A |
268 |
243 |
2.68 |
|
|
|
| 31 |
A |
228 |
207 |
0.93 |
|
|
|
| 32 |
A |
194 |
176 |
1.34 |
|
|
|
| 33 |
A |
145 |
132 |
0.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26374.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 23906.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.159 |
|
|
|
| 2 |
C |
-0.077 |
|
|
|
| 3 |
C |
-0.065 |
|
|
|
| 4 |
C |
-0.061 |
|
|
|
| 5 |
C |
-0.128 |
|
|
|
| 6 |
H |
0.062 |
|
|
|
| 7 |
H |
0.070 |
|
|
|
| 8 |
H |
0.067 |
|
|
|
| 9 |
H |
0.051 |
|
|
|
| 10 |
H |
0.060 |
|
|
|
| 11 |
H |
0.058 |
|
|
|
| 12 |
H |
0.060 |
|
|
|
| 13 |
H |
0.062 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.482 |
-0.036 |
0.042 |
0.485 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.324 |
0.350 |
0.583 |
| y |
0.350 |
-30.085 |
-0.037 |
| z |
0.583 |
-0.037 |
-34.706 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.071 |
0.350 |
0.583 |
| y |
0.350 |
2.430 |
-0.037 |
| z |
0.583 |
-0.037 |
-4.502 |
|
| Polar |
|---|
| 3z2-r2 | -9.003 |
|---|
| x2-y2 | -0.239 |
|---|
| xy | 0.350 |
|---|
| xz | 0.583 |
|---|
| yz | -0.037 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.120 |
-0.077 |
-0.334 |
| y |
-0.077 |
8.658 |
-0.007 |
| z |
-0.334 |
-0.007 |
6.289 |
<r2> (average value of r
2) Å
2
| <r2> |
159.688 |
| (<r2>)1/2 |
12.637 |