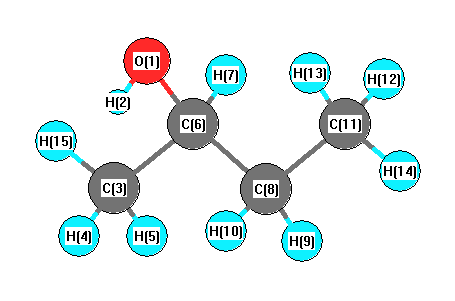
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4155 |
3766 |
37.00 |
|
|
|
| 2 |
A |
3239 |
2936 |
58.56 |
|
|
|
| 3 |
A |
3234 |
2931 |
31.63 |
|
|
|
| 4 |
A |
3216 |
2915 |
118.27 |
|
|
|
| 5 |
A |
3210 |
2910 |
20.65 |
|
|
|
| 6 |
A |
3184 |
2886 |
21.05 |
|
|
|
| 7 |
A |
3172 |
2875 |
18.67 |
|
|
|
| 8 |
A |
3159 |
2864 |
32.03 |
|
|
|
| 9 |
A |
3150 |
2855 |
33.74 |
|
|
|
| 10 |
A |
3130 |
2837 |
22.56 |
|
|
|
| 11 |
A |
1624 |
1472 |
6.32 |
|
|
|
| 12 |
A |
1614 |
1463 |
9.09 |
|
|
|
| 13 |
A |
1612 |
1461 |
2.34 |
|
|
|
| 14 |
A |
1610 |
1460 |
2.58 |
|
|
|
| 15 |
A |
1600 |
1450 |
0.23 |
|
|
|
| 16 |
A |
1549 |
1404 |
35.49 |
|
|
|
| 17 |
A |
1537 |
1393 |
11.77 |
|
|
|
| 18 |
A |
1533 |
1389 |
13.90 |
|
|
|
| 19 |
A |
1520 |
1378 |
4.09 |
|
|
|
| 20 |
A |
1446 |
1310 |
2.34 |
|
|
|
| 21 |
A |
1437 |
1302 |
6.81 |
|
|
|
| 22 |
A |
1361 |
1234 |
8.44 |
|
|
|
| 23 |
A |
1285 |
1165 |
27.87 |
|
|
|
| 24 |
A |
1225 |
1111 |
32.69 |
|
|
|
| 25 |
A |
1186 |
1075 |
75.27 |
|
|
|
| 26 |
A |
1115 |
1010 |
7.95 |
|
|
|
| 27 |
A |
1076 |
975 |
7.37 |
|
|
|
| 28 |
A |
1058 |
959 |
8.91 |
|
|
|
| 29 |
A |
996 |
903 |
10.89 |
|
|
|
| 30 |
A |
875 |
794 |
5.61 |
|
|
|
| 31 |
A |
830 |
753 |
0.91 |
|
|
|
| 32 |
A |
531 |
481 |
10.06 |
|
|
|
| 33 |
A |
496 |
450 |
7.84 |
|
|
|
| 34 |
A |
403 |
365 |
6.63 |
|
|
|
| 35 |
A |
288 |
261 |
122.68 |
|
|
|
| 36 |
A |
273 |
248 |
0.73 |
|
|
|
| 37 |
A |
252 |
228 |
4.93 |
|
|
|
| 38 |
A |
236 |
214 |
0.07 |
|
|
|
| 39 |
A |
119 |
108 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31767.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 28794.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.443 |
|
|
|
| 2 |
H |
0.192 |
|
|
|
| 3 |
C |
-0.209 |
|
|
|
| 4 |
H |
0.039 |
|
|
|
| 5 |
H |
0.047 |
|
|
|
| 6 |
C |
0.316 |
|
|
|
| 7 |
H |
0.015 |
|
|
|
| 8 |
C |
-0.071 |
|
|
|
| 9 |
H |
0.029 |
|
|
|
| 10 |
H |
0.018 |
|
|
|
| 11 |
C |
-0.178 |
|
|
|
| 12 |
H |
0.044 |
|
|
|
| 13 |
H |
0.066 |
|
|
|
| 14 |
H |
0.058 |
|
|
|
| 15 |
H |
0.078 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.167 |
-1.340 |
0.952 |
1.652 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.877 |
0.218 |
-0.740 |
| y |
0.218 |
-35.969 |
2.466 |
| z |
-0.740 |
2.466 |
-31.002 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.392 |
0.218 |
-0.740 |
| y |
0.218 |
-3.529 |
2.466 |
| z |
-0.740 |
2.466 |
3.921 |
|
| Polar |
|---|
| 3z2-r2 | 7.842 |
|---|
| x2-y2 | 2.092 |
|---|
| xy | 0.218 |
|---|
| xz | -0.740 |
|---|
| yz | 2.466 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.371 |
0.128 |
-0.068 |
| y |
0.128 |
7.303 |
0.061 |
| z |
-0.068 |
0.061 |
7.119 |
<r2> (average value of r
2) Å
2
| <r2> |
145.800 |
| (<r2>)1/2 |
12.075 |