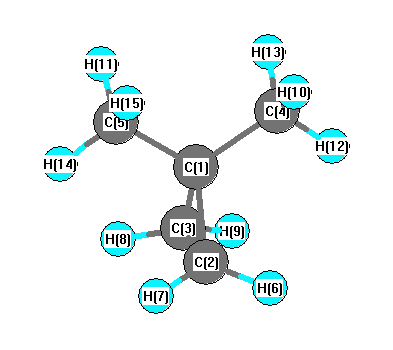
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3265 |
2959 |
15.46 |
|
|
|
| 2 |
A1 |
3218 |
2917 |
50.85 |
|
|
|
| 3 |
A1 |
3152 |
2857 |
53.89 |
|
|
|
| 4 |
A1 |
1648 |
1494 |
1.09 |
|
|
|
| 5 |
A1 |
1623 |
1471 |
5.48 |
|
|
|
| 6 |
A1 |
1549 |
1404 |
0.00 |
|
|
|
| 7 |
A1 |
1458 |
1321 |
1.70 |
|
|
|
| 8 |
A1 |
1171 |
1062 |
1.18 |
|
|
|
| 9 |
A1 |
1150 |
1042 |
3.89 |
|
|
|
| 10 |
A1 |
1009 |
915 |
15.55 |
|
|
|
| 11 |
A1 |
726 |
658 |
2.46 |
|
|
|
| 12 |
A1 |
376 |
340 |
0.09 |
|
|
|
| 13 |
A2 |
3329 |
3017 |
0.00 |
|
|
|
| 14 |
A2 |
3194 |
2895 |
0.00 |
|
|
|
| 15 |
A2 |
1599 |
1450 |
0.00 |
|
|
|
| 16 |
A2 |
1286 |
1166 |
0.00 |
|
|
|
| 17 |
A2 |
1155 |
1047 |
0.00 |
|
|
|
| 18 |
A2 |
975 |
884 |
0.00 |
|
|
|
| 19 |
A2 |
356 |
323 |
0.00 |
|
|
|
| 20 |
A2 |
232 |
211 |
0.00 |
|
|
|
| 21 |
B1 |
3256 |
2952 |
34.66 |
|
|
|
| 22 |
B1 |
3200 |
2900 |
90.73 |
|
|
|
| 23 |
B1 |
1618 |
1467 |
5.09 |
|
|
|
| 24 |
B1 |
1592 |
1443 |
4.21 |
|
|
|
| 25 |
B1 |
1260 |
1142 |
4.80 |
|
|
|
| 26 |
B1 |
1196 |
1084 |
0.12 |
|
|
|
| 27 |
B1 |
919 |
833 |
11.48 |
|
|
|
| 28 |
B1 |
427 |
387 |
0.91 |
|
|
|
| 29 |
B1 |
265 |
240 |
0.00 |
|
|
|
| 30 |
B2 |
3345 |
3032 |
39.49 |
|
|
|
| 31 |
B2 |
3215 |
2914 |
28.47 |
|
|
|
| 32 |
B2 |
3145 |
2851 |
37.15 |
|
|
|
| 33 |
B2 |
1610 |
1460 |
3.84 |
|
|
|
| 34 |
B2 |
1539 |
1395 |
5.70 |
|
|
|
| 35 |
B2 |
1421 |
1288 |
1.01 |
|
|
|
| 36 |
B2 |
1052 |
953 |
0.37 |
|
|
|
| 37 |
B2 |
1014 |
919 |
0.18 |
|
|
|
| 38 |
B2 |
847 |
767 |
0.02 |
|
|
|
| 39 |
B2 |
386 |
350 |
0.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31888.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 28903.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.182 |
|
|
|
| 2 |
C |
-0.153 |
|
|
|
| 3 |
C |
-0.153 |
|
|
|
| 4 |
C |
-0.184 |
|
|
|
| 5 |
C |
-0.184 |
|
|
|
| 6 |
H |
0.041 |
|
|
|
| 7 |
H |
0.041 |
|
|
|
| 8 |
H |
0.041 |
|
|
|
| 9 |
H |
0.041 |
|
|
|
| 10 |
H |
0.056 |
|
|
|
| 11 |
H |
0.056 |
|
|
|
| 12 |
H |
0.056 |
|
|
|
| 13 |
H |
0.056 |
|
|
|
| 14 |
H |
0.051 |
|
|
|
| 15 |
H |
0.051 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.138 |
0.138 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.800 |
0.000 |
0.000 |
| y |
0.000 |
-32.716 |
0.000 |
| z |
0.000 |
0.000 |
-34.517 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.183 |
0.000 |
0.000 |
| y |
0.000 |
1.443 |
0.000 |
| z |
0.000 |
0.000 |
-1.259 |
|
| Polar |
|---|
| 3z2-r2 | -2.519 |
|---|
| x2-y2 | -1.084 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.962 |
0.000 |
0.000 |
| y |
0.000 |
8.175 |
0.000 |
| z |
0.000 |
0.000 |
8.783 |
<r2> (average value of r
2) Å
2
| <r2> |
122.026 |
| (<r2>)1/2 |
11.047 |