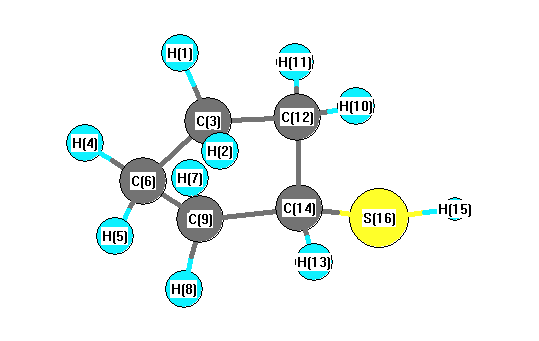
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3240 |
2936 |
109.22 |
|
|
|
| 2 |
A |
3224 |
2922 |
31.17 |
|
|
|
| 3 |
A |
3219 |
2918 |
18.29 |
|
|
|
| 4 |
A |
3204 |
2904 |
41.88 |
|
|
|
| 5 |
A |
3194 |
2895 |
26.28 |
|
|
|
| 6 |
A |
3187 |
2889 |
48.21 |
|
|
|
| 7 |
A |
3175 |
2878 |
19.68 |
|
|
|
| 8 |
A |
3171 |
2874 |
38.63 |
|
|
|
| 9 |
A |
3170 |
2873 |
1.26 |
|
|
|
| 10 |
A |
2850 |
2583 |
5.99 |
|
|
|
| 11 |
A |
1646 |
1492 |
0.69 |
|
|
|
| 12 |
A |
1621 |
1470 |
3.56 |
|
|
|
| 13 |
A |
1620 |
1468 |
3.08 |
|
|
|
| 14 |
A |
1608 |
1458 |
0.26 |
|
|
|
| 15 |
A |
1489 |
1350 |
5.19 |
|
|
|
| 16 |
A |
1471 |
1333 |
0.49 |
|
|
|
| 17 |
A |
1460 |
1323 |
0.52 |
|
|
|
| 18 |
A |
1443 |
1308 |
3.33 |
|
|
|
| 19 |
A |
1415 |
1282 |
0.76 |
|
|
|
| 20 |
A |
1392 |
1262 |
17.01 |
|
|
|
| 21 |
A |
1374 |
1245 |
1.59 |
|
|
|
| 22 |
A |
1339 |
1214 |
3.92 |
|
|
|
| 23 |
A |
1310 |
1187 |
0.52 |
|
|
|
| 24 |
A |
1272 |
1153 |
4.83 |
|
|
|
| 25 |
A |
1199 |
1086 |
1.00 |
|
|
|
| 26 |
A |
1122 |
1017 |
1.31 |
|
|
|
| 27 |
A |
1070 |
969 |
0.11 |
|
|
|
| 28 |
A |
1049 |
951 |
0.40 |
|
|
|
| 29 |
A |
1019 |
924 |
1.40 |
|
|
|
| 30 |
A |
995 |
902 |
1.49 |
|
|
|
| 31 |
A |
952 |
862 |
6.07 |
|
|
|
| 32 |
A |
943 |
854 |
0.64 |
|
|
|
| 33 |
A |
888 |
804 |
2.03 |
|
|
|
| 34 |
A |
865 |
784 |
3.33 |
|
|
|
| 35 |
A |
802 |
726 |
1.76 |
|
|
|
| 36 |
A |
668 |
605 |
0.25 |
|
|
|
| 37 |
A |
493 |
447 |
0.09 |
|
|
|
| 38 |
A |
393 |
356 |
1.36 |
|
|
|
| 39 |
A |
268 |
243 |
2.51 |
|
|
|
| 40 |
A |
219 |
198 |
18.04 |
|
|
|
| 41 |
A |
177 |
160 |
0.51 |
|
|
|
| 42 |
A |
40 |
36 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32624.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 29571.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.057 |
|
|
|
| 2 |
H |
0.049 |
|
|
|
| 3 |
C |
-0.105 |
|
|
|
| 4 |
H |
0.056 |
|
|
|
| 5 |
H |
0.054 |
|
|
|
| 6 |
C |
-0.104 |
|
|
|
| 7 |
H |
0.061 |
|
|
|
| 8 |
H |
0.070 |
|
|
|
| 9 |
C |
-0.137 |
|
|
|
| 10 |
H |
0.049 |
|
|
|
| 11 |
H |
0.048 |
|
|
|
| 12 |
C |
-0.106 |
|
|
|
| 13 |
H |
0.045 |
|
|
|
| 14 |
C |
0.068 |
|
|
|
| 15 |
H |
0.055 |
|
|
|
| 16 |
S |
-0.160 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.606 |
0.730 |
0.696 |
1.896 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.235 |
2.671 |
1.374 |
| y |
2.671 |
-45.486 |
1.071 |
| z |
1.374 |
1.071 |
-46.052 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.466 |
2.671 |
1.374 |
| y |
2.671 |
2.158 |
1.071 |
| z |
1.374 |
1.071 |
1.309 |
|
| Polar |
|---|
| 3z2-r2 | 2.617 |
|---|
| x2-y2 | -3.749 |
|---|
| xy | 2.671 |
|---|
| xz | 1.374 |
|---|
| yz | 1.071 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.853 |
0.168 |
-0.111 |
| y |
0.168 |
10.236 |
0.168 |
| z |
-0.111 |
0.168 |
8.922 |
<r2> (average value of r
2) Å
2
| <r2> |
229.389 |
| (<r2>)1/2 |
15.146 |