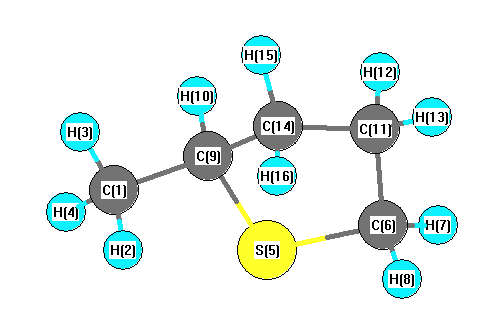
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3251 |
2947 |
28.00 |
|
|
|
| 2 |
A |
3235 |
2932 |
54.99 |
|
|
|
| 3 |
A |
3215 |
2914 |
67.61 |
|
|
|
| 4 |
A |
3214 |
2913 |
52.24 |
|
|
|
| 5 |
A |
3205 |
2905 |
23.81 |
|
|
|
| 6 |
A |
3195 |
2896 |
29.30 |
|
|
|
| 7 |
A |
3180 |
2882 |
6.69 |
|
|
|
| 8 |
A |
3168 |
2872 |
24.98 |
|
|
|
| 9 |
A |
3161 |
2865 |
58.75 |
|
|
|
| 10 |
A |
3155 |
2859 |
13.65 |
|
|
|
| 11 |
A |
1632 |
1479 |
0.47 |
|
|
|
| 12 |
A |
1615 |
1464 |
5.80 |
|
|
|
| 13 |
A |
1612 |
1461 |
3.40 |
|
|
|
| 14 |
A |
1609 |
1459 |
5.26 |
|
|
|
| 15 |
A |
1606 |
1456 |
2.70 |
|
|
|
| 16 |
A |
1538 |
1394 |
2.40 |
|
|
|
| 17 |
A |
1507 |
1366 |
1.82 |
|
|
|
| 18 |
A |
1478 |
1339 |
0.79 |
|
|
|
| 19 |
A |
1443 |
1308 |
0.07 |
|
|
|
| 20 |
A |
1423 |
1290 |
3.89 |
|
|
|
| 21 |
A |
1402 |
1270 |
26.66 |
|
|
|
| 22 |
A |
1345 |
1219 |
3.32 |
|
|
|
| 23 |
A |
1307 |
1184 |
4.12 |
|
|
|
| 24 |
A |
1257 |
1140 |
0.86 |
|
|
|
| 25 |
A |
1188 |
1077 |
0.09 |
|
|
|
| 26 |
A |
1162 |
1054 |
1.99 |
|
|
|
| 27 |
A |
1118 |
1014 |
4.07 |
|
|
|
| 28 |
A |
1095 |
993 |
0.23 |
|
|
|
| 29 |
A |
1051 |
952 |
1.80 |
|
|
|
| 30 |
A |
1026 |
930 |
2.01 |
|
|
|
| 31 |
A |
954 |
865 |
0.53 |
|
|
|
| 32 |
A |
929 |
842 |
1.60 |
|
|
|
| 33 |
A |
905 |
820 |
1.47 |
|
|
|
| 34 |
A |
749 |
679 |
3.27 |
|
|
|
| 35 |
A |
689 |
624 |
3.83 |
|
|
|
| 36 |
A |
567 |
514 |
0.45 |
|
|
|
| 37 |
A |
497 |
450 |
0.05 |
|
|
|
| 38 |
A |
441 |
399 |
0.52 |
|
|
|
| 39 |
A |
306 |
277 |
0.80 |
|
|
|
| 40 |
A |
271 |
246 |
0.13 |
|
|
|
| 41 |
A |
257 |
233 |
0.15 |
|
|
|
| 42 |
A |
95 |
86 |
1.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33025.5 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 29934.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.209 |
|
|
|
| 2 |
H |
0.056 |
|
|
|
| 3 |
H |
0.055 |
|
|
|
| 4 |
H |
0.070 |
|
|
|
| 5 |
S |
-0.133 |
|
|
|
| 6 |
C |
-0.089 |
|
|
|
| 7 |
H |
0.071 |
|
|
|
| 8 |
H |
0.063 |
|
|
|
| 9 |
C |
0.081 |
|
|
|
| 10 |
H |
0.042 |
|
|
|
| 11 |
C |
-0.083 |
|
|
|
| 12 |
H |
0.049 |
|
|
|
| 13 |
H |
0.052 |
|
|
|
| 14 |
C |
-0.124 |
|
|
|
| 15 |
H |
0.050 |
|
|
|
| 16 |
H |
0.048 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.395 |
2.129 |
-0.001 |
2.165 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.006 |
-0.385 |
1.044 |
| y |
-0.385 |
-49.048 |
-0.374 |
| z |
1.044 |
-0.374 |
-46.329 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.682 |
-0.385 |
1.044 |
| y |
-0.385 |
-4.380 |
-0.374 |
| z |
1.044 |
-0.374 |
-0.302 |
|
| Polar |
|---|
| 3z2-r2 | -0.604 |
|---|
| x2-y2 | 6.042 |
|---|
| xy | -0.385 |
|---|
| xz | 1.044 |
|---|
| yz | -0.374 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.222 |
-0.094 |
0.249 |
| y |
-0.094 |
11.034 |
-0.046 |
| z |
0.249 |
-0.046 |
8.886 |
<r2> (average value of r
2) Å
2
| <r2> |
202.887 |
| (<r2>)1/2 |
14.244 |