All results from a given calculation for CHBr2F (dibromofluoromethane)
using model chemistry: QCISD(T)=FULL/Def2TZVPP
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at QCISD(T)=FULL/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -5283.944883 |
| Energy at 298.15K | |
| HF Energy | -5282.757201 |
| Nuclear repulsion energy | 483.188020 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/Def2TZVPP
Geometric Data calculated at QCISD(T)=FULL/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.103 |
0.783 |
0.000 |
| H2 |
-1.005 |
1.378 |
0.000 |
| F3 |
0.977 |
1.591 |
0.000 |
| Br4 |
-0.103 |
-0.291 |
1.599 |
| Br5 |
-0.103 |
-0.291 |
-1.599 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
F3 |
Br4 |
Br5 |
| C1 | | 1.0807 | 1.3484 | 1.9266 | 1.9266 |
H2 | 1.0807 | | 1.9934 | 2.4816 | 2.4816 | F3 | 1.3484 | 1.9934 | | 2.6955 | 2.6955 | Br4 | 1.9266 | 2.4816 | 2.6955 | | 3.1981 | Br5 | 1.9266 | 2.4816 | 2.6955 | 3.1981 | |
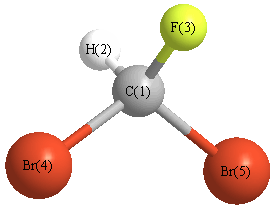 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| H2 |
C1 |
F3 |
109.803 |
|
H2 |
C1 |
Br4 |
107.879 |
| H2 |
C1 |
Br5 |
107.879 |
|
F3 |
C1 |
Br4 |
109.518 |
| F3 |
C1 |
Br5 |
109.518 |
|
Br4 |
C1 |
Br5 |
112.199 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
