Jump to
S1C2
S1C3
Energy calculated at PBE1PBE/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -1592.239181 |
| Energy at 298.15K | |
| HF Energy | -1592.239181 |
| Nuclear repulsion energy | 353.850516 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
544 |
544 |
0.00 |
36.37 |
0.04 |
0.09 |
| 2 |
A1 |
211 |
211 |
0.00 |
2.35 |
0.74 |
0.85 |
| 3 |
B1 |
483 |
483 |
0.00 |
15.89 |
0.75 |
0.86 |
| 4 |
B2 |
303 |
303 |
0.00 |
7.94 |
0.75 |
0.86 |
| 5 |
E |
454 |
454 |
0.00 |
4.06 |
0.75 |
0.86 |
| 5 |
E |
454 |
454 |
0.00 |
4.06 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1224.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1224.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/Def2TZVPP
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.416 |
0.314 |
| S2 |
0.000 |
-1.416 |
0.314 |
| S3 |
-1.416 |
0.000 |
-0.314 |
| S4 |
1.416 |
0.000 |
-0.314 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8322 | 2.0988 | 2.0988 |
S2 | 2.8322 | | 2.0988 | 2.0988 | S3 | 2.0988 | 2.0988 | | 2.8322 | S4 | 2.0988 | 2.0988 | 2.8322 | |
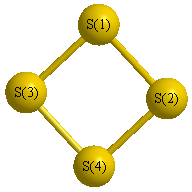 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.567 |
|
S1 |
S3 |
S4 |
47.567 |
| S2 |
S1 |
S3 |
47.567 |
|
S2 |
S4 |
S3 |
47.567 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.000 |
|
|
|
| 2 |
S |
0.000 |
|
|
|
| 3 |
S |
0.000 |
|
|
|
| 4 |
S |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.570 |
0.000 |
0.000 |
| y |
0.000 |
-48.570 |
0.000 |
| z |
0.000 |
0.000 |
-53.975 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.702 |
0.000 |
0.000 |
| y |
0.000 |
2.702 |
0.000 |
| z |
0.000 |
0.000 |
-5.405 |
|
| Polar |
|---|
| 3z2-r2 | -10.810 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.180 |
0.000 |
0.000 |
| y |
0.000 |
11.180 |
0.000 |
| z |
0.000 |
0.000 |
6.574 |
<r2> (average value of r
2) Å
2
| <r2> |
166.110 |
| (<r2>)1/2 |
12.888 |
Jump to
S1C1
S1C3
Energy calculated at PBE1PBE/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -1592.262663 |
| Energy at 298.15K | |
| HF Energy | -1592.262663 |
| Nuclear repulsion energy | 335.732644 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
711 |
711 |
7.73 |
24.32 |
0.16 |
0.27 |
| 2 |
A1 |
420 |
420 |
4.91 |
40.97 |
0.19 |
0.32 |
| 3 |
A1 |
118 |
118 |
0.33 |
10.26 |
0.45 |
0.62 |
| 4 |
A2 |
224 |
224 |
0.00 |
0.12 |
0.75 |
0.86 |
| 5 |
B2 |
680 |
680 |
123.19 |
3.88 |
0.75 |
0.86 |
| 6 |
B2 |
347 |
347 |
0.21 |
11.05 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1249.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1249.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/Def2TZVPP
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.037 |
0.908 |
| S2 |
0.000 |
-1.037 |
0.908 |
| S3 |
0.000 |
1.588 |
-0.908 |
| S4 |
0.000 |
-1.588 |
-0.908 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.0733 | 1.8970 | 3.1910 |
S2 | 2.0733 | | 3.1910 | 1.8970 | S3 | 1.8970 | 3.1910 | | 3.1755 | S4 | 3.1910 | 1.8970 | 3.1755 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
106.888 |
|
S1 |
S3 |
S4 |
73.112 |
| S2 |
S1 |
S3 |
106.888 |
|
S2 |
S4 |
S3 |
73.112 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.139 |
|
|
|
| 2 |
S |
0.139 |
|
|
|
| 3 |
S |
-0.139 |
|
|
|
| 4 |
S |
-0.139 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.441 |
1.441 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.934 |
0.000 |
0.000 |
| y |
0.000 |
-52.992 |
0.000 |
| z |
0.000 |
0.000 |
-48.198 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.339 |
0.000 |
0.000 |
| y |
0.000 |
-3.425 |
0.000 |
| z |
0.000 |
0.000 |
3.764 |
|
| Polar |
|---|
| 3z2-r2 | 7.529 |
|---|
| x2-y2 | 2.058 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.723 |
0.000 |
0.000 |
| y |
0.000 |
18.003 |
0.000 |
| z |
0.000 |
0.000 |
11.693 |
<r2> (average value of r
2) Å
2
| <r2> |
199.447 |
| (<r2>)1/2 |
14.123 |
Jump to
S1C1
S1C2
Energy calculated at PBE1PBE/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -1592.255447 |
| Energy at 298.15K | |
| HF Energy | -1592.255447 |
| Nuclear repulsion energy | 340.245657 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
764 |
764 |
0.00 |
32.17 |
0.20 |
0.34 |
| 2 |
Ag |
312 |
312 |
0.00 |
36.39 |
0.24 |
0.39 |
| 3 |
Au |
263 |
263 |
0.00 |
0.00 |
0.00 |
0.00 |
| 4 |
B1u |
701 |
701 |
125.29 |
0.00 |
0.00 |
0.00 |
| 5 |
B2u |
169i |
169i |
18.44 |
0.00 |
0.00 |
0.00 |
| 6 |
B3g |
355 |
355 |
0.00 |
12.76 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1113.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1113.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/Def2TZVPP
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.939 |
1.242 |
| S2 |
0.000 |
0.939 |
-1.242 |
| S3 |
0.000 |
-0.939 |
1.242 |
| S4 |
0.000 |
-0.939 |
-1.242 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.4849 | 1.8785 | 3.1150 |
S2 | 2.4849 | | 3.1150 | 1.8785 | S3 | 1.8785 | 3.1150 | | 2.4849 | S4 | 3.1150 | 1.8785 | 2.4849 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.000 |
|
|
|
| 2 |
S |
0.000 |
|
|
|
| 3 |
S |
0.000 |
|
|
|
| 4 |
S |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.284 |
0.000 |
0.000 |
| y |
0.000 |
-47.468 |
0.000 |
| z |
0.000 |
0.000 |
-52.592 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.253 |
0.000 |
0.000 |
| y |
0.000 |
4.470 |
0.000 |
| z |
0.000 |
0.000 |
-3.216 |
|
| Polar |
|---|
| 3z2-r2 | -6.432 |
|---|
| x2-y2 | -3.815 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.698 |
0.000 |
0.000 |
| y |
0.000 |
12.204 |
0.000 |
| z |
0.000 |
0.000 |
16.978 |
<r2> (average value of r
2) Å
2
| <r2> |
186.763 |
| (<r2>)1/2 |
13.666 |
