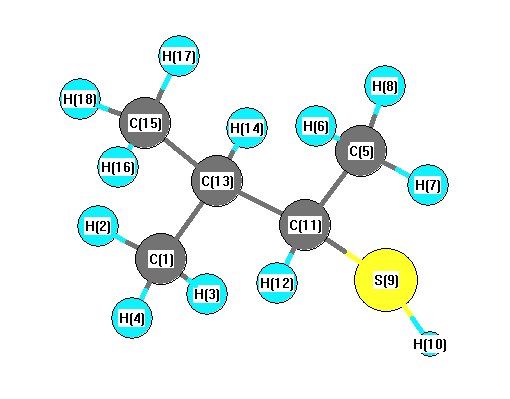
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3248 |
2944 |
29.95 |
|
|
|
| 2 |
A |
3245 |
2941 |
45.57 |
|
|
|
| 3 |
A |
3235 |
2932 |
51.72 |
|
|
|
| 4 |
A |
3228 |
2926 |
43.96 |
|
|
|
| 5 |
A |
3220 |
2919 |
82.04 |
|
|
|
| 6 |
A |
3208 |
2908 |
1.49 |
|
|
|
| 7 |
A |
3187 |
2888 |
20.87 |
|
|
|
| 8 |
A |
3170 |
2873 |
22.74 |
|
|
|
| 9 |
A |
3161 |
2865 |
38.77 |
|
|
|
| 10 |
A |
3155 |
2859 |
16.83 |
|
|
|
| 11 |
A |
3146 |
2852 |
4.03 |
|
|
|
| 12 |
A |
2861 |
2593 |
6.27 |
|
|
|
| 13 |
A |
1639 |
1486 |
2.48 |
|
|
|
| 14 |
A |
1623 |
1471 |
9.07 |
|
|
|
| 15 |
A |
1622 |
1470 |
11.86 |
|
|
|
| 16 |
A |
1611 |
1460 |
2.57 |
|
|
|
| 17 |
A |
1610 |
1459 |
2.83 |
|
|
|
| 18 |
A |
1604 |
1454 |
1.17 |
|
|
|
| 19 |
A |
1551 |
1405 |
4.63 |
|
|
|
| 20 |
A |
1540 |
1396 |
5.13 |
|
|
|
| 21 |
A |
1531 |
1388 |
2.48 |
|
|
|
| 22 |
A |
1510 |
1369 |
1.28 |
|
|
|
| 23 |
A |
1470 |
1332 |
0.49 |
|
|
|
| 24 |
A |
1438 |
1304 |
4.00 |
|
|
|
| 25 |
A |
1368 |
1240 |
29.25 |
|
|
|
| 26 |
A |
1294 |
1172 |
2.52 |
|
|
|
| 27 |
A |
1278 |
1159 |
7.60 |
|
|
|
| 28 |
A |
1237 |
1121 |
0.96 |
|
|
|
| 29 |
A |
1167 |
1058 |
2.90 |
|
|
|
| 30 |
A |
1118 |
1013 |
7.31 |
|
|
|
| 31 |
A |
1064 |
965 |
1.11 |
|
|
|
| 32 |
A |
1038 |
941 |
0.08 |
|
|
|
| 33 |
A |
1006 |
911 |
0.49 |
|
|
|
| 34 |
A |
982 |
891 |
1.77 |
|
|
|
| 35 |
A |
973 |
881 |
2.98 |
|
|
|
| 36 |
A |
843 |
764 |
2.82 |
|
|
|
| 37 |
A |
739 |
670 |
3.43 |
|
|
|
| 38 |
A |
508 |
461 |
0.42 |
|
|
|
| 39 |
A |
434 |
394 |
0.51 |
|
|
|
| 40 |
A |
400 |
363 |
0.12 |
|
|
|
| 41 |
A |
386 |
350 |
0.16 |
|
|
|
| 42 |
A |
357 |
324 |
1.17 |
|
|
|
| 43 |
A |
276 |
250 |
1.00 |
|
|
|
| 44 |
A |
255 |
231 |
3.45 |
|
|
|
| 45 |
A |
245 |
222 |
0.33 |
|
|
|
| 46 |
A |
210 |
191 |
1.22 |
|
|
|
| 47 |
A |
188 |
170 |
13.40 |
|
|
|
| 48 |
A |
61 |
55 |
1.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37118.1 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 33643.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.201 |
|
|
|
| 2 |
H |
0.057 |
|
|
|
| 3 |
H |
0.077 |
|
|
|
| 4 |
H |
0.047 |
|
|
|
| 5 |
C |
-0.208 |
|
|
|
| 6 |
H |
0.057 |
|
|
|
| 7 |
H |
0.058 |
|
|
|
| 8 |
H |
0.066 |
|
|
|
| 9 |
S |
-0.172 |
|
|
|
| 10 |
H |
0.057 |
|
|
|
| 11 |
C |
0.058 |
|
|
|
| 12 |
H |
0.039 |
|
|
|
| 13 |
C |
0.087 |
|
|
|
| 14 |
H |
0.022 |
|
|
|
| 15 |
C |
-0.227 |
|
|
|
| 16 |
H |
0.054 |
|
|
|
| 17 |
H |
0.064 |
|
|
|
| 18 |
H |
0.066 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.967 |
1.338 |
0.636 |
1.769 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.668 |
2.575 |
1.930 |
| y |
2.575 |
-49.041 |
0.765 |
| z |
1.930 |
0.765 |
-48.324 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.014 |
2.575 |
1.930 |
| y |
2.575 |
-1.045 |
0.765 |
| z |
1.930 |
0.765 |
0.031 |
|
| Polar |
|---|
| 3z2-r2 | 0.061 |
|---|
| x2-y2 | 1.373 |
|---|
| xy | 2.575 |
|---|
| xz | 1.930 |
|---|
| yz | 0.765 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.902 |
-0.330 |
-0.016 |
| y |
-0.330 |
11.568 |
0.187 |
| z |
-0.016 |
0.187 |
9.403 |
<r2> (average value of r
2) Å
2
| <r2> |
249.804 |
| (<r2>)1/2 |
15.805 |