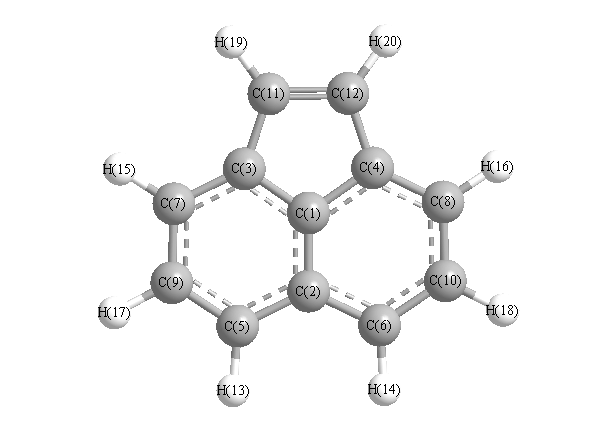
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3377 |
3061 |
16.92 |
|
|
|
| 2 |
A1 |
3343 |
3030 |
13.82 |
|
|
|
| 3 |
A1 |
3328 |
3017 |
28.40 |
|
|
|
| 4 |
A1 |
3315 |
3005 |
1.36 |
|
|
|
| 5 |
A1 |
1802 |
1633 |
0.32 |
|
|
|
| 6 |
A1 |
1790 |
1622 |
0.15 |
|
|
|
| 7 |
A1 |
1696 |
1537 |
15.88 |
|
|
|
| 8 |
A1 |
1566 |
1419 |
3.21 |
|
|
|
| 9 |
A1 |
1532 |
1389 |
28.99 |
|
|
|
| 10 |
A1 |
1482 |
1343 |
9.93 |
|
|
|
| 11 |
A1 |
1364 |
1236 |
0.75 |
|
|
|
| 12 |
A1 |
1286 |
1166 |
1.53 |
|
|
|
| 13 |
A1 |
1195 |
1083 |
6.29 |
|
|
|
| 14 |
A1 |
1114 |
1010 |
3.86 |
|
|
|
| 15 |
A1 |
1089 |
987 |
1.60 |
|
|
|
| 16 |
A1 |
870 |
788 |
2.28 |
|
|
|
| 17 |
A1 |
703 |
637 |
0.53 |
|
|
|
| 18 |
A1 |
598 |
542 |
3.10 |
|
|
|
| 19 |
A1 |
449 |
407 |
2.01 |
|
|
|
| 20 |
A2 |
1098 |
995 |
0.00 |
|
|
|
| 21 |
A2 |
1072 |
971 |
0.00 |
|
|
|
| 22 |
A2 |
1031 |
935 |
0.00 |
|
|
|
| 23 |
A2 |
852 |
772 |
0.00 |
|
|
|
| 24 |
A2 |
736 |
667 |
0.00 |
|
|
|
| 25 |
A2 |
632 |
573 |
0.00 |
|
|
|
| 26 |
A2 |
410 |
371 |
0.00 |
|
|
|
| 27 |
A2 |
233 |
212 |
0.00 |
|
|
|
| 28 |
B1 |
1105 |
1001 |
1.04 |
|
|
|
| 29 |
B1 |
1042 |
945 |
4.84 |
|
|
|
| 30 |
B1 |
944 |
856 |
54.46 |
|
|
|
| 31 |
B1 |
877 |
795 |
50.20 |
|
|
|
| 32 |
B1 |
828 |
751 |
61.18 |
|
|
|
| 33 |
B1 |
668 |
605 |
0.31 |
|
|
|
| 34 |
B1 |
506 |
458 |
0.00 |
|
|
|
| 35 |
B1 |
253 |
230 |
7.01 |
|
|
|
| 36 |
B1 |
177 |
160 |
7.20 |
|
|
|
| 37 |
B2 |
3355 |
3041 |
1.57 |
|
|
|
| 38 |
B2 |
3341 |
3029 |
51.45 |
|
|
|
| 39 |
B2 |
3327 |
3016 |
5.07 |
|
|
|
| 40 |
B2 |
3315 |
3004 |
0.75 |
|
|
|
| 41 |
B2 |
1814 |
1644 |
1.10 |
|
|
|
| 42 |
B2 |
1634 |
1481 |
24.60 |
|
|
|
| 43 |
B2 |
1612 |
1461 |
11.13 |
|
|
|
| 44 |
B2 |
1516 |
1374 |
3.54 |
|
|
|
| 45 |
B2 |
1424 |
1290 |
0.03 |
|
|
|
| 46 |
B2 |
1330 |
1206 |
0.53 |
|
|
|
| 47 |
B2 |
1304 |
1182 |
3.41 |
|
|
|
| 48 |
B2 |
1222 |
1108 |
13.85 |
|
|
|
| 49 |
B2 |
1188 |
1076 |
2.67 |
|
|
|
| 50 |
B2 |
1053 |
954 |
2.41 |
|
|
|
| 51 |
B2 |
937 |
849 |
4.04 |
|
|
|
| 52 |
B2 |
734 |
665 |
0.28 |
|
|
|
| 53 |
B2 |
553 |
502 |
1.43 |
|
|
|
| 54 |
B2 |
499 |
452 |
1.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37256.6 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 33769.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.328 |
|
|
|
| 2 |
C |
-0.126 |
|
|
|
| 3 |
C |
-0.107 |
|
|
|
| 4 |
C |
-0.107 |
|
|
|
| 5 |
C |
-0.154 |
|
|
|
| 6 |
C |
-0.154 |
|
|
|
| 7 |
C |
-0.090 |
|
|
|
| 8 |
C |
-0.090 |
|
|
|
| 9 |
C |
-0.022 |
|
|
|
| 10 |
C |
-0.022 |
|
|
|
| 11 |
C |
-0.121 |
|
|
|
| 12 |
C |
-0.121 |
|
|
|
| 13 |
H |
0.093 |
|
|
|
| 14 |
H |
0.093 |
|
|
|
| 15 |
H |
0.094 |
|
|
|
| 16 |
H |
0.094 |
|
|
|
| 17 |
H |
0.098 |
|
|
|
| 18 |
H |
0.098 |
|
|
|
| 19 |
H |
0.108 |
|
|
|
| 20 |
H |
0.108 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.081 |
0.081 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-76.360 |
0.000 |
0.000 |
| y |
0.000 |
-59.745 |
0.000 |
| z |
0.000 |
0.000 |
-61.290 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-15.843 |
0.000 |
0.000 |
| y |
0.000 |
9.080 |
0.000 |
| z |
0.000 |
0.000 |
6.762 |
|
| Polar |
|---|
| 3z2-r2 | 13.525 |
|---|
| x2-y2 | -16.615 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.045 |
0.000 |
0.000 |
| y |
0.000 |
26.082 |
0.000 |
| z |
0.000 |
0.000 |
21.719 |
<r2> (average value of r
2) Å
2
| <r2> |
456.683 |
| (<r2>)1/2 |
21.370 |