Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -192.264808 |
| Energy at 298.15K | -192.269692 |
| HF Energy | -192.264808 |
| Nuclear repulsion energy | 149.441547 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3388 |
3071 |
0.17 |
|
|
|
| 2 |
A1 |
3367 |
3052 |
23.16 |
|
|
|
| 3 |
A1 |
3350 |
3036 |
5.05 |
|
|
|
| 4 |
A1 |
1565 |
1418 |
16.84 |
|
|
|
| 5 |
A1 |
1504 |
1363 |
33.12 |
|
|
|
| 6 |
A1 |
1197 |
1085 |
2.38 |
|
|
|
| 7 |
A1 |
1094 |
992 |
0.70 |
|
|
|
| 8 |
A1 |
971 |
880 |
0.65 |
|
|
|
| 9 |
A1 |
895 |
811 |
0.22 |
|
|
|
| 10 |
A2 |
961 |
871 |
0.00 |
|
|
|
| 11 |
A2 |
768 |
696 |
0.00 |
|
|
|
| 12 |
A2 |
537 |
486 |
0.00 |
|
|
|
| 13 |
B1 |
967 |
877 |
9.85 |
|
|
|
| 14 |
B1 |
836 |
758 |
1.57 |
|
|
|
| 15 |
B1 |
714 |
648 |
126.79 |
|
|
|
| 16 |
B1 |
543 |
492 |
5.73 |
|
|
|
| 17 |
B2 |
3377 |
3061 |
17.58 |
|
|
|
| 18 |
B2 |
3356 |
3042 |
0.13 |
|
|
|
| 19 |
B2 |
1599 |
1449 |
0.11 |
|
|
|
| 20 |
B2 |
1408 |
1277 |
0.17 |
|
|
|
| 21 |
B2 |
1333 |
1208 |
0.39 |
|
|
|
| 22 |
B2 |
1114 |
1010 |
10.37 |
|
|
|
| 23 |
B2 |
995 |
902 |
0.55 |
|
|
|
| 24 |
B2 |
704 |
638 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18271.3 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 16561.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.187 |
| C2 |
0.000 |
1.162 |
0.349 |
| C3 |
0.000 |
0.742 |
-0.942 |
| C4 |
0.000 |
-0.742 |
-0.942 |
| C5 |
0.000 |
-1.162 |
0.349 |
| H6 |
0.000 |
0.000 |
2.258 |
| H7 |
0.000 |
2.173 |
0.693 |
| H8 |
0.000 |
1.353 |
-1.821 |
| H9 |
0.000 |
-1.353 |
-1.821 |
| H10 |
0.000 |
-2.173 |
0.693 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4329 | 2.2547 | 2.2547 | 1.4329 | 1.0710 | 2.2288 | 3.2982 | 3.2982 | 2.2288 |
C2 | 1.4329 | | 1.3579 | 2.3004 | 2.3244 | 2.2350 | 1.0681 | 2.1781 | 3.3220 | 3.3533 | C3 | 2.2547 | 1.3579 | | 1.4834 | 2.3004 | 3.2851 | 2.1735 | 1.0705 | 2.2718 | 3.3424 | C4 | 2.2547 | 2.3004 | 1.4834 | | 1.3579 | 3.2851 | 3.3424 | 2.2718 | 1.0705 | 2.1735 | C5 | 1.4329 | 2.3244 | 2.3004 | 1.3579 | | 2.2350 | 3.3533 | 3.3220 | 2.1781 | 1.0681 | H6 | 1.0710 | 2.2350 | 3.2851 | 3.2851 | 2.2350 | | 2.6782 | 4.2974 | 4.2974 | 2.6782 | H7 | 2.2288 | 1.0681 | 2.1735 | 3.3424 | 3.3533 | 2.6782 | | 2.6442 | 4.3310 | 4.3467 | H8 | 3.2982 | 2.1781 | 1.0705 | 2.2718 | 3.3220 | 4.2974 | 2.6442 | | 2.7068 | 4.3310 | H9 | 3.2982 | 3.3220 | 2.2718 | 1.0705 | 2.1781 | 4.2974 | 4.3310 | 2.7068 | | 2.6442 | H10 | 2.2288 | 3.3533 | 3.3424 | 2.1735 | 1.0681 | 2.6782 | 4.3467 | 4.3310 | 2.6442 | |
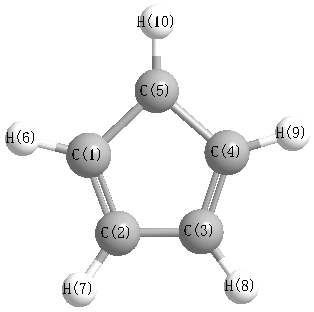 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
107.757 |
|
C1 |
C2 |
H7 |
125.408 |
| C1 |
C5 |
C4 |
107.757 |
|
C1 |
C5 |
H10 |
125.408 |
| C2 |
C1 |
C5 |
108.407 |
|
C2 |
C1 |
H6 |
125.796 |
| C2 |
C3 |
C4 |
108.040 |
|
C2 |
C3 |
H8 |
127.109 |
| C3 |
C2 |
H7 |
126.835 |
|
C3 |
C4 |
C5 |
108.040 |
| C3 |
C4 |
H9 |
124.851 |
|
C4 |
C3 |
H8 |
124.851 |
| C4 |
C5 |
H10 |
126.835 |
|
C5 |
C1 |
H6 |
125.796 |
| C5 |
C4 |
H9 |
127.109 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.008 |
|
|
|
| 2 |
C |
-0.202 |
|
|
|
| 3 |
C |
-0.054 |
|
|
|
| 4 |
C |
-0.054 |
|
|
|
| 5 |
C |
-0.202 |
|
|
|
| 6 |
H |
0.107 |
|
|
|
| 7 |
H |
0.092 |
|
|
|
| 8 |
H |
0.107 |
|
|
|
| 9 |
H |
0.107 |
|
|
|
| 10 |
H |
0.092 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.157 |
0.157 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.697 |
0.000 |
0.000 |
| y |
0.000 |
-27.904 |
0.000 |
| z |
0.000 |
0.000 |
-25.775 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.857 |
0.000 |
0.000 |
| y |
0.000 |
1.832 |
0.000 |
| z |
0.000 |
0.000 |
5.026 |
|
| Polar |
|---|
| 3z2-r2 | 10.051 |
|---|
| x2-y2 | -5.793 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.926 |
0.000 |
0.000 |
| y |
0.000 |
7.994 |
0.000 |
| z |
0.000 |
0.000 |
8.969 |
<r2> (average value of r
2) Å
2
| <r2> |
87.412 |
| (<r2>)1/2 |
9.349 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -192.263849 |
| Energy at 298.15K | |
| HF Energy | -192.263849 |
| Nuclear repulsion energy | 149.594346 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3389 |
3072 |
0.13 |
|
|
|
| 2 |
A1 |
3380 |
3063 |
16.11 |
|
|
|
| 3 |
A1 |
3353 |
3039 |
0.60 |
|
|
|
| 4 |
A1 |
1714 |
1553 |
0.02 |
|
|
|
| 5 |
A1 |
1547 |
1402 |
0.31 |
|
|
|
| 6 |
A1 |
1220 |
1106 |
0.40 |
|
|
|
| 7 |
A1 |
1129 |
1023 |
8.07 |
|
|
|
| 8 |
A1 |
1076 |
976 |
3.35 |
|
|
|
| 9 |
A1 |
896 |
812 |
0.35 |
|
|
|
| 10 |
A2 |
1048 |
950 |
0.00 |
|
|
|
| 11 |
A2 |
827 |
750 |
0.00 |
|
|
|
| 12 |
A2 |
564 |
511 |
0.00 |
|
|
|
| 13 |
B1 |
929 |
842 |
14.37 |
|
|
|
| 14 |
B1 |
803 |
728 |
19.80 |
|
|
|
| 15 |
B1 |
715 |
648 |
104.34 |
|
|
|
| 16 |
B1 |
524 |
475 |
7.51 |
|
|
|
| 17 |
B2 |
3365 |
3050 |
38.23 |
|
|
|
| 18 |
B2 |
3354 |
3040 |
6.26 |
|
|
|
| 19 |
B2 |
1503 |
1363 |
45.76 |
|
|
|
| 20 |
B2 |
1410 |
1278 |
0.01 |
|
|
|
| 21 |
B2 |
1266 |
1148 |
12.27 |
|
|
|
| 22 |
B2 |
1013 |
918 |
0.84 |
|
|
|
| 23 |
B2 |
922 |
836 |
0.01 |
|
|
|
| 24 |
B2 |
2223i |
2015i |
2036.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16862.1 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 15283.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.220 |
| C2 |
0.000 |
1.119 |
0.392 |
| C3 |
0.000 |
0.667 |
-1.001 |
| C4 |
0.000 |
-0.667 |
-1.001 |
| C5 |
0.000 |
-1.119 |
0.392 |
| H6 |
0.000 |
0.000 |
2.288 |
| H7 |
0.000 |
2.142 |
0.706 |
| H8 |
0.000 |
1.308 |
-1.856 |
| H9 |
0.000 |
-1.308 |
-1.856 |
| H10 |
0.000 |
-2.142 |
0.706 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.3920 | 2.3191 | 2.3191 | 1.3920 | 1.0680 | 2.2031 | 3.3427 | 3.3427 | 2.2031 |
C2 | 1.3920 | | 1.4645 | 2.2649 | 2.2378 | 2.2016 | 1.0707 | 2.2561 | 3.3080 | 3.2764 | C3 | 2.3191 | 1.4645 | | 1.3338 | 2.2649 | 3.3561 | 2.2567 | 1.0686 | 2.1518 | 3.2875 | C4 | 2.3191 | 2.2649 | 1.3338 | | 1.4645 | 3.3561 | 3.2875 | 2.1518 | 1.0686 | 2.2567 | C5 | 1.3920 | 2.2378 | 2.2649 | 1.4645 | | 2.2016 | 3.2764 | 3.3080 | 2.2561 | 1.0707 | H6 | 1.0680 | 2.2016 | 3.3561 | 3.3561 | 2.2016 | | 2.6630 | 4.3458 | 4.3458 | 2.6630 | H7 | 2.2031 | 1.0707 | 2.2567 | 3.2875 | 3.2764 | 2.6630 | | 2.6952 | 4.2977 | 4.2848 | H8 | 3.3427 | 2.2561 | 1.0686 | 2.1518 | 3.3080 | 4.3458 | 2.6952 | | 2.6154 | 4.2977 | H9 | 3.3427 | 3.3080 | 2.1518 | 1.0686 | 2.2561 | 4.3458 | 4.2977 | 2.6154 | | 2.6952 | H10 | 2.2031 | 3.2764 | 3.2875 | 2.2567 | 1.0707 | 2.6630 | 4.2848 | 4.2977 | 2.6952 | |
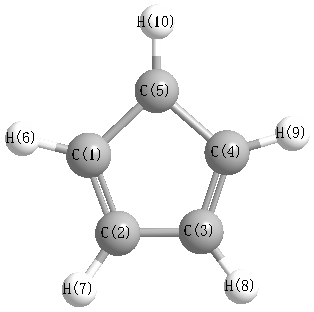 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
108.531 |
|
C1 |
C2 |
H7 |
126.417 |
| C1 |
C5 |
C4 |
108.531 |
|
C1 |
C5 |
H10 |
126.417 |
| C2 |
C1 |
C5 |
106.988 |
|
C2 |
C1 |
H6 |
126.506 |
| C2 |
C3 |
C4 |
107.975 |
|
C2 |
C3 |
H8 |
125.180 |
| C3 |
C2 |
H7 |
125.052 |
|
C3 |
C4 |
C5 |
107.975 |
| C3 |
C4 |
H9 |
126.845 |
|
C4 |
C3 |
H8 |
126.845 |
| C4 |
C5 |
H10 |
125.052 |
|
C5 |
C1 |
H6 |
126.506 |
| C5 |
C4 |
H9 |
125.180 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.228 |
|
|
|
| 2 |
C |
-0.001 |
|
|
|
| 3 |
C |
-0.137 |
|
|
|
| 4 |
C |
-0.137 |
|
|
|
| 5 |
C |
-0.001 |
|
|
|
| 6 |
H |
0.092 |
|
|
|
| 7 |
H |
0.109 |
|
|
|
| 8 |
H |
0.098 |
|
|
|
| 9 |
H |
0.098 |
|
|
|
| 10 |
H |
0.109 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.030 |
0.030 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.743 |
0.000 |
0.000 |
| y |
0.000 |
-25.710 |
0.000 |
| z |
0.000 |
0.000 |
-27.921 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.927 |
0.000 |
0.000 |
| y |
0.000 |
5.122 |
0.000 |
| z |
0.000 |
0.000 |
1.805 |
|
| Polar |
|---|
| 3z2-r2 | 3.611 |
|---|
| x2-y2 | -8.033 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.940 |
0.000 |
0.000 |
| y |
0.000 |
10.443 |
0.000 |
| z |
0.000 |
0.000 |
8.021 |
<r2> (average value of r
2) Å
2
| <r2> |
87.282 |
| (<r2>)1/2 |
9.343 |
