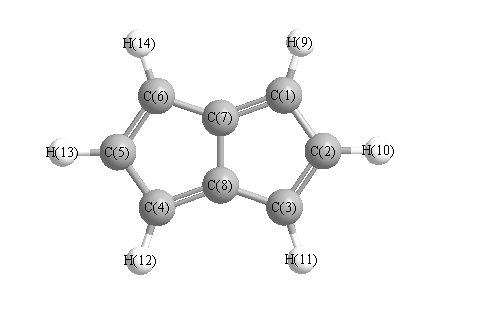
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3386 |
3069 |
0.00 |
|
|
|
| 2 |
Ag |
3363 |
3049 |
0.00 |
|
|
|
| 3 |
Ag |
3351 |
3038 |
0.00 |
|
|
|
| 4 |
Ag |
1825 |
1654 |
0.00 |
|
|
|
| 5 |
Ag |
1728 |
1567 |
0.00 |
|
|
|
| 6 |
Ag |
1492 |
1353 |
0.00 |
|
|
|
| 7 |
Ag |
1448 |
1312 |
0.00 |
|
|
|
| 8 |
Ag |
1287 |
1166 |
0.00 |
|
|
|
| 9 |
Ag |
1201 |
1089 |
0.00 |
|
|
|
| 10 |
Ag |
1006 |
912 |
0.00 |
|
|
|
| 11 |
Ag |
899 |
815 |
0.00 |
|
|
|
| 12 |
Ag |
771 |
699 |
0.00 |
|
|
|
| 13 |
Ag |
693 |
628 |
0.00 |
|
|
|
| 14 |
Au |
1064 |
965 |
2.29 |
|
|
|
| 15 |
Au |
997 |
904 |
31.04 |
|
|
|
| 16 |
Au |
823 |
746 |
113.14 |
|
|
|
| 17 |
Au |
562 |
509 |
11.23 |
|
|
|
| 18 |
Au |
319 |
290 |
3.27 |
|
|
|
| 19 |
Au |
191 |
173 |
8.95 |
|
|
|
| 20 |
Bg |
1065 |
966 |
0.00 |
|
|
|
| 21 |
Bg |
1013 |
918 |
0.00 |
|
|
|
| 22 |
Bg |
874 |
792 |
0.00 |
|
|
|
| 23 |
Bg |
732 |
663 |
0.00 |
|
|
|
| 24 |
Bg |
589 |
534 |
0.00 |
|
|
|
| 25 |
Bu |
3386 |
3069 |
24.00 |
|
|
|
| 26 |
Bu |
3364 |
3049 |
27.10 |
|
|
|
| 27 |
Bu |
3352 |
3039 |
10.18 |
|
|
|
| 28 |
Bu |
1816 |
1646 |
17.62 |
|
|
|
| 29 |
Bu |
1694 |
1536 |
34.40 |
|
|
|
| 30 |
Bu |
1462 |
1325 |
17.28 |
|
|
|
| 31 |
Bu |
1374 |
1245 |
16.18 |
|
|
|
| 32 |
Bu |
1209 |
1095 |
25.38 |
|
|
|
| 33 |
Bu |
1116 |
1012 |
28.12 |
|
|
|
| 34 |
Bu |
998 |
905 |
0.01 |
|
|
|
| 35 |
Bu |
907 |
822 |
11.60 |
|
|
|
| 36 |
Bu |
477 |
432 |
4.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25917.5 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 23491.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.077 |
|
|
|
| 2 |
C |
0.083 |
|
|
|
| 3 |
C |
-0.369 |
|
|
|
| 4 |
C |
-0.077 |
|
|
|
| 5 |
C |
0.083 |
|
|
|
| 6 |
C |
-0.369 |
|
|
|
| 7 |
C |
0.053 |
|
|
|
| 8 |
C |
0.053 |
|
|
|
| 9 |
H |
0.105 |
|
|
|
| 10 |
H |
0.110 |
|
|
|
| 11 |
H |
0.096 |
|
|
|
| 12 |
H |
0.105 |
|
|
|
| 13 |
H |
0.110 |
|
|
|
| 14 |
H |
0.096 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.563 |
1.613 |
0.000 |
| y |
1.613 |
-41.159 |
0.000 |
| z |
0.000 |
0.000 |
-51.852 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.942 |
1.613 |
0.000 |
| y |
1.613 |
5.048 |
0.000 |
| z |
0.000 |
0.000 |
-10.991 |
|
| Polar |
|---|
| 3z2-r2 | -21.982 |
|---|
| x2-y2 | 0.596 |
|---|
| xy | 1.613 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
18.060 |
1.598 |
0.000 |
| y |
1.598 |
12.403 |
0.000 |
| z |
0.000 |
0.000 |
7.192 |
<r2> (average value of r
2) Å
2
| <r2> |
215.677 |
| (<r2>)1/2 |
14.686 |