Jump to
S1C2
Energy calculated at QCISD/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -188.817618 |
| Energy at 298.15K | -188.818707 |
| HF Energy | -188.214699 |
| Nuclear repulsion energy | 63.252549 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3714 |
3714 |
37.99 |
|
|
|
| 2 |
A' |
1875 |
1875 |
306.01 |
|
|
|
| 3 |
A' |
1331 |
1331 |
0.14 |
|
|
|
| 4 |
A' |
1096 |
1096 |
188.65 |
|
|
|
| 5 |
A' |
609 |
609 |
34.27 |
|
|
|
| 6 |
A" |
577 |
577 |
112.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4600.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 4600.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.443 |
0.000 |
| O2 |
-1.061 |
-0.358 |
0.000 |
| O3 |
1.154 |
0.185 |
0.000 |
| H4 |
-0.746 |
-1.275 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
| C1 | | 1.3298 | 1.1828 | 1.8732 |
O2 | 1.3298 | | 2.2808 | 0.9694 | O3 | 1.1828 | 2.2808 | | 2.3960 | H4 | 1.8732 | 0.9694 | 2.3960 | |
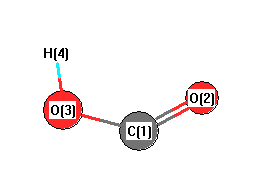 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
108.095 |
|
O2 |
C1 |
O3 |
130.300 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -188.820329 |
| Energy at 298.15K | -188.821395 |
| HF Energy | -188.215503 |
| Nuclear repulsion energy | 63.017891 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3871 |
3871 |
124.94 |
|
|
|
| 2 |
A' |
1919 |
1919 |
223.73 |
|
|
|
| 3 |
A' |
1276 |
1276 |
242.35 |
|
|
|
| 4 |
A' |
1101 |
1101 |
76.04 |
|
|
|
| 5 |
A' |
628 |
628 |
5.27 |
|
|
|
| 6 |
A" |
530 |
530 |
87.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4662.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 4662.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.408 |
0.000 |
| O2 |
-0.942 |
-0.548 |
0.000 |
| O3 |
1.167 |
0.256 |
0.000 |
| H4 |
-1.800 |
-0.114 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
| C1 | | 1.3422 | 1.1772 | 1.8738 |
O2 | 1.3422 | | 2.2579 | 0.9606 | O3 | 1.1772 | 2.2579 | | 2.9900 | H4 | 1.8738 | 0.9606 | 2.9900 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
107.761 |
|
O2 |
C1 |
O3 |
127.200 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
