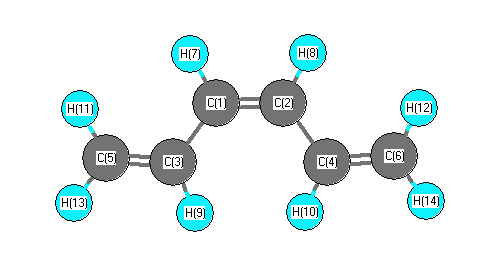
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3371 |
3056 |
36.59 |
|
|
|
| 2 |
A1 |
3335 |
3023 |
8.08 |
|
|
|
| 3 |
A1 |
3314 |
3004 |
16.10 |
|
|
|
| 4 |
A1 |
3284 |
2977 |
1.43 |
|
|
|
| 5 |
A1 |
1867 |
1692 |
0.01 |
|
|
|
| 6 |
A1 |
1762 |
1597 |
0.59 |
|
|
|
| 7 |
A1 |
1552 |
1407 |
0.92 |
|
|
|
| 8 |
A1 |
1456 |
1319 |
2.07 |
|
|
|
| 9 |
A1 |
1382 |
1253 |
0.34 |
|
|
|
| 10 |
A1 |
1180 |
1070 |
0.05 |
|
|
|
| 11 |
A1 |
942 |
854 |
0.20 |
|
|
|
| 12 |
A1 |
420 |
381 |
0.07 |
|
|
|
| 13 |
A1 |
180 |
163 |
0.13 |
|
|
|
| 14 |
A2 |
1144 |
1037 |
0.00 |
|
|
|
| 15 |
A2 |
1105 |
1002 |
0.00 |
|
|
|
| 16 |
A2 |
1064 |
964 |
0.00 |
|
|
|
| 17 |
A2 |
803 |
727 |
0.00 |
|
|
|
| 18 |
A2 |
372 |
337 |
0.00 |
|
|
|
| 19 |
A2 |
146 |
133 |
0.00 |
|
|
|
| 20 |
B1 |
1128 |
1022 |
19.62 |
|
|
|
| 21 |
B1 |
1069 |
969 |
126.84 |
|
|
|
| 22 |
B1 |
927 |
841 |
4.60 |
|
|
|
| 23 |
B1 |
657 |
595 |
37.08 |
|
|
|
| 24 |
B1 |
100 |
91 |
0.30 |
|
|
|
| 25 |
B2 |
3370 |
3054 |
0.28 |
|
|
|
| 26 |
B2 |
3316 |
3005 |
0.18 |
|
|
|
| 27 |
B2 |
3291 |
2983 |
5.29 |
|
|
|
| 28 |
B2 |
3283 |
2975 |
11.16 |
|
|
|
| 29 |
B2 |
1826 |
1655 |
25.33 |
|
|
|
| 30 |
B2 |
1605 |
1455 |
14.25 |
|
|
|
| 31 |
B2 |
1499 |
1359 |
1.13 |
|
|
|
| 32 |
B2 |
1415 |
1283 |
2.46 |
|
|
|
| 33 |
B2 |
1293 |
1172 |
1.74 |
|
|
|
| 34 |
B2 |
1025 |
929 |
5.25 |
|
|
|
| 35 |
B2 |
728 |
660 |
0.67 |
|
|
|
| 36 |
B2 |
377 |
342 |
16.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27794.5 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 25193.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.025 |
|
|
|
| 2 |
C |
-0.025 |
|
|
|
| 3 |
C |
-0.100 |
|
|
|
| 4 |
C |
-0.100 |
|
|
|
| 5 |
C |
-0.137 |
|
|
|
| 6 |
C |
-0.137 |
|
|
|
| 7 |
H |
0.069 |
|
|
|
| 8 |
H |
0.069 |
|
|
|
| 9 |
H |
0.062 |
|
|
|
| 10 |
H |
0.062 |
|
|
|
| 11 |
H |
0.057 |
|
|
|
| 12 |
H |
0.057 |
|
|
|
| 13 |
H |
0.075 |
|
|
|
| 14 |
H |
0.075 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.045 |
0.045 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.906 |
0.000 |
0.000 |
| y |
0.000 |
-33.823 |
0.000 |
| z |
0.000 |
0.000 |
-33.888 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.050 |
0.000 |
0.000 |
| y |
0.000 |
4.574 |
0.000 |
| z |
0.000 |
0.000 |
4.476 |
|
| Polar |
|---|
| 3z2-r2 | 8.952 |
|---|
| x2-y2 | -9.083 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.486 |
0.000 |
0.000 |
| y |
0.000 |
22.052 |
0.000 |
| z |
0.000 |
0.000 |
8.554 |
<r2> (average value of r
2) Å
2
| <r2> |
230.943 |
| (<r2>)1/2 |
15.197 |