Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -343.176236 |
| Energy at 298.15K | -343.193856 |
| HF Energy | -343.176236 |
| Nuclear repulsion energy | 428.119895 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3195 |
2896 |
0.00 |
|
|
|
| 2 |
A1' |
1648 |
1494 |
0.00 |
|
|
|
| 3 |
A1' |
1498 |
1358 |
0.00 |
|
|
|
| 4 |
A1' |
1052 |
953 |
0.00 |
|
|
|
| 5 |
A1' |
874 |
792 |
0.00 |
|
|
|
| 6 |
A1' |
653 |
592 |
0.00 |
|
|
|
| 7 |
A1" |
3194 |
2895 |
0.00 |
|
|
|
| 8 |
A1" |
1372 |
1243 |
0.00 |
|
|
|
| 9 |
A1" |
1119 |
1015 |
0.00 |
|
|
|
| 10 |
A1" |
63 |
57 |
0.00 |
|
|
|
| 11 |
A2' |
3220 |
2919 |
0.00 |
|
|
|
| 12 |
A2' |
1288 |
1168 |
0.00 |
|
|
|
| 13 |
A2' |
870 |
789 |
0.00 |
|
|
|
| 14 |
A2" |
3175 |
2878 |
127.35 |
|
|
|
| 15 |
A2" |
1634 |
1481 |
6.11 |
|
|
|
| 16 |
A2" |
1517 |
1375 |
6.85 |
|
|
|
| 17 |
A2" |
1076 |
976 |
34.01 |
|
|
|
| 18 |
A2" |
845 |
766 |
70.69 |
|
|
|
| 19 |
E' |
3232 |
2929 |
136.70 |
|
|
|
| 19 |
E' |
3232 |
2929 |
136.70 |
|
|
|
| 20 |
E' |
3182 |
2884 |
131.99 |
|
|
|
| 20 |
E' |
3182 |
2884 |
131.99 |
|
|
|
| 21 |
E' |
1630 |
1477 |
4.99 |
|
|
|
| 21 |
E' |
1630 |
1477 |
4.99 |
|
|
|
| 22 |
E' |
1476 |
1338 |
18.99 |
|
|
|
| 22 |
E' |
1476 |
1338 |
18.99 |
|
|
|
| 23 |
E' |
1448 |
1312 |
1.18 |
|
|
|
| 23 |
E' |
1448 |
1312 |
1.18 |
|
|
|
| 24 |
E' |
1188 |
1077 |
54.26 |
|
|
|
| 24 |
E' |
1188 |
1077 |
54.26 |
|
|
|
| 25 |
E' |
965 |
874 |
6.73 |
|
|
|
| 25 |
E' |
965 |
874 |
6.73 |
|
|
|
| 26 |
E' |
898 |
814 |
5.26 |
|
|
|
| 26 |
E' |
898 |
814 |
5.26 |
|
|
|
| 27 |
E' |
460 |
417 |
0.07 |
|
|
|
| 27 |
E' |
460 |
417 |
0.07 |
|
|
|
| 28 |
E" |
3200 |
2901 |
0.00 |
|
|
|
| 28 |
E" |
3200 |
2901 |
0.00 |
|
|
|
| 29 |
E" |
3168 |
2871 |
0.00 |
|
|
|
| 29 |
E" |
3168 |
2871 |
0.00 |
|
|
|
| 30 |
E" |
1616 |
1465 |
0.00 |
|
|
|
| 30 |
E" |
1616 |
1465 |
0.00 |
|
|
|
| 31 |
E" |
1476 |
1338 |
0.00 |
|
|
|
| 31 |
E" |
1476 |
1338 |
0.00 |
|
|
|
| 32 |
E" |
1438 |
1304 |
0.00 |
|
|
|
| 32 |
E" |
1438 |
1304 |
0.00 |
|
|
|
| 33 |
E" |
1326 |
1202 |
0.00 |
|
|
|
| 33 |
E" |
1326 |
1202 |
0.00 |
|
|
|
| 34 |
E" |
1135 |
1029 |
0.00 |
|
|
|
| 34 |
E" |
1135 |
1029 |
0.00 |
|
|
|
| 35 |
E" |
639 |
579 |
0.00 |
|
|
|
| 35 |
E" |
639 |
579 |
0.00 |
|
|
|
| 36 |
E" |
356 |
323 |
0.00 |
|
|
|
| 36 |
E" |
356 |
323 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42977.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 38955.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.269 |
| N2 |
0.000 |
0.000 |
-1.269 |
| C3 |
0.000 |
1.371 |
0.776 |
| C4 |
1.187 |
-0.685 |
0.776 |
| C5 |
-1.187 |
-0.685 |
0.776 |
| C6 |
0.000 |
1.371 |
-0.776 |
| C7 |
-1.187 |
-0.685 |
-0.776 |
| C8 |
1.187 |
-0.685 |
-0.776 |
| H9 |
0.871 |
1.879 |
1.170 |
| H10 |
-0.871 |
1.879 |
1.170 |
| H11 |
1.192 |
-1.694 |
1.170 |
| H12 |
2.063 |
-0.185 |
1.170 |
| H13 |
-2.063 |
-0.185 |
1.170 |
| H14 |
-1.192 |
-1.694 |
1.170 |
| H15 |
-0.871 |
1.879 |
-1.170 |
| H16 |
0.871 |
1.879 |
-1.170 |
| H17 |
-1.192 |
-1.694 |
-1.170 |
| H18 |
-2.063 |
-0.185 |
-1.170 |
| H19 |
2.063 |
-0.185 |
-1.170 |
| H20 |
1.192 |
-1.694 |
-1.170 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5372 | 1.4568 | 1.4568 | 1.4568 | 2.4616 | 2.4616 | 2.4616 | 2.0739 | 2.0739 | 2.0739 | 2.0739 | 2.0739 | 2.0739 | 3.1994 | 3.1994 | 3.1994 | 3.1994 | 3.1994 | 3.1994 |
N2 | 2.5372 | | 2.4616 | 2.4616 | 2.4616 | 1.4568 | 1.4568 | 1.4568 | 3.1994 | 3.1994 | 3.1994 | 3.1994 | 3.1994 | 3.1994 | 2.0739 | 2.0739 | 2.0739 | 2.0739 | 2.0739 | 2.0739 | C3 | 1.4568 | 2.4616 | | 2.3745 | 2.3745 | 1.5518 | 2.8366 | 2.8366 | 1.0829 | 1.0829 | 3.3122 | 2.6140 | 2.6140 | 3.3122 | 2.1915 | 2.1915 | 3.8211 | 3.2347 | 3.2347 | 3.8211 | C4 | 1.4568 | 2.4616 | 2.3745 | | 2.3745 | 2.8366 | 2.8366 | 1.5518 | 2.6140 | 3.3122 | 1.0829 | 1.0829 | 3.3122 | 2.6140 | 3.8211 | 3.2347 | 3.2347 | 3.8211 | 2.1915 | 2.1915 | C5 | 1.4568 | 2.4616 | 2.3745 | 2.3745 | | 2.8366 | 1.5518 | 2.8366 | 3.3122 | 2.6140 | 2.6140 | 3.3122 | 1.0829 | 1.0829 | 3.2347 | 3.8211 | 2.1915 | 2.1915 | 3.8211 | 3.2347 | C6 | 2.4616 | 1.4568 | 1.5518 | 2.8366 | 2.8366 | | 2.3745 | 2.3745 | 2.1915 | 2.1915 | 3.8211 | 3.2347 | 3.2347 | 3.8211 | 1.0829 | 1.0829 | 3.3122 | 2.6140 | 2.6140 | 3.3122 | C7 | 2.4616 | 1.4568 | 2.8366 | 2.8366 | 1.5518 | 2.3745 | | 2.3745 | 3.8211 | 3.2347 | 3.2347 | 3.8211 | 2.1915 | 2.1915 | 2.6140 | 3.3122 | 1.0829 | 1.0829 | 3.3122 | 2.6140 | C8 | 2.4616 | 1.4568 | 2.8366 | 1.5518 | 2.8366 | 2.3745 | 2.3745 | | 3.2347 | 3.8211 | 2.1915 | 2.1915 | 3.8211 | 3.2347 | 3.3122 | 2.6140 | 2.6140 | 3.3122 | 1.0829 | 1.0829 | H9 | 2.0739 | 3.1994 | 1.0829 | 2.6140 | 3.3122 | 2.1915 | 3.8211 | 3.2347 | | 1.7425 | 3.5879 | 2.3839 | 3.5879 | 4.1264 | 2.9169 | 2.3392 | 4.7433 | 4.2831 | 3.3399 | 4.2831 | H10 | 2.0739 | 3.1994 | 1.0829 | 3.3122 | 2.6140 | 2.1915 | 3.2347 | 3.8211 | 1.7425 | | 4.1264 | 3.5879 | 2.3839 | 3.5879 | 2.3392 | 2.9169 | 4.2831 | 3.3399 | 4.2831 | 4.7433 | H11 | 2.0739 | 3.1994 | 3.3122 | 1.0829 | 2.6140 | 3.8211 | 3.2347 | 2.1915 | 3.5879 | 4.1264 | | 1.7425 | 3.5879 | 2.3839 | 4.7433 | 4.2831 | 3.3399 | 4.2831 | 2.9169 | 2.3392 | H12 | 2.0739 | 3.1994 | 2.6140 | 1.0829 | 3.3122 | 3.2347 | 3.8211 | 2.1915 | 2.3839 | 3.5879 | 1.7425 | | 4.1264 | 3.5879 | 4.2831 | 3.3399 | 4.2831 | 4.7433 | 2.3392 | 2.9169 | H13 | 2.0739 | 3.1994 | 2.6140 | 3.3122 | 1.0829 | 3.2347 | 2.1915 | 3.8211 | 3.5879 | 2.3839 | 3.5879 | 4.1264 | | 1.7425 | 3.3399 | 4.2831 | 2.9169 | 2.3392 | 4.7433 | 4.2831 | H14 | 2.0739 | 3.1994 | 3.3122 | 2.6140 | 1.0829 | 3.8211 | 2.1915 | 3.2347 | 4.1264 | 3.5879 | 2.3839 | 3.5879 | 1.7425 | | 4.2831 | 4.7433 | 2.3392 | 2.9169 | 4.2831 | 3.3399 | H15 | 3.1994 | 2.0739 | 2.1915 | 3.8211 | 3.2347 | 1.0829 | 2.6140 | 3.3122 | 2.9169 | 2.3392 | 4.7433 | 4.2831 | 3.3399 | 4.2831 | | 1.7425 | 3.5879 | 2.3839 | 3.5879 | 4.1264 | H16 | 3.1994 | 2.0739 | 2.1915 | 3.2347 | 3.8211 | 1.0829 | 3.3122 | 2.6140 | 2.3392 | 2.9169 | 4.2831 | 3.3399 | 4.2831 | 4.7433 | 1.7425 | | 4.1264 | 3.5879 | 2.3839 | 3.5879 | H17 | 3.1994 | 2.0739 | 3.8211 | 3.2347 | 2.1915 | 3.3122 | 1.0829 | 2.6140 | 4.7433 | 4.2831 | 3.3399 | 4.2831 | 2.9169 | 2.3392 | 3.5879 | 4.1264 | | 1.7425 | 3.5879 | 2.3839 | H18 | 3.1994 | 2.0739 | 3.2347 | 3.8211 | 2.1915 | 2.6140 | 1.0829 | 3.3122 | 4.2831 | 3.3399 | 4.2831 | 4.7433 | 2.3392 | 2.9169 | 2.3839 | 3.5879 | 1.7425 | | 4.1264 | 3.5879 | H19 | 3.1994 | 2.0739 | 3.2347 | 2.1915 | 3.8211 | 2.6140 | 3.3122 | 1.0829 | 3.3399 | 4.2831 | 2.9169 | 2.3392 | 4.7433 | 4.2831 | 3.5879 | 2.3839 | 3.5879 | 4.1264 | | 1.7425 | H20 | 3.1994 | 2.0739 | 3.8211 | 2.1915 | 3.2347 | 3.3122 | 2.6140 | 1.0829 | 4.2831 | 4.7433 | 2.3392 | 2.9169 | 4.2831 | 3.3399 | 4.1264 | 3.5879 | 2.3839 | 3.5879 | 1.7425 | |
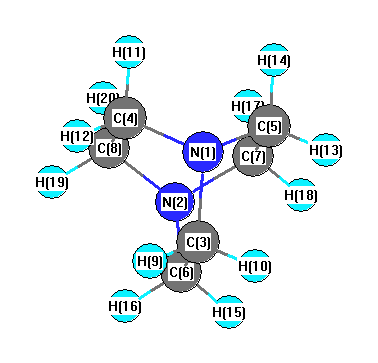 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
109.767 |
|
N1 |
C3 |
H9 |
108.597 |
| N1 |
C3 |
H10 |
108.597 |
|
N1 |
C4 |
C8 |
109.767 |
| N1 |
C4 |
H11 |
108.597 |
|
N1 |
C4 |
H12 |
108.597 |
| N1 |
C5 |
C7 |
109.767 |
|
N1 |
C5 |
H13 |
108.597 |
| N1 |
C5 |
H14 |
108.597 |
|
N2 |
C6 |
C3 |
109.767 |
| N2 |
C6 |
H15 |
108.597 |
|
N2 |
C6 |
H16 |
108.597 |
| N2 |
C7 |
C5 |
109.767 |
|
N2 |
C7 |
H17 |
108.597 |
| N2 |
C7 |
H18 |
108.597 |
|
N2 |
C8 |
C4 |
109.767 |
| N2 |
C8 |
H19 |
108.597 |
|
N2 |
C8 |
H20 |
108.597 |
| C3 |
N1 |
C4 |
109.173 |
|
C3 |
N1 |
C5 |
109.173 |
| C3 |
C6 |
H15 |
111.318 |
|
C3 |
C6 |
H16 |
111.318 |
| C4 |
N1 |
C5 |
109.173 |
|
C4 |
C8 |
H19 |
111.318 |
| C4 |
C8 |
H20 |
111.318 |
|
C5 |
C6 |
H15 |
101.684 |
| C5 |
C6 |
H16 |
151.167 |
|
C6 |
N2 |
C7 |
109.173 |
| C6 |
N2 |
C8 |
109.173 |
|
C6 |
C3 |
H9 |
111.318 |
| C6 |
C3 |
H10 |
111.318 |
|
C7 |
N2 |
C8 |
109.173 |
| C7 |
C5 |
H13 |
111.318 |
|
C7 |
C5 |
H14 |
111.318 |
| C8 |
C4 |
H11 |
111.318 |
|
C8 |
C4 |
H12 |
111.318 |
| H9 |
C3 |
H10 |
107.142 |
|
H11 |
C4 |
H12 |
107.142 |
| H13 |
C5 |
H14 |
107.142 |
|
H15 |
C6 |
H16 |
107.142 |
| H17 |
C7 |
H18 |
107.142 |
|
H19 |
C8 |
H20 |
107.142 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.158 |
|
|
|
| 2 |
N |
-0.158 |
|
|
|
| 3 |
C |
-0.054 |
|
|
|
| 4 |
C |
-0.054 |
|
|
|
| 5 |
C |
-0.054 |
|
|
|
| 6 |
C |
-0.054 |
|
|
|
| 7 |
C |
-0.054 |
|
|
|
| 8 |
C |
-0.054 |
|
|
|
| 9 |
H |
0.053 |
|
|
|
| 10 |
H |
0.053 |
|
|
|
| 11 |
H |
0.053 |
|
|
|
| 12 |
H |
0.053 |
|
|
|
| 13 |
H |
0.053 |
|
|
|
| 14 |
H |
0.053 |
|
|
|
| 15 |
H |
0.053 |
|
|
|
| 16 |
H |
0.053 |
|
|
|
| 17 |
H |
0.053 |
|
|
|
| 18 |
H |
0.053 |
|
|
|
| 19 |
H |
0.053 |
|
|
|
| 20 |
H |
0.053 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.032 |
0.000 |
0.000 |
| y |
0.000 |
-47.032 |
0.000 |
| z |
0.000 |
0.000 |
-58.287 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.627 |
0.000 |
0.000 |
| y |
0.000 |
5.627 |
0.000 |
| z |
0.000 |
0.000 |
-11.255 |
|
| Polar |
|---|
| 3z2-r2 | -22.510 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.289 |
0.000 |
0.000 |
| y |
0.000 |
11.289 |
0.000 |
| z |
0.000 |
0.000 |
10.371 |
<r2> (average value of r
2) Å
2
| <r2> |
211.529 |
| (<r2>)1/2 |
14.544 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -343.176236 |
| Energy at 298.15K | -343.193865 |
| HF Energy | -343.176236 |
| Nuclear repulsion energy | 428.072539 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3194 |
2895 |
0.00 |
|
|
|
| 2 |
A1 |
3193 |
2894 |
0.00 |
|
|
|
| 3 |
A1 |
1648 |
1494 |
0.00 |
|
|
|
| 4 |
A1 |
1498 |
1358 |
0.00 |
|
|
|
| 5 |
A1 |
1372 |
1243 |
0.00 |
|
|
|
| 6 |
A1 |
1120 |
1015 |
0.00 |
|
|
|
| 7 |
A1 |
1051 |
953 |
0.00 |
|
|
|
| 8 |
A1 |
874 |
792 |
0.00 |
|
|
|
| 9 |
A1 |
653 |
592 |
0.00 |
|
|
|
| 10 |
A1 |
68 |
61 |
0.00 |
|
|
|
| 11 |
A2 |
3219 |
2918 |
0.00 |
|
|
|
| 12 |
A2 |
3174 |
2877 |
127.52 |
|
|
|
| 13 |
A2 |
1634 |
1481 |
6.11 |
|
|
|
| 14 |
A2 |
1517 |
1375 |
6.85 |
|
|
|
| 15 |
A2 |
1288 |
1167 |
0.00 |
|
|
|
| 16 |
A2 |
1076 |
975 |
34.07 |
|
|
|
| 17 |
A2 |
871 |
789 |
0.00 |
|
|
|
| 18 |
A2 |
844 |
765 |
70.79 |
|
|
|
| 19 |
E |
3231 |
2928 |
136.78 |
|
|
|
| 19 |
E |
3231 |
2928 |
136.78 |
|
|
|
| 20 |
E |
3199 |
2900 |
0.00 |
|
|
|
| 20 |
E |
3199 |
2900 |
0.00 |
|
|
|
| 21 |
E |
3181 |
2883 |
131.99 |
|
|
|
| 21 |
E |
3181 |
2883 |
131.99 |
|
|
|
| 22 |
E |
3167 |
2870 |
0.00 |
|
|
|
| 22 |
E |
3167 |
2870 |
0.00 |
|
|
|
| 23 |
E |
1630 |
1477 |
4.95 |
|
|
|
| 23 |
E |
1630 |
1477 |
4.95 |
|
|
|
| 24 |
E |
1617 |
1465 |
0.00 |
|
|
|
| 24 |
E |
1617 |
1465 |
0.00 |
|
|
|
| 25 |
E |
1476 |
1338 |
19.01 |
|
|
|
| 25 |
E |
1476 |
1338 |
19.01 |
|
|
|
| 26 |
E |
1476 |
1337 |
0.00 |
|
|
|
| 26 |
E |
1476 |
1337 |
0.00 |
|
|
|
| 27 |
E |
1448 |
1312 |
1.17 |
|
|
|
| 27 |
E |
1448 |
1312 |
1.17 |
|
|
|
| 28 |
E |
1438 |
1303 |
0.00 |
|
|
|
| 28 |
E |
1438 |
1303 |
0.00 |
|
|
|
| 29 |
E |
1326 |
1202 |
0.00 |
|
|
|
| 29 |
E |
1326 |
1202 |
0.00 |
|
|
|
| 30 |
E |
1187 |
1076 |
54.46 |
|
|
|
| 30 |
E |
1187 |
1076 |
54.46 |
|
|
|
| 31 |
E |
1135 |
1028 |
0.00 |
|
|
|
| 31 |
E |
1135 |
1028 |
0.00 |
|
|
|
| 32 |
E |
964 |
874 |
6.67 |
|
|
|
| 32 |
E |
964 |
874 |
6.67 |
|
|
|
| 33 |
E |
898 |
814 |
5.29 |
|
|
|
| 33 |
E |
898 |
814 |
5.29 |
|
|
|
| 34 |
E |
639 |
579 |
0.00 |
|
|
|
| 34 |
E |
639 |
579 |
0.00 |
|
|
|
| 35 |
E |
460 |
417 |
0.07 |
|
|
|
| 35 |
E |
460 |
417 |
0.07 |
|
|
|
| 36 |
E |
356 |
323 |
0.00 |
|
|
|
| 36 |
E |
356 |
323 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42972.2 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 38950.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is D3
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.269 |
| N2 |
0.000 |
0.000 |
-1.269 |
| C3 |
-0.000 |
1.373 |
0.774 |
| C4 |
1.189 |
-0.686 |
0.774 |
| C5 |
-1.188 |
-0.687 |
0.774 |
| C6 |
0.000 |
1.373 |
-0.774 |
| C7 |
-1.189 |
-0.686 |
-0.774 |
| C8 |
1.188 |
-0.687 |
-0.774 |
| H9 |
0.870 |
1.881 |
1.167 |
| H10 |
-0.872 |
1.880 |
1.166 |
| H11 |
1.195 |
-1.694 |
1.167 |
| H12 |
2.064 |
-0.185 |
1.166 |
| H13 |
-2.064 |
-0.188 |
1.167 |
| H14 |
-1.193 |
-1.695 |
1.166 |
| H15 |
-0.870 |
1.881 |
-1.167 |
| H16 |
0.872 |
1.880 |
-1.166 |
| H17 |
-1.195 |
-1.694 |
-1.167 |
| H18 |
-2.064 |
-0.185 |
-1.166 |
| H19 |
2.064 |
-0.188 |
-1.167 |
| H20 |
1.193 |
-1.695 |
-1.166 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5390 | 1.4593 | 1.4593 | 1.4593 | 2.4614 | 2.4614 | 2.4614 | 2.0753 | 2.0752 | 2.0753 | 2.0752 | 2.0753 | 2.0752 | 3.1991 | 3.1982 | 3.1991 | 3.1982 | 3.1991 | 3.1982 |
N2 | 2.5390 | | 2.4614 | 2.4614 | 2.4614 | 1.4593 | 1.4593 | 1.4593 | 3.1991 | 3.1982 | 3.1991 | 3.1982 | 3.1991 | 3.1982 | 2.0753 | 2.0752 | 2.0753 | 2.0752 | 2.0753 | 2.0752 | C3 | 1.4593 | 2.4614 | | 2.3773 | 2.3773 | 1.5473 | 2.8361 | 2.8369 | 1.0822 | 1.0822 | 3.3145 | 2.6161 | 2.6168 | 3.3145 | 2.1868 | 2.1868 | 3.8205 | 3.2325 | 3.2349 | 3.8206 | C4 | 1.4593 | 2.4614 | 2.3773 | | 2.3773 | 2.8361 | 2.8369 | 1.5473 | 2.6168 | 3.3145 | 1.0822 | 1.0822 | 3.3145 | 2.6161 | 3.8205 | 3.2325 | 3.2349 | 3.8206 | 2.1868 | 2.1868 | C5 | 1.4593 | 2.4614 | 2.3773 | 2.3773 | | 2.8369 | 1.5473 | 2.8361 | 3.3145 | 2.6161 | 2.6168 | 3.3145 | 1.0822 | 1.0822 | 3.2349 | 3.8206 | 2.1868 | 2.1868 | 3.8205 | 3.2325 | C6 | 2.4614 | 1.4593 | 1.5473 | 2.8361 | 2.8369 | | 2.3773 | 2.3773 | 2.1868 | 2.1868 | 3.8205 | 3.2325 | 3.2349 | 3.8206 | 1.0822 | 1.0822 | 3.3145 | 2.6161 | 2.6168 | 3.3145 | C7 | 2.4614 | 1.4593 | 2.8361 | 2.8369 | 1.5473 | 2.3773 | | 2.3773 | 3.8205 | 3.2325 | 3.2349 | 3.8206 | 2.1868 | 2.1868 | 2.6168 | 3.3145 | 1.0822 | 1.0822 | 3.3145 | 2.6161 | C8 | 2.4614 | 1.4593 | 2.8369 | 1.5473 | 2.8361 | 2.3773 | 2.3773 | | 3.2349 | 3.8206 | 2.1868 | 2.1868 | 3.8205 | 3.2325 | 3.3145 | 2.6161 | 2.6168 | 3.3145 | 1.0822 | 1.0822 | H9 | 2.0753 | 3.1991 | 1.0822 | 2.6168 | 3.3145 | 2.1868 | 3.8205 | 3.2349 | | 1.7415 | 3.5901 | 2.3870 | 3.5901 | 4.1286 | 2.9115 | 2.3336 | 4.7429 | 4.2808 | 3.3404 | 4.2828 | H10 | 2.0752 | 3.1982 | 1.0822 | 3.3145 | 2.6161 | 2.1868 | 3.2325 | 3.8206 | 1.7415 | | 4.1286 | 3.5899 | 2.3870 | 3.5899 | 2.3336 | 2.9122 | 4.2808 | 3.3360 | 4.2828 | 4.7420 | H11 | 2.0753 | 3.1991 | 3.3145 | 1.0822 | 2.6168 | 3.8205 | 3.2349 | 2.1868 | 3.5901 | 4.1286 | | 1.7415 | 3.5901 | 2.3870 | 4.7429 | 4.2808 | 3.3404 | 4.2828 | 2.9115 | 2.3336 | H12 | 2.0752 | 3.1982 | 2.6161 | 1.0822 | 3.3145 | 3.2325 | 3.8206 | 2.1868 | 2.3870 | 3.5899 | 1.7415 | | 4.1286 | 3.5899 | 4.2808 | 3.3360 | 4.2828 | 4.7420 | 2.3336 | 2.9122 | H13 | 2.0753 | 3.1991 | 2.6168 | 3.3145 | 1.0822 | 3.2349 | 2.1868 | 3.8205 | 3.5901 | 2.3870 | 3.5901 | 4.1286 | | 1.7415 | 3.3404 | 4.2828 | 2.9115 | 2.3336 | 4.7429 | 4.2808 | H14 | 2.0752 | 3.1982 | 3.3145 | 2.6161 | 1.0822 | 3.8206 | 2.1868 | 3.2325 | 4.1286 | 3.5899 | 2.3870 | 3.5899 | 1.7415 | | 4.2828 | 4.7420 | 2.3336 | 2.9122 | 4.2808 | 3.3360 | H15 | 3.1991 | 2.0753 | 2.1868 | 3.8205 | 3.2349 | 1.0822 | 2.6168 | 3.3145 | 2.9115 | 2.3336 | 4.7429 | 4.2808 | 3.3404 | 4.2828 | | 1.7415 | 3.5901 | 2.3870 | 3.5901 | 4.1286 | H16 | 3.1982 | 2.0752 | 2.1868 | 3.2325 | 3.8206 | 1.0822 | 3.3145 | 2.6161 | 2.3336 | 2.9122 | 4.2808 | 3.3360 | 4.2828 | 4.7420 | 1.7415 | | 4.1286 | 3.5899 | 2.3870 | 3.5899 | H17 | 3.1991 | 2.0753 | 3.8205 | 3.2349 | 2.1868 | 3.3145 | 1.0822 | 2.6168 | 4.7429 | 4.2808 | 3.3404 | 4.2828 | 2.9115 | 2.3336 | 3.5901 | 4.1286 | | 1.7415 | 3.5901 | 2.3870 | H18 | 3.1982 | 2.0752 | 3.2325 | 3.8206 | 2.1868 | 2.6161 | 1.0822 | 3.3145 | 4.2808 | 3.3360 | 4.2828 | 4.7420 | 2.3336 | 2.9122 | 2.3870 | 3.5899 | 1.7415 | | 4.1286 | 3.5899 | H19 | 3.1991 | 2.0753 | 3.2349 | 2.1868 | 3.8205 | 2.6168 | 3.3145 | 1.0822 | 3.3404 | 4.2828 | 2.9115 | 2.3336 | 4.7429 | 4.2808 | 3.5901 | 2.3870 | 3.5901 | 4.1286 | | 1.7415 | H20 | 3.1982 | 2.0752 | 3.8206 | 2.1868 | 3.2325 | 3.3145 | 2.6161 | 1.0822 | 4.2828 | 4.7420 | 2.3336 | 2.9122 | 4.2808 | 3.3360 | 4.1286 | 3.5899 | 2.3870 | 3.5899 | 1.7415 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
109.863 |
|
N1 |
C3 |
H9 |
108.567 |
| N1 |
C3 |
H10 |
108.561 |
|
N1 |
C4 |
C8 |
109.863 |
| N1 |
C4 |
H11 |
108.567 |
|
N1 |
C4 |
H12 |
108.561 |
| N1 |
C5 |
C7 |
109.863 |
|
N1 |
C5 |
H13 |
108.567 |
| N1 |
C5 |
H14 |
108.561 |
|
N2 |
C6 |
C3 |
109.863 |
| N2 |
C6 |
H15 |
108.567 |
|
N2 |
C6 |
H16 |
108.561 |
| N2 |
C7 |
C5 |
109.863 |
|
N2 |
C7 |
H17 |
108.567 |
| N2 |
C7 |
H18 |
108.561 |
|
N2 |
C8 |
C4 |
109.863 |
| N2 |
C8 |
H19 |
108.567 |
|
N2 |
C8 |
H20 |
108.561 |
| C3 |
N1 |
C4 |
109.077 |
|
C3 |
N1 |
C5 |
109.077 |
| C3 |
C6 |
H15 |
111.302 |
|
C3 |
C6 |
H16 |
111.300 |
| C4 |
N1 |
C5 |
109.077 |
|
C4 |
C8 |
H19 |
111.302 |
| C4 |
C8 |
H20 |
111.300 |
|
C5 |
C6 |
H15 |
101.700 |
| C5 |
C6 |
H16 |
151.148 |
|
C6 |
N2 |
C7 |
109.077 |
| C6 |
N2 |
C8 |
109.077 |
|
C6 |
C3 |
H9 |
111.302 |
| C6 |
C3 |
H10 |
111.300 |
|
C7 |
N2 |
C8 |
109.077 |
| C7 |
C5 |
H13 |
111.302 |
|
C7 |
C5 |
H14 |
111.300 |
| C8 |
C4 |
H11 |
111.302 |
|
C8 |
C4 |
H12 |
111.300 |
| H9 |
C3 |
H10 |
107.143 |
|
H11 |
C4 |
H12 |
107.143 |
| H13 |
C5 |
H14 |
107.143 |
|
H15 |
C6 |
H16 |
107.143 |
| H17 |
C7 |
H18 |
107.143 |
|
H19 |
C8 |
H20 |
107.143 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.158 |
|
|
|
| 2 |
N |
-0.158 |
|
|
|
| 3 |
C |
-0.054 |
|
|
|
| 4 |
C |
-0.054 |
|
|
|
| 5 |
C |
-0.054 |
|
|
|
| 6 |
C |
-0.054 |
|
|
|
| 7 |
C |
-0.054 |
|
|
|
| 8 |
C |
-0.054 |
|
|
|
| 9 |
H |
0.053 |
|
|
|
| 10 |
H |
0.053 |
|
|
|
| 11 |
H |
0.053 |
|
|
|
| 12 |
H |
0.053 |
|
|
|
| 13 |
H |
0.053 |
|
|
|
| 14 |
H |
0.053 |
|
|
|
| 15 |
H |
0.053 |
|
|
|
| 16 |
H |
0.053 |
|
|
|
| 17 |
H |
0.053 |
|
|
|
| 18 |
H |
0.053 |
|
|
|
| 19 |
H |
0.053 |
|
|
|
| 20 |
H |
0.053 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.029 |
0.000 |
0.000 |
| y |
0.000 |
-47.029 |
0.000 |
| z |
0.000 |
0.000 |
-58.285 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.628 |
0.000 |
0.000 |
| y |
0.000 |
5.628 |
0.000 |
| z |
0.000 |
0.000 |
-11.256 |
|
| Polar |
|---|
| 3z2-r2 | -22.511 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.293 |
0.000 |
0.000 |
| y |
0.000 |
11.293 |
0.000 |
| z |
0.000 |
0.000 |
10.375 |
<r2> (average value of r
2) Å
2
| <r2> |
211.576 |
| (<r2>)1/2 |
14.546 |
