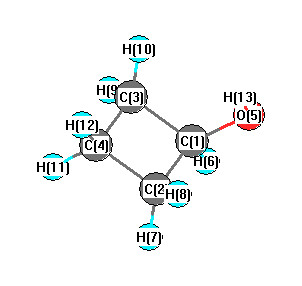
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
4166 |
3776 |
41.08 |
|
|
|
| 2 |
A' |
3251 |
2947 |
111.29 |
|
|
|
| 3 |
A' |
3235 |
2932 |
19.27 |
|
|
|
| 4 |
A' |
3211 |
2911 |
10.37 |
|
|
|
| 5 |
A' |
3194 |
2895 |
32.14 |
|
|
|
| 6 |
A' |
3174 |
2877 |
31.35 |
|
|
|
| 7 |
A' |
1641 |
1487 |
11.15 |
|
|
|
| 8 |
A' |
1608 |
1458 |
3.08 |
|
|
|
| 9 |
A' |
1562 |
1416 |
83.72 |
|
|
|
| 10 |
A' |
1436 |
1301 |
0.52 |
|
|
|
| 11 |
A' |
1391 |
1261 |
17.87 |
|
|
|
| 12 |
A' |
1315 |
1192 |
0.54 |
|
|
|
| 13 |
A' |
1248 |
1131 |
157.27 |
|
|
|
| 14 |
A' |
1178 |
1068 |
27.67 |
|
|
|
| 15 |
A' |
1043 |
945 |
8.69 |
|
|
|
| 16 |
A' |
973 |
882 |
0.99 |
|
|
|
| 17 |
A' |
810 |
735 |
5.12 |
|
|
|
| 18 |
A' |
658 |
596 |
3.63 |
|
|
|
| 19 |
A' |
491 |
445 |
4.44 |
|
|
|
| 20 |
A' |
192 |
174 |
1.67 |
|
|
|
| 21 |
A" |
3230 |
2927 |
53.51 |
|
|
|
| 22 |
A" |
3169 |
2872 |
49.19 |
|
|
|
| 23 |
A" |
1599 |
1450 |
1.85 |
|
|
|
| 24 |
A" |
1424 |
1291 |
0.91 |
|
|
|
| 25 |
A" |
1376 |
1247 |
0.19 |
|
|
|
| 26 |
A" |
1359 |
1232 |
0.39 |
|
|
|
| 27 |
A" |
1297 |
1176 |
0.02 |
|
|
|
| 28 |
A" |
1133 |
1027 |
10.01 |
|
|
|
| 29 |
A" |
1014 |
919 |
5.33 |
|
|
|
| 30 |
A" |
977 |
885 |
0.45 |
|
|
|
| 31 |
A" |
848 |
768 |
0.14 |
|
|
|
| 32 |
A" |
422 |
382 |
29.20 |
|
|
|
| 33 |
A" |
298 |
270 |
95.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26960.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 24437.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.255 |
|
|
|
| 2 |
C |
-0.108 |
|
|
|
| 3 |
C |
-0.108 |
|
|
|
| 4 |
C |
-0.082 |
|
|
|
| 5 |
O |
-0.431 |
|
|
|
| 6 |
H |
0.034 |
|
|
|
| 7 |
H |
0.046 |
|
|
|
| 8 |
H |
0.028 |
|
|
|
| 9 |
H |
0.046 |
|
|
|
| 10 |
H |
0.028 |
|
|
|
| 11 |
H |
0.048 |
|
|
|
| 12 |
H |
0.045 |
|
|
|
| 13 |
H |
0.198 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.252 |
-1.668 |
0.000 |
1.687 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.841 |
1.370 |
0.000 |
| y |
1.370 |
-36.873 |
0.000 |
| z |
0.000 |
0.000 |
-32.346 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.768 |
1.370 |
0.000 |
| y |
1.370 |
-6.779 |
0.000 |
| z |
0.000 |
0.000 |
0.011 |
|
| Polar |
|---|
| 3z2-r2 | 0.022 |
|---|
| x2-y2 | 9.031 |
|---|
| xy | 1.370 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.803 |
-0.400 |
0.000 |
| y |
-0.400 |
7.125 |
0.000 |
| z |
0.000 |
0.000 |
7.074 |
<r2> (average value of r
2) Å
2
| <r2> |
114.452 |
| (<r2>)1/2 |
10.698 |