Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -269.197726 |
| Energy at 298.15K | |
| HF Energy | -269.197726 |
| Nuclear repulsion energy | 258.735345 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3333 |
3021 |
0.53 |
|
|
|
| 2 |
A1 |
3321 |
3010 |
66.11 |
|
|
|
| 3 |
A1 |
3308 |
2999 |
2.51 |
|
|
|
| 4 |
A1 |
3286 |
2979 |
1.33 |
|
|
|
| 5 |
A1 |
1822 |
1651 |
1.46 |
|
|
|
| 6 |
A1 |
1643 |
1489 |
0.66 |
|
|
|
| 7 |
A1 |
1583 |
1435 |
3.61 |
|
|
|
| 8 |
A1 |
1376 |
1247 |
0.40 |
|
|
|
| 9 |
A1 |
1287 |
1167 |
0.34 |
|
|
|
| 10 |
A1 |
998 |
905 |
0.72 |
|
|
|
| 11 |
A1 |
974 |
883 |
1.86 |
|
|
|
| 12 |
A1 |
849 |
770 |
0.12 |
|
|
|
| 13 |
A1 |
440 |
399 |
0.46 |
|
|
|
| 14 |
A2 |
1123 |
1018 |
0.00 |
|
|
|
| 15 |
A2 |
1025 |
929 |
0.00 |
|
|
|
| 16 |
A2 |
785 |
712 |
0.00 |
|
|
|
| 17 |
A2 |
575 |
521 |
0.00 |
|
|
|
| 18 |
A2 |
304 |
275 |
0.00 |
|
|
|
| 19 |
B1 |
1018 |
923 |
0.19 |
|
|
|
| 20 |
B1 |
887 |
804 |
3.31 |
|
|
|
| 21 |
B1 |
812 |
736 |
2.06 |
|
|
|
| 22 |
B1 |
668 |
606 |
104.57 |
|
|
|
| 23 |
B1 |
536 |
486 |
17.34 |
|
|
|
| 24 |
B1 |
149 |
135 |
0.01 |
|
|
|
| 25 |
B2 |
3323 |
3012 |
67.28 |
|
|
|
| 26 |
B2 |
3294 |
2986 |
4.13 |
|
|
|
| 27 |
B2 |
3287 |
2979 |
0.98 |
|
|
|
| 28 |
B2 |
1616 |
1465 |
1.95 |
|
|
|
| 29 |
B2 |
1574 |
1427 |
4.06 |
|
|
|
| 30 |
B2 |
1523 |
1380 |
0.09 |
|
|
|
| 31 |
B2 |
1395 |
1264 |
0.21 |
|
|
|
| 32 |
B2 |
1232 |
1117 |
0.14 |
|
|
|
| 33 |
B2 |
980 |
888 |
0.05 |
|
|
|
| 34 |
B2 |
966 |
876 |
5.03 |
|
|
|
| 35 |
B2 |
469 |
425 |
0.52 |
|
|
|
| 36 |
B2 |
2769i |
2510i |
35.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24495.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 22202.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.619 |
| C2 |
0.000 |
1.267 |
0.999 |
| C3 |
0.000 |
-1.267 |
0.999 |
| C4 |
0.000 |
1.582 |
-0.337 |
| C5 |
0.000 |
-1.582 |
-0.337 |
| C6 |
0.000 |
0.665 |
-1.472 |
| C7 |
0.000 |
-0.665 |
-1.472 |
| H8 |
0.000 |
0.000 |
2.693 |
| H9 |
0.000 |
2.105 |
1.672 |
| H10 |
0.000 |
-2.105 |
1.672 |
| H11 |
0.000 |
2.627 |
-0.583 |
| H12 |
0.000 |
-2.627 |
-0.583 |
| H13 |
0.000 |
1.140 |
-2.436 |
| H14 |
0.000 |
-1.140 |
-2.436 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.4105 | 1.4105 | 2.5163 | 2.5163 | 3.1618 | 3.1618 | 1.0734 | 2.1058 | 2.1058 | 3.4282 | 3.4282 | 4.2122 | 4.2122 |
C2 | 1.4105 | | 2.5334 | 1.3729 | 3.1468 | 2.5429 | 3.1362 | 2.1150 | 1.0753 | 3.4384 | 2.0866 | 4.2029 | 3.4370 | 4.1939 | C3 | 1.4105 | 2.5334 | | 3.1468 | 1.3729 | 3.1362 | 2.5429 | 2.1150 | 3.4384 | 1.0753 | 4.2029 | 2.0866 | 4.1939 | 3.4370 | C4 | 2.5163 | 1.3729 | 3.1468 | | 3.1647 | 1.4591 | 2.5174 | 3.4182 | 2.0763 | 4.1994 | 1.0732 | 4.2164 | 2.1447 | 3.4372 | C5 | 2.5163 | 3.1468 | 1.3729 | 3.1647 | | 2.5174 | 1.4591 | 3.4182 | 4.1994 | 2.0763 | 4.2164 | 1.0732 | 3.4372 | 2.1447 | C6 | 3.1618 | 2.5429 | 3.1362 | 1.4591 | 2.5174 | | 1.3299 | 4.2172 | 3.4581 | 4.1902 | 2.1537 | 3.4096 | 1.0746 | 2.0461 | C7 | 3.1618 | 3.1362 | 2.5429 | 2.5174 | 1.4591 | 1.3299 | | 4.2172 | 4.1902 | 3.4581 | 3.4096 | 2.1537 | 2.0461 | 1.0746 | H8 | 1.0734 | 2.1150 | 2.1150 | 3.4182 | 3.4182 | 4.2172 | 4.2172 | | 2.3394 | 2.3394 | 4.1992 | 4.1992 | 5.2536 | 5.2536 | H9 | 2.1058 | 1.0753 | 3.4384 | 2.0763 | 4.1994 | 3.4581 | 4.1902 | 2.3394 | | 4.2102 | 2.3152 | 5.2421 | 4.2198 | 5.2350 | H10 | 2.1058 | 3.4384 | 1.0753 | 4.1994 | 2.0763 | 4.1902 | 3.4581 | 2.3394 | 4.2102 | | 5.2421 | 2.3152 | 5.2350 | 4.2198 | H11 | 3.4282 | 2.0866 | 4.2029 | 1.0732 | 4.2164 | 2.1537 | 3.4096 | 4.1992 | 2.3152 | 5.2421 | | 5.2538 | 2.3753 | 4.1976 | H12 | 3.4282 | 4.2029 | 2.0866 | 4.2164 | 1.0732 | 3.4096 | 2.1537 | 4.1992 | 5.2421 | 2.3152 | 5.2538 | | 4.1976 | 2.3753 | H13 | 4.2122 | 3.4370 | 4.1939 | 2.1447 | 3.4372 | 1.0746 | 2.0461 | 5.2536 | 4.2198 | 5.2350 | 2.3753 | 4.1976 | | 2.2798 | H14 | 4.2122 | 4.1939 | 3.4370 | 3.4372 | 2.1447 | 2.0461 | 1.0746 | 5.2536 | 5.2350 | 4.2198 | 4.1976 | 2.3753 | 2.2798 | |
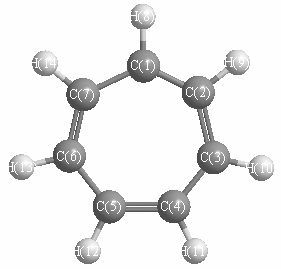 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
26.094 |
|
C1 |
C2 |
H9 |
115.139 |
| C1 |
C7 |
C6 |
77.860 |
|
C1 |
C7 |
H14 |
165.909 |
| C2 |
C1 |
C7 |
76.046 |
|
C2 |
C1 |
H8 |
116.094 |
| C2 |
C3 |
C4 |
25.125 |
|
C2 |
C3 |
H10 |
141.234 |
| C3 |
C2 |
H9 |
141.234 |
|
C3 |
C4 |
C5 |
25.125 |
| C3 |
C4 |
H11 |
168.140 |
|
C4 |
C3 |
H10 |
166.359 |
| C4 |
C5 |
C6 |
26.787 |
|
C4 |
C5 |
H12 |
166.735 |
| C5 |
C4 |
H11 |
166.735 |
|
C5 |
C6 |
C7 |
26.787 |
| C5 |
C6 |
H13 |
143.018 |
|
C6 |
C5 |
H12 |
139.947 |
| C6 |
C7 |
H14 |
116.231 |
|
C7 |
C1 |
H8 |
167.860 |
| C7 |
C6 |
H13 |
116.231 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.066 |
|
|
|
| 2 |
C |
-0.094 |
|
|
|
| 3 |
C |
-0.094 |
|
|
|
| 4 |
C |
-0.064 |
|
|
|
| 5 |
C |
-0.064 |
|
|
|
| 6 |
C |
-0.094 |
|
|
|
| 7 |
C |
-0.094 |
|
|
|
| 8 |
H |
0.083 |
|
|
|
| 9 |
H |
0.080 |
|
|
|
| 10 |
H |
0.080 |
|
|
|
| 11 |
H |
0.084 |
|
|
|
| 12 |
H |
0.084 |
|
|
|
| 13 |
H |
0.080 |
|
|
|
| 14 |
H |
0.080 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.085 |
0.085 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.018 |
0.000 |
0.000 |
| y |
0.000 |
-37.438 |
0.000 |
| z |
0.000 |
0.000 |
-37.653 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.473 |
0.000 |
0.000 |
| y |
0.000 |
4.898 |
0.000 |
| z |
0.000 |
0.000 |
4.575 |
|
| Polar |
|---|
| 3z2-r2 | 9.149 |
|---|
| x2-y2 | -9.580 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.676 |
0.000 |
0.000 |
| y |
0.000 |
13.292 |
0.000 |
| z |
0.000 |
0.000 |
12.754 |
<r2> (average value of r
2) Å
2
| <r2> |
185.872 |
| (<r2>)1/2 |
13.634 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -269.198389 |
| Energy at 298.15K | -269.205449 |
| HF Energy | -269.198389 |
| Nuclear repulsion energy | 258.518276 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3332 |
3020 |
1.92 |
|
|
|
| 2 |
A' |
3322 |
3011 |
66.24 |
|
|
|
| 3 |
A' |
3320 |
3010 |
70.90 |
|
|
|
| 4 |
A' |
3307 |
2998 |
1.72 |
|
|
|
| 5 |
A' |
3296 |
2988 |
3.84 |
|
|
|
| 6 |
A' |
3288 |
2980 |
2.08 |
|
|
|
| 7 |
A' |
3286 |
2978 |
0.57 |
|
|
|
| 8 |
A' |
1721 |
1560 |
0.00 |
|
|
|
| 9 |
A' |
1666 |
1510 |
0.84 |
|
|
|
| 10 |
A' |
1604 |
1454 |
0.91 |
|
|
|
| 11 |
A' |
1589 |
1440 |
2.43 |
|
|
|
| 12 |
A' |
1573 |
1426 |
5.29 |
|
|
|
| 13 |
A' |
1522 |
1379 |
0.11 |
|
|
|
| 14 |
A' |
1419 |
1286 |
1.02 |
|
|
|
| 15 |
A' |
1365 |
1238 |
0.10 |
|
|
|
| 16 |
A' |
1287 |
1167 |
0.22 |
|
|
|
| 17 |
A' |
1220 |
1106 |
0.09 |
|
|
|
| 18 |
A' |
1002 |
908 |
2.78 |
|
|
|
| 19 |
A' |
991 |
898 |
8.42 |
|
|
|
| 20 |
A' |
975 |
884 |
2.17 |
|
|
|
| 21 |
A' |
949 |
860 |
0.02 |
|
|
|
| 22 |
A' |
926 |
840 |
0.91 |
|
|
|
| 23 |
A' |
837 |
758 |
0.40 |
|
|
|
| 24 |
A' |
465 |
421 |
0.48 |
|
|
|
| 25 |
A' |
438 |
397 |
0.22 |
|
|
|
| 26 |
A" |
1070 |
969 |
0.32 |
|
|
|
| 27 |
A" |
1057 |
958 |
0.00 |
|
|
|
| 28 |
A" |
1002 |
908 |
0.07 |
|
|
|
| 29 |
A" |
868 |
787 |
0.00 |
|
|
|
| 30 |
A" |
800 |
725 |
2.60 |
|
|
|
| 31 |
A" |
794 |
719 |
0.00 |
|
|
|
| 32 |
A" |
670 |
608 |
105.56 |
|
|
|
| 33 |
A" |
561 |
509 |
0.00 |
|
|
|
| 34 |
A" |
539 |
488 |
18.08 |
|
|
|
| 35 |
A" |
300 |
272 |
0.05 |
|
|
|
| 36 |
A" |
150 |
136 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26254.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 23797.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.788 |
-1.390 |
0.000 |
| C2 |
-0.813 |
-1.117 |
0.000 |
| C3 |
1.643 |
-0.503 |
0.000 |
| C4 |
-1.452 |
-0.207 |
0.000 |
| C5 |
1.154 |
1.128 |
0.000 |
| C6 |
-1.239 |
0.722 |
0.000 |
| C7 |
0.000 |
1.363 |
0.000 |
| H8 |
1.060 |
-2.435 |
0.000 |
| H9 |
-1.310 |
-2.093 |
0.000 |
| H10 |
2.710 |
-0.624 |
0.000 |
| H11 |
-2.531 |
-0.158 |
0.000 |
| H12 |
2.077 |
1.728 |
0.000 |
| H13 |
-2.179 |
1.141 |
0.000 |
| H14 |
-0.322 |
2.466 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.6240 | 1.2312 | 2.5330 | 2.5435 | 2.9267 | 2.8631 | 1.0800 | 2.2136 | 2.0685 | 3.5406 | 3.3734 | 3.8994 | 4.0117 |
C2 | 1.6240 | | 2.5310 | 1.1121 | 2.9842 | 1.8872 | 2.6095 | 2.2900 | 1.0960 | 3.5569 | 1.9679 | 4.0553 | 2.6389 | 3.6160 | C3 | 1.2312 | 2.5310 | | 3.1085 | 1.7024 | 3.1308 | 2.4861 | 2.0176 | 3.3541 | 1.0739 | 4.1883 | 2.2729 | 4.1600 | 3.5597 | C4 | 2.5330 | 1.1121 | 3.1085 | | 2.9272 | 0.9523 | 2.1378 | 3.3576 | 1.8920 | 4.1823 | 1.0808 | 4.0244 | 1.5311 | 2.9013 | C5 | 2.5435 | 2.9842 | 1.7024 | 2.9272 | | 2.4268 | 1.1777 | 3.5635 | 4.0556 | 2.3427 | 3.9032 | 1.1014 | 3.3325 | 1.9918 | C6 | 2.9267 | 1.8872 | 3.1308 | 0.9523 | 2.4268 | | 1.3948 | 3.9047 | 2.8159 | 4.1714 | 1.5637 | 3.4654 | 1.0292 | 1.9704 | C7 | 2.8631 | 2.6095 | 2.4861 | 2.1378 | 1.1777 | 1.3948 | | 3.9428 | 3.6963 | 3.3601 | 2.9532 | 2.1093 | 2.1898 | 1.1487 | H8 | 1.0800 | 2.2900 | 2.0176 | 3.3576 | 3.5635 | 3.9047 | 3.9428 | | 2.3951 | 2.4496 | 4.2521 | 4.2851 | 4.8244 | 5.0914 | H9 | 2.2136 | 1.0960 | 3.3541 | 1.8920 | 4.0556 | 2.8159 | 3.6963 | 2.3951 | | 4.2804 | 2.2879 | 5.1068 | 3.3488 | 4.6650 | H10 | 2.0685 | 3.5569 | 1.0739 | 4.1823 | 2.3427 | 4.1714 | 3.3601 | 2.4496 | 4.2804 | | 5.2618 | 2.4353 | 5.1972 | 4.3283 | H11 | 3.5406 | 1.9679 | 4.1883 | 1.0808 | 3.9032 | 1.5637 | 2.9532 | 4.2521 | 2.2879 | 5.2618 | | 4.9798 | 1.3463 | 3.4305 | H12 | 3.3734 | 4.0553 | 2.2729 | 4.0244 | 1.1014 | 3.4654 | 2.1093 | 4.2851 | 5.1068 | 2.4353 | 4.9798 | | 4.2963 | 2.5099 | H13 | 3.8994 | 2.6389 | 4.1600 | 1.5311 | 3.3325 | 1.0292 | 2.1898 | 4.8244 | 3.3488 | 5.1972 | 1.3463 | 4.2963 | | 2.2811 | H14 | 4.0117 | 3.6160 | 3.5597 | 2.9013 | 1.9918 | 1.9704 | 1.1487 | 5.0914 | 4.6650 | 4.3283 | 3.4305 | 2.5099 | 2.2811 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
23.699 |
|
C1 |
C2 |
H9 |
107.348 |
| C1 |
C7 |
C6 |
78.610 |
|
C1 |
C7 |
H14 |
179.729 |
| C2 |
C1 |
C7 |
64.354 |
|
C2 |
C1 |
H8 |
114.243 |
| C2 |
C3 |
C4 |
19.509 |
|
C2 |
C3 |
H10 |
159.509 |
| C3 |
C2 |
H9 |
131.047 |
|
C3 |
C4 |
C5 |
32.589 |
| C3 |
C4 |
H11 |
177.085 |
|
C4 |
C3 |
H10 |
179.018 |
| C4 |
C5 |
C6 |
17.485 |
|
C4 |
C5 |
H12 |
174.090 |
| C5 |
C4 |
H11 |
150.327 |
|
C5 |
C6 |
C7 |
17.742 |
| C5 |
C6 |
H13 |
146.325 |
|
C6 |
C5 |
H12 |
156.605 |
| C6 |
C7 |
H14 |
101.118 |
|
C7 |
C1 |
H8 |
178.597 |
| C7 |
C6 |
H13 |
128.583 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.100 |
|
|
|
| 2 |
C |
-0.064 |
|
|
|
| 3 |
C |
-0.076 |
|
|
|
| 4 |
C |
-0.091 |
|
|
|
| 5 |
C |
-0.076 |
|
|
|
| 6 |
C |
-0.064 |
|
|
|
| 7 |
C |
-0.100 |
|
|
|
| 8 |
H |
0.079 |
|
|
|
| 9 |
H |
0.083 |
|
|
|
| 10 |
H |
0.084 |
|
|
|
| 11 |
H |
0.080 |
|
|
|
| 12 |
H |
0.084 |
|
|
|
| 13 |
H |
0.083 |
|
|
|
| 14 |
H |
0.079 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
-0.042 |
0.000 |
0.042 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.674 |
0.000 |
0.000 |
| y |
0.000 |
-37.465 |
0.000 |
| z |
0.000 |
0.000 |
-46.980 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.548 |
0.000 |
0.000 |
| y |
0.000 |
4.862 |
0.000 |
| z |
0.000 |
0.000 |
-9.411 |
|
| Polar |
|---|
| 3z2-r2 | -18.821 |
|---|
| x2-y2 | -0.209 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.750 |
0.001 |
0.000 |
| y |
0.001 |
13.158 |
0.000 |
| z |
0.000 |
0.000 |
6.664 |
<r2> (average value of r
2) Å
2
| <r2> |
186.141 |
| (<r2>)1/2 |
13.643 |
