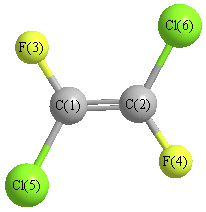
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1956 |
1773 |
0.00 |
|
|
|
| 2 |
Ag |
1332 |
1207 |
0.00 |
|
|
|
| 3 |
Ag |
701 |
635 |
0.00 |
|
|
|
| 4 |
Ag |
466 |
423 |
0.00 |
|
|
|
| 5 |
Ag |
315 |
285 |
0.00 |
|
|
|
| 6 |
Au |
423 |
383 |
1.65 |
|
|
|
| 7 |
Au |
150 |
136 |
0.22 |
|
|
|
| 8 |
Bg |
672 |
609 |
0.00 |
|
|
|
| 9 |
Bu |
1353 |
1226 |
349.31 |
|
|
|
| 10 |
Bu |
956 |
866 |
196.54 |
|
|
|
| 11 |
Bu |
464 |
421 |
3.41 |
|
|
|
| 12 |
Bu |
194 |
176 |
2.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4490.3 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 4070.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.236 |
|
|
|
| 2 |
C |
0.236 |
|
|
|
| 3 |
F |
-0.218 |
|
|
|
| 4 |
F |
-0.218 |
|
|
|
| 5 |
Cl |
-0.017 |
|
|
|
| 6 |
Cl |
-0.017 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.396 |
2.599 |
0.000 |
| y |
2.599 |
-45.765 |
0.000 |
| z |
0.000 |
0.000 |
-45.227 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.900 |
2.599 |
0.000 |
| y |
2.599 |
0.046 |
0.000 |
| z |
0.000 |
0.000 |
0.854 |
|
| Polar |
|---|
| 3z2-r2 | 1.707 |
|---|
| x2-y2 | -0.631 |
|---|
| xy | 2.599 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.028 |
1.961 |
0.000 |
| y |
1.961 |
9.252 |
0.000 |
| z |
0.000 |
0.000 |
4.414 |
<r2> (average value of r
2) Å
2
| <r2> |
243.265 |
| (<r2>)1/2 |
15.597 |