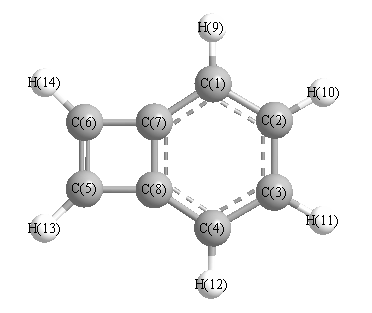
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3397 |
3079 |
8.29 |
|
|
|
| 2 |
A1 |
3347 |
3034 |
16.56 |
|
|
|
| 3 |
A1 |
3329 |
3018 |
13.21 |
|
|
|
| 4 |
A1 |
1840 |
1668 |
3.66 |
|
|
|
| 5 |
A1 |
1711 |
1551 |
2.92 |
|
|
|
| 6 |
A1 |
1595 |
1446 |
11.64 |
|
|
|
| 7 |
A1 |
1545 |
1401 |
10.83 |
|
|
|
| 8 |
A1 |
1274 |
1155 |
0.00 |
|
|
|
| 9 |
A1 |
1173 |
1063 |
1.25 |
|
|
|
| 10 |
A1 |
1158 |
1050 |
0.11 |
|
|
|
| 11 |
A1 |
992 |
899 |
6.56 |
|
|
|
| 12 |
A1 |
873 |
792 |
0.45 |
|
|
|
| 13 |
A1 |
584 |
529 |
0.54 |
|
|
|
| 14 |
A2 |
1088 |
987 |
0.00 |
|
|
|
| 15 |
A2 |
1058 |
959 |
0.00 |
|
|
|
| 16 |
A2 |
988 |
895 |
0.00 |
|
|
|
| 17 |
A2 |
851 |
772 |
0.00 |
|
|
|
| 18 |
A2 |
636 |
577 |
0.00 |
|
|
|
| 19 |
A2 |
326 |
296 |
0.00 |
|
|
|
| 20 |
B1 |
1026 |
930 |
13.28 |
|
|
|
| 21 |
B1 |
846 |
766 |
122.83 |
|
|
|
| 22 |
B1 |
811 |
735 |
9.00 |
|
|
|
| 23 |
B1 |
393 |
356 |
3.95 |
|
|
|
| 24 |
B1 |
263 |
239 |
15.72 |
|
|
|
| 25 |
B2 |
3366 |
3051 |
1.08 |
|
|
|
| 26 |
B2 |
3339 |
3026 |
34.56 |
|
|
|
| 27 |
B2 |
3315 |
3004 |
0.25 |
|
|
|
| 28 |
B2 |
1809 |
1640 |
3.07 |
|
|
|
| 29 |
B2 |
1572 |
1425 |
6.40 |
|
|
|
| 30 |
B2 |
1421 |
1288 |
5.52 |
|
|
|
| 31 |
B2 |
1374 |
1245 |
9.00 |
|
|
|
| 32 |
B2 |
1163 |
1054 |
3.30 |
|
|
|
| 33 |
B2 |
1130 |
1024 |
0.01 |
|
|
|
| 34 |
B2 |
955 |
865 |
1.04 |
|
|
|
| 35 |
B2 |
704 |
638 |
0.48 |
|
|
|
| 36 |
B2 |
455 |
412 |
4.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25852.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 23432.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.077 |
|
|
|
| 2 |
C |
-0.081 |
|
|
|
| 3 |
C |
-0.081 |
|
|
|
| 4 |
C |
-0.077 |
|
|
|
| 5 |
C |
-0.146 |
|
|
|
| 6 |
C |
-0.146 |
|
|
|
| 7 |
C |
-0.017 |
|
|
|
| 8 |
C |
-0.017 |
|
|
|
| 9 |
H |
0.090 |
|
|
|
| 10 |
H |
0.089 |
|
|
|
| 11 |
H |
0.089 |
|
|
|
| 12 |
H |
0.090 |
|
|
|
| 13 |
H |
0.141 |
|
|
|
| 14 |
H |
0.141 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.885 |
0.885 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.834 |
0.000 |
0.000 |
| y |
0.000 |
-41.017 |
0.000 |
| z |
0.000 |
0.000 |
-40.875 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.888 |
0.000 |
0.000 |
| y |
0.000 |
5.338 |
0.000 |
| z |
0.000 |
0.000 |
5.551 |
|
| Polar |
|---|
| 3z2-r2 | 11.101 |
|---|
| x2-y2 | -10.818 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.187 |
0.000 |
0.000 |
| y |
0.000 |
13.840 |
0.000 |
| z |
0.000 |
0.000 |
15.971 |
<r2> (average value of r
2) Å
2
| <r2> |
217.881 |
| (<r2>)1/2 |
14.761 |