Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -234.266781 |
| Energy at 298.15K | |
| HF Energy | -234.266781 |
| Nuclear repulsion energy | 226.512967 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3267 |
2961 |
51.39 |
|
|
|
| 2 |
A' |
3258 |
2953 |
13.66 |
|
|
|
| 3 |
A' |
3224 |
2922 |
15.84 |
|
|
|
| 4 |
A' |
3213 |
2913 |
60.65 |
|
|
|
| 5 |
A' |
3155 |
2859 |
55.16 |
|
|
|
| 6 |
A' |
3151 |
2856 |
55.51 |
|
|
|
| 7 |
A' |
3150 |
2855 |
28.72 |
|
|
|
| 8 |
A' |
3143 |
2849 |
14.60 |
|
|
|
| 9 |
A' |
1874 |
1699 |
1.03 |
|
|
|
| 10 |
A' |
1627 |
1474 |
4.92 |
|
|
|
| 11 |
A' |
1610 |
1460 |
0.26 |
|
|
|
| 12 |
A' |
1610 |
1459 |
9.38 |
|
|
|
| 13 |
A' |
1604 |
1454 |
0.85 |
|
|
|
| 14 |
A' |
1540 |
1395 |
1.69 |
|
|
|
| 15 |
A' |
1538 |
1394 |
4.07 |
|
|
|
| 16 |
A' |
1523 |
1380 |
0.58 |
|
|
|
| 17 |
A' |
1459 |
1322 |
2.39 |
|
|
|
| 18 |
A' |
1445 |
1310 |
2.40 |
|
|
|
| 19 |
A' |
1379 |
1250 |
11.68 |
|
|
|
| 20 |
A' |
1247 |
1131 |
0.98 |
|
|
|
| 21 |
A' |
1166 |
1056 |
0.84 |
|
|
|
| 22 |
A' |
1142 |
1036 |
5.65 |
|
|
|
| 23 |
A' |
1096 |
994 |
0.35 |
|
|
|
| 24 |
A' |
986 |
894 |
1.99 |
|
|
|
| 25 |
A' |
960 |
870 |
2.89 |
|
|
|
| 26 |
A' |
552 |
501 |
0.15 |
|
|
|
| 27 |
A' |
411 |
373 |
1.08 |
|
|
|
| 28 |
A' |
333 |
302 |
0.50 |
|
|
|
| 29 |
A' |
148 |
134 |
0.03 |
|
|
|
| 30 |
A" |
3211 |
2911 |
101.49 |
|
|
|
| 31 |
A" |
3191 |
2892 |
34.16 |
|
|
|
| 32 |
A" |
3182 |
2884 |
22.70 |
|
|
|
| 33 |
A" |
3157 |
2862 |
0.24 |
|
|
|
| 34 |
A" |
1614 |
1463 |
6.06 |
|
|
|
| 35 |
A" |
1597 |
1447 |
5.50 |
|
|
|
| 36 |
A" |
1435 |
1301 |
0.14 |
|
|
|
| 37 |
A" |
1367 |
1239 |
0.00 |
|
|
|
| 38 |
A" |
1219 |
1105 |
0.27 |
|
|
|
| 39 |
A" |
1174 |
1064 |
4.15 |
|
|
|
| 40 |
A" |
1095 |
993 |
39.81 |
|
|
|
| 41 |
A" |
963 |
873 |
0.06 |
|
|
|
| 42 |
A" |
849 |
770 |
0.33 |
|
|
|
| 43 |
A" |
769 |
697 |
1.46 |
|
|
|
| 44 |
A" |
282 |
255 |
3.05 |
|
|
|
| 45 |
A" |
255 |
231 |
0.20 |
|
|
|
| 46 |
A" |
212 |
193 |
1.19 |
|
|
|
| 47 |
A" |
101 |
91 |
0.08 |
|
|
|
| 48 |
A" |
118i |
107i |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38181.6 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 34607.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.634 |
3.179 |
0.000 |
| C2 |
0.914 |
1.707 |
0.000 |
| C3 |
0.000 |
0.765 |
0.000 |
| C4 |
0.277 |
-0.721 |
0.000 |
| C5 |
-0.993 |
-1.569 |
0.000 |
| C6 |
-0.712 |
-3.067 |
0.000 |
| H7 |
-0.429 |
3.379 |
0.000 |
| H8 |
1.068 |
3.659 |
0.872 |
| H9 |
1.068 |
3.659 |
-0.872 |
| H10 |
1.953 |
1.418 |
0.000 |
| H11 |
-1.035 |
1.063 |
0.000 |
| H12 |
0.876 |
-0.981 |
0.867 |
| H13 |
0.876 |
-0.981 |
-0.867 |
| H14 |
-1.592 |
-1.313 |
-0.869 |
| H15 |
-1.592 |
-1.313 |
0.869 |
| H16 |
-1.631 |
-3.641 |
0.000 |
| H17 |
-0.142 |
-3.358 |
-0.875 |
| H18 |
-0.142 |
-3.358 |
0.875 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.4987 | 2.4958 | 3.9164 | 5.0195 | 6.3897 | 1.0821 | 1.0851 | 1.0851 | 2.2002 | 2.6955 | 4.2564 | 4.2564 | 5.0888 | 5.0888 | 7.1867 | 6.6409 | 6.6409 |
C2 | 1.4987 | | 1.3123 | 2.5099 | 3.7906 | 5.0433 | 2.1450 | 2.1431 | 2.1431 | 1.0780 | 2.0529 | 2.8245 | 2.8245 | 4.0197 | 4.0197 | 5.9226 | 5.2472 | 5.2472 | C3 | 2.4958 | 1.3123 | | 1.5117 | 2.5369 | 3.8979 | 2.6488 | 3.2048 | 3.2048 | 2.0586 | 1.0771 | 2.1375 | 2.1375 | 2.7588 | 2.7588 | 4.6986 | 4.2175 | 4.2175 | C4 | 3.9164 | 2.5099 | 1.5117 | | 1.5270 | 2.5462 | 4.1603 | 4.5348 | 4.5348 | 2.7166 | 2.2144 | 1.0857 | 1.0857 | 2.1447 | 2.1447 | 3.4883 | 2.8099 | 2.8099 | C5 | 5.0195 | 3.7906 | 2.5369 | 1.5270 | | 1.5236 | 4.9805 | 5.6865 | 5.6865 | 4.1947 | 2.6327 | 2.1425 | 2.1425 | 1.0861 | 1.0861 | 2.1679 | 2.1653 | 2.1653 | C6 | 6.3897 | 5.0433 | 3.8979 | 2.5462 | 1.5236 | | 6.4523 | 7.0115 | 7.0115 | 5.2165 | 4.1425 | 2.7616 | 2.7616 | 2.1457 | 2.1457 | 1.0832 | 1.0844 | 1.0844 | H7 | 1.0821 | 2.1450 | 2.6488 | 4.1603 | 4.9805 | 6.4523 | | 1.7544 | 1.7544 | 3.0855 | 2.3941 | 4.6331 | 4.6331 | 4.9119 | 4.9119 | 7.1224 | 6.7997 | 6.7997 | H8 | 1.0851 | 2.1431 | 3.2048 | 4.5348 | 5.6865 | 7.0115 | 1.7544 | | 1.7434 | 2.5623 | 3.4524 | 4.6435 | 4.9585 | 5.9012 | 5.6388 | 7.8312 | 7.3312 | 7.1200 | H9 | 1.0851 | 2.1431 | 3.2048 | 4.5348 | 5.6865 | 7.0115 | 1.7544 | 1.7434 | | 2.5623 | 3.4524 | 4.9585 | 4.6435 | 5.6388 | 5.9012 | 7.8312 | 7.1200 | 7.3312 | H10 | 2.2002 | 1.0780 | 2.0586 | 2.7166 | 4.1947 | 5.2165 | 3.0855 | 2.5623 | 2.5623 | | 3.0087 | 2.7683 | 2.7683 | 4.5583 | 4.5583 | 6.1993 | 5.2875 | 5.2875 | H11 | 2.6955 | 2.0529 | 1.0771 | 2.2144 | 2.6327 | 4.1425 | 2.3941 | 3.4524 | 3.4524 | 3.0087 | | 2.9296 | 2.9296 | 2.5907 | 2.5907 | 4.7417 | 4.5944 | 4.5944 | H12 | 4.2564 | 2.8245 | 2.1375 | 1.0857 | 2.1425 | 2.7616 | 4.6331 | 4.6435 | 4.9585 | 2.7683 | 2.9296 | | 1.7346 | 3.0359 | 2.4907 | 3.7568 | 3.1181 | 2.5858 | H13 | 4.2564 | 2.8245 | 2.1375 | 1.0857 | 2.1425 | 2.7616 | 4.6331 | 4.9585 | 4.6435 | 2.7683 | 2.9296 | 1.7346 | | 2.4907 | 3.0359 | 3.7568 | 2.5858 | 3.1181 | H14 | 5.0888 | 4.0197 | 2.7588 | 2.1447 | 1.0861 | 2.1457 | 4.9119 | 5.9012 | 5.6388 | 4.5583 | 2.5907 | 3.0359 | 2.4907 | | 1.7372 | 2.4848 | 2.5068 | 3.0536 | H15 | 5.0888 | 4.0197 | 2.7588 | 2.1447 | 1.0861 | 2.1457 | 4.9119 | 5.6388 | 5.9012 | 4.5583 | 2.5907 | 2.4907 | 3.0359 | 1.7372 | | 2.4848 | 3.0536 | 2.5068 | H16 | 7.1867 | 5.9226 | 4.6986 | 3.4883 | 2.1679 | 1.0832 | 7.1224 | 7.8312 | 7.8312 | 6.1993 | 4.7417 | 3.7568 | 3.7568 | 2.4848 | 2.4848 | | 1.7502 | 1.7502 | H17 | 6.6409 | 5.2472 | 4.2175 | 2.8099 | 2.1653 | 1.0844 | 6.7997 | 7.3312 | 7.1200 | 5.2875 | 4.5944 | 3.1181 | 2.5858 | 2.5068 | 3.0536 | 1.7502 | | 1.7503 | H18 | 6.6409 | 5.2472 | 4.2175 | 2.8099 | 2.1653 | 1.0844 | 6.7997 | 7.1200 | 7.3312 | 5.2875 | 4.5944 | 2.5858 | 3.1181 | 3.0536 | 2.5068 | 1.7502 | 1.7503 | |
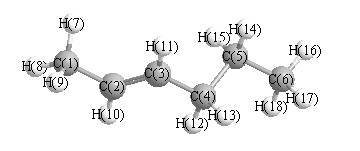 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.088 |
|
C1 |
C2 |
H10 |
116.326 |
| C2 |
C1 |
H7 |
111.406 |
|
C2 |
C1 |
H8 |
111.073 |
| C2 |
C1 |
H9 |
111.073 |
|
C2 |
C3 |
C4 |
125.288 |
| C2 |
C3 |
H11 |
118.111 |
|
C3 |
C2 |
H10 |
118.587 |
| C3 |
C4 |
C5 |
113.199 |
|
C3 |
C4 |
H12 |
109.670 |
| C3 |
C4 |
H13 |
109.670 |
|
C4 |
C3 |
H11 |
116.601 |
| C4 |
C5 |
C6 |
113.156 |
|
C4 |
C5 |
H14 |
109.155 |
| C4 |
C5 |
H15 |
109.155 |
|
C5 |
C4 |
H12 |
109.003 |
| C5 |
C4 |
H13 |
109.003 |
|
C5 |
C6 |
H16 |
111.408 |
| C5 |
C6 |
H17 |
111.131 |
|
C5 |
C6 |
H18 |
111.131 |
| C6 |
C5 |
H14 |
109.466 |
|
C6 |
C5 |
H15 |
109.466 |
| H7 |
C1 |
H8 |
108.105 |
|
H7 |
C1 |
H9 |
108.105 |
| H8 |
C1 |
H9 |
106.899 |
|
H12 |
C4 |
H13 |
106.042 |
| H14 |
C5 |
H15 |
106.205 |
|
H16 |
C6 |
H17 |
107.688 |
| H16 |
C6 |
H18 |
107.688 |
|
H17 |
C6 |
H18 |
107.615 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.134 |
|
|
|
| 2 |
C |
-0.085 |
|
|
|
| 3 |
C |
-0.137 |
|
|
|
| 4 |
C |
-0.008 |
|
|
|
| 5 |
C |
-0.009 |
|
|
|
| 6 |
C |
-0.190 |
|
|
|
| 7 |
H |
0.055 |
|
|
|
| 8 |
H |
0.058 |
|
|
|
| 9 |
H |
0.058 |
|
|
|
| 10 |
H |
0.040 |
|
|
|
| 11 |
H |
0.045 |
|
|
|
| 12 |
H |
0.044 |
|
|
|
| 13 |
H |
0.044 |
|
|
|
| 14 |
H |
0.031 |
|
|
|
| 15 |
H |
0.031 |
|
|
|
| 16 |
H |
0.061 |
|
|
|
| 17 |
H |
0.048 |
|
|
|
| 18 |
H |
0.048 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.031 |
-0.034 |
0.000 |
0.047 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.151 |
0.441 |
0.000 |
| y |
0.441 |
-39.956 |
0.000 |
| z |
0.000 |
0.000 |
-41.388 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.521 |
0.441 |
0.000 |
| y |
0.441 |
0.313 |
0.000 |
| z |
0.000 |
0.000 |
-1.834 |
|
| Polar |
|---|
| 3z2-r2 | -3.668 |
|---|
| x2-y2 | 0.806 |
|---|
| xy | 0.441 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.169 |
1.690 |
0.000 |
| y |
1.690 |
13.539 |
0.000 |
| z |
0.000 |
0.000 |
8.377 |
<r2> (average value of r
2) Å
2
| <r2> |
303.413 |
| (<r2>)1/2 |
17.419 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -234.270405 |
| Energy at 298.15K | -234.283009 |
| HF Energy | -234.270405 |
| Nuclear repulsion energy | 227.487945 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3269 |
2963 |
57.81 |
|
|
|
| 2 |
A |
3258 |
2953 |
15.34 |
|
|
|
| 3 |
A |
3225 |
2924 |
14.46 |
|
|
|
| 4 |
A |
3213 |
2913 |
68.88 |
|
|
|
| 5 |
A |
3213 |
2912 |
94.30 |
|
|
|
| 6 |
A |
3192 |
2894 |
33.86 |
|
|
|
| 7 |
A |
3188 |
2889 |
10.10 |
|
|
|
| 8 |
A |
3169 |
2872 |
12.94 |
|
|
|
| 9 |
A |
3154 |
2858 |
15.60 |
|
|
|
| 10 |
A |
3152 |
2857 |
63.07 |
|
|
|
| 11 |
A |
3149 |
2854 |
52.05 |
|
|
|
| 12 |
A |
3136 |
2843 |
22.32 |
|
|
|
| 13 |
A |
1879 |
1703 |
0.46 |
|
|
|
| 14 |
A |
1626 |
1474 |
6.07 |
|
|
|
| 15 |
A |
1614 |
1463 |
4.71 |
|
|
|
| 16 |
A |
1611 |
1460 |
5.20 |
|
|
|
| 17 |
A |
1609 |
1459 |
5.71 |
|
|
|
| 18 |
A |
1603 |
1453 |
1.28 |
|
|
|
| 19 |
A |
1597 |
1448 |
5.99 |
|
|
|
| 20 |
A |
1539 |
1395 |
2.99 |
|
|
|
| 21 |
A |
1537 |
1393 |
1.29 |
|
|
|
| 22 |
A |
1519 |
1376 |
0.21 |
|
|
|
| 23 |
A |
1461 |
1324 |
0.58 |
|
|
|
| 24 |
A |
1447 |
1311 |
0.58 |
|
|
|
| 25 |
A |
1426 |
1292 |
0.27 |
|
|
|
| 26 |
A |
1405 |
1273 |
8.62 |
|
|
|
| 27 |
A |
1357 |
1230 |
0.39 |
|
|
|
| 28 |
A |
1271 |
1152 |
0.06 |
|
|
|
| 29 |
A |
1206 |
1093 |
0.40 |
|
|
|
| 30 |
A |
1174 |
1064 |
3.45 |
|
|
|
| 31 |
A |
1154 |
1046 |
0.28 |
|
|
|
| 32 |
A |
1127 |
1022 |
11.11 |
|
|
|
| 33 |
A |
1101 |
998 |
35.04 |
|
|
|
| 34 |
A |
1089 |
988 |
4.42 |
|
|
|
| 35 |
A |
989 |
896 |
3.13 |
|
|
|
| 36 |
A |
968 |
878 |
1.22 |
|
|
|
| 37 |
A |
939 |
851 |
0.58 |
|
|
|
| 38 |
A |
857 |
777 |
0.26 |
|
|
|
| 39 |
A |
796 |
722 |
1.55 |
|
|
|
| 40 |
A |
570 |
517 |
0.15 |
|
|
|
| 41 |
A |
413 |
374 |
1.31 |
|
|
|
| 42 |
A |
332 |
301 |
0.79 |
|
|
|
| 43 |
A |
306 |
278 |
1.63 |
|
|
|
| 44 |
A |
262 |
237 |
0.03 |
|
|
|
| 45 |
A |
225 |
204 |
0.59 |
|
|
|
| 46 |
A |
156 |
141 |
0.33 |
|
|
|
| 47 |
A |
99 |
89 |
0.09 |
|
|
|
| 48 |
A |
85 |
77 |
0.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38332.1 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 34744.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
3.170 |
0.222 |
-0.155 |
| C2 |
1.789 |
-0.357 |
-0.227 |
| C3 |
0.748 |
0.087 |
0.438 |
| C4 |
-0.639 |
-0.482 |
0.360 |
| C5 |
-1.674 |
0.521 |
-0.157 |
| C6 |
-3.089 |
-0.045 |
-0.191 |
| H7 |
3.211 |
1.068 |
0.519 |
| H8 |
3.885 |
-0.519 |
0.190 |
| H9 |
3.506 |
0.552 |
-1.132 |
| H10 |
1.659 |
-1.204 |
-0.879 |
| H11 |
0.879 |
0.934 |
1.092 |
| H12 |
-0.944 |
-0.817 |
1.348 |
| H13 |
-0.636 |
-1.361 |
-0.275 |
| H14 |
-1.385 |
0.845 |
-1.151 |
| H15 |
-1.655 |
1.409 |
0.469 |
| H16 |
-3.798 |
0.684 |
-0.562 |
| H17 |
-3.145 |
-0.916 |
-0.834 |
| H18 |
-3.413 |
-0.347 |
0.799 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.4987 | 2.4973 | 3.9073 | 4.8535 | 6.2646 | 1.0821 | 1.0850 | 1.0850 | 2.2004 | 2.7035 | 4.5016 | 4.1243 | 4.7043 | 5.0084 | 6.9953 | 6.4528 | 6.6766 |
C2 | 1.4987 | | 1.3128 | 2.5011 | 3.5737 | 4.8881 | 2.1458 | 2.1424 | 2.1427 | 1.0772 | 2.0573 | 3.1883 | 2.6261 | 3.5179 | 3.9328 | 5.6934 | 5.0031 | 5.3031 | C3 | 2.4973 | 1.3128 | | 1.5009 | 2.5318 | 3.8902 | 2.6522 | 3.2039 | 3.2075 | 2.0567 | 1.0780 | 2.1230 | 2.1266 | 2.7662 | 2.7433 | 4.6931 | 4.2168 | 4.1995 | C4 | 3.9073 | 2.5011 | 1.5009 | | 1.5316 | 2.5487 | 4.1528 | 4.5265 | 4.5245 | 2.7079 | 2.2012 | 1.0874 | 1.0846 | 2.1450 | 2.1496 | 3.4916 | 2.8099 | 2.8126 | C5 | 4.8535 | 3.5737 | 2.5318 | 1.5316 | | 1.5242 | 4.9615 | 5.6656 | 5.2708 | 3.8216 | 2.8722 | 2.1419 | 2.1529 | 1.0849 | 1.0861 | 2.1684 | 2.1655 | 2.1663 | C6 | 6.2646 | 4.8881 | 3.8902 | 2.5487 | 1.5242 | | 6.4362 | 6.9995 | 6.6879 | 4.9348 | 4.2835 | 2.7498 | 2.7843 | 2.1489 | 2.1456 | 1.0833 | 1.0844 | 1.0844 | H7 | 1.0821 | 2.1458 | 2.6522 | 4.1528 | 4.9615 | 6.4362 | | 1.7543 | 1.7544 | 3.0859 | 2.4046 | 4.6371 | 4.6186 | 4.8947 | 4.8783 | 7.1019 | 6.7943 | 6.7793 | H8 | 1.0850 | 2.1424 | 3.2039 | 4.5265 | 5.6656 | 6.9995 | 1.7543 | | 1.7434 | 2.5629 | 3.4572 | 4.9744 | 4.6223 | 5.6060 | 5.8721 | 7.8124 | 7.1149 | 7.3253 | H9 | 1.0850 | 2.1427 | 3.2075 | 4.5245 | 5.2708 | 6.6879 | 1.7544 | 1.7434 | | 2.5612 | 3.4626 | 5.2751 | 4.6424 | 4.8994 | 5.4712 | 7.3270 | 6.8174 | 7.2396 | H10 | 2.2004 | 1.0772 | 2.0567 | 2.7079 | 3.8216 | 4.9348 | 3.0859 | 2.5629 | 2.5612 | | 3.0102 | 3.4474 | 2.3782 | 3.6792 | 4.4300 | 5.7827 | 4.8124 | 5.4107 | H11 | 2.7035 | 2.0573 | 1.0780 | 2.2012 | 2.8722 | 4.2835 | 2.4046 | 3.4572 | 3.4626 | 3.0102 | | 2.5406 | 3.0716 | 3.1884 | 2.6530 | 4.9674 | 4.8299 | 4.4893 | H12 | 4.5016 | 3.1883 | 2.1230 | 1.0874 | 2.1419 | 2.7498 | 4.6371 | 4.9744 | 5.2751 | 3.4474 | 2.5406 | | 1.7400 | 3.0335 | 2.4961 | 3.7478 | 3.1009 | 2.5728 | H13 | 4.1243 | 2.6261 | 2.1266 | 1.0846 | 2.1529 | 2.7843 | 4.6186 | 4.6223 | 4.6424 | 2.3782 | 3.0716 | 1.7400 | | 2.4891 | 3.0439 | 3.7766 | 2.6083 | 3.1458 | H14 | 4.7043 | 3.5179 | 2.7662 | 2.1450 | 1.0849 | 2.1489 | 4.8947 | 5.6060 | 4.8994 | 3.6792 | 3.1884 | 3.0335 | 2.4891 | | 1.7365 | 2.4890 | 2.5103 | 3.0559 | H15 | 5.0084 | 3.9328 | 2.7433 | 2.1496 | 1.0861 | 2.1456 | 4.8783 | 5.8721 | 5.4712 | 4.4300 | 2.6530 | 2.4961 | 3.0439 | 1.7365 | | 2.4857 | 3.0534 | 2.5063 | H16 | 6.9953 | 5.6934 | 4.6931 | 3.4916 | 2.1684 | 1.0833 | 7.1019 | 7.8124 | 7.3270 | 5.7827 | 4.9674 | 3.7478 | 3.7766 | 2.4890 | 2.4857 | | 1.7501 | 1.7504 | H17 | 6.4528 | 5.0031 | 4.2168 | 2.8099 | 2.1655 | 1.0844 | 6.7943 | 7.1149 | 6.8174 | 4.8124 | 4.8299 | 3.1009 | 2.6083 | 2.5103 | 3.0534 | 1.7501 | | 1.7503 | H18 | 6.6766 | 5.3031 | 4.1995 | 2.8126 | 2.1663 | 1.0844 | 6.7793 | 7.3253 | 7.2396 | 5.4107 | 4.4893 | 2.5728 | 3.1458 | 3.0559 | 2.5063 | 1.7504 | 1.7503 | |
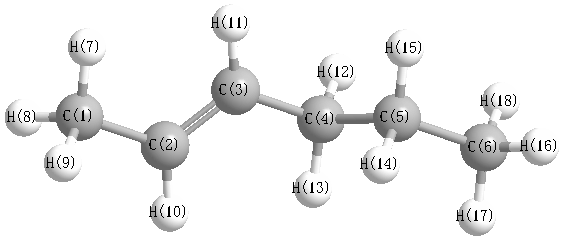 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.182 |
|
C1 |
C2 |
H10 |
116.395 |
| C2 |
C1 |
H7 |
111.482 |
|
C2 |
C1 |
H8 |
111.019 |
| C2 |
C1 |
H9 |
111.043 |
|
C2 |
C3 |
C4 |
125.339 |
| C2 |
C3 |
H11 |
118.416 |
|
C3 |
C2 |
H10 |
118.422 |
| C3 |
C4 |
C5 |
113.208 |
|
C3 |
C4 |
H12 |
109.173 |
| C3 |
C4 |
H13 |
109.628 |
|
C4 |
C3 |
H11 |
116.242 |
| C4 |
C5 |
C6 |
113.037 |
|
C4 |
C5 |
H14 |
108.936 |
| C4 |
C5 |
H15 |
109.230 |
|
C5 |
C4 |
H12 |
108.552 |
| C5 |
C4 |
H13 |
109.574 |
|
C5 |
C6 |
H16 |
111.401 |
| C5 |
C6 |
H17 |
111.104 |
|
C5 |
C6 |
H18 |
111.165 |
| C6 |
C5 |
H14 |
109.750 |
|
C6 |
C5 |
H15 |
109.421 |
| H7 |
C1 |
H8 |
108.096 |
|
H7 |
C1 |
H9 |
108.105 |
| H8 |
C1 |
H9 |
106.915 |
|
H12 |
C4 |
H13 |
106.478 |
| H14 |
C5 |
H15 |
106.239 |
|
H16 |
C6 |
H17 |
107.675 |
| H16 |
C6 |
H18 |
107.700 |
|
H17 |
C6 |
H18 |
107.616 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.137 |
|
|
|
| 2 |
C |
-0.071 |
|
|
|
| 3 |
C |
-0.134 |
|
|
|
| 4 |
C |
-0.012 |
|
|
|
| 5 |
C |
-0.011 |
|
|
|
| 6 |
C |
-0.187 |
|
|
|
| 7 |
H |
0.056 |
|
|
|
| 8 |
H |
0.059 |
|
|
|
| 9 |
H |
0.058 |
|
|
|
| 10 |
H |
0.042 |
|
|
|
| 11 |
H |
0.042 |
|
|
|
| 12 |
H |
0.043 |
|
|
|
| 13 |
H |
0.039 |
|
|
|
| 14 |
H |
0.034 |
|
|
|
| 15 |
H |
0.026 |
|
|
|
| 16 |
H |
0.060 |
|
|
|
| 17 |
H |
0.047 |
|
|
|
| 18 |
H |
0.047 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.005 |
0.039 |
-0.046 |
0.061 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.887 |
-0.157 |
-0.709 |
| y |
-0.157 |
-40.306 |
1.406 |
| z |
-0.709 |
1.406 |
-40.371 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.452 |
-0.157 |
-0.709 |
| y |
-0.157 |
-0.177 |
1.406 |
| z |
-0.709 |
1.406 |
-0.275 |
|
| Polar |
|---|
| 3z2-r2 | -0.550 |
|---|
| x2-y2 | 0.419 |
|---|
| xy | -0.157 |
|---|
| xz | -0.709 |
|---|
| yz | 1.406 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.961 |
-0.326 |
-0.861 |
| y |
-0.326 |
9.150 |
0.410 |
| z |
-0.861 |
0.410 |
9.049 |
<r2> (average value of r
2) Å
2
| <r2> |
294.446 |
| (<r2>)1/2 |
17.159 |
