Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -375.958483 |
| Energy at 298.15K | |
| HF Energy | -375.958483 |
| Nuclear repulsion energy | 193.359673 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3282 |
2975 |
44.64 |
24.39 |
0.36 |
0.53 |
| 2 |
A |
3260 |
2955 |
16.48 |
96.48 |
0.38 |
0.55 |
| 3 |
A |
3221 |
2919 |
21.91 |
112.53 |
0.07 |
0.13 |
| 4 |
A |
1630 |
1478 |
7.53 |
3.81 |
0.75 |
0.85 |
| 5 |
A |
1608 |
1457 |
13.13 |
1.98 |
0.32 |
0.48 |
| 6 |
A |
1539 |
1395 |
27.48 |
2.30 |
0.74 |
0.85 |
| 7 |
A |
1476 |
1338 |
24.63 |
1.44 |
0.75 |
0.86 |
| 8 |
A |
1382 |
1252 |
21.39 |
4.64 |
0.70 |
0.83 |
| 9 |
A |
1279 |
1159 |
116.22 |
3.60 |
0.68 |
0.81 |
| 10 |
A |
1244 |
1128 |
147.85 |
3.27 |
0.75 |
0.86 |
| 11 |
A |
1229 |
1114 |
118.29 |
1.49 |
0.27 |
0.42 |
| 12 |
A |
1199 |
1087 |
84.03 |
4.56 |
0.43 |
0.60 |
| 13 |
A |
990 |
897 |
36.44 |
4.49 |
0.27 |
0.42 |
| 14 |
A |
636 |
576 |
5.44 |
1.50 |
0.31 |
0.47 |
| 15 |
A |
519 |
470 |
23.66 |
0.48 |
0.73 |
0.85 |
| 16 |
A |
465 |
421 |
6.04 |
1.29 |
0.43 |
0.60 |
| 17 |
A |
267 |
242 |
9.35 |
0.05 |
0.56 |
0.72 |
| 18 |
A |
129 |
117 |
9.74 |
0.02 |
0.71 |
0.83 |
Unscaled Zero Point Vibrational Energy (zpe) 12676.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 11490.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.781 |
-0.578 |
-0.280 |
| C2 |
0.467 |
0.019 |
0.322 |
| F3 |
-1.858 |
0.122 |
0.151 |
| F4 |
1.505 |
-0.756 |
-0.006 |
| F5 |
0.693 |
1.231 |
-0.182 |
| H6 |
-0.728 |
-0.524 |
-1.358 |
| H7 |
-0.879 |
-1.606 |
0.037 |
| H8 |
0.415 |
0.101 |
1.397 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5088 | 1.3554 | 2.3091 | 2.3360 | 1.0808 | 1.0805 | 2.1688 |
C2 | 1.5088 | | 2.3334 | 1.3362 | 1.3321 | 2.1326 | 2.1296 | 1.0797 | F3 | 1.3554 | 2.3334 | | 3.4793 | 2.8015 | 1.9933 | 1.9899 | 2.5915 | F4 | 2.3091 | 1.3362 | 3.4793 | | 2.1536 | 2.6211 | 2.5318 | 1.9732 | F5 | 2.3360 | 1.3321 | 2.8015 | 2.1536 | | 2.5473 | 3.2515 | 1.9617 | H6 | 1.0808 | 2.1326 | 1.9933 | 2.6211 | 2.5473 | | 1.7722 | 3.0481 | H7 | 1.0805 | 2.1296 | 1.9899 | 2.5318 | 3.2515 | 1.7722 | | 2.5373 | H8 | 2.1688 | 1.0797 | 2.5915 | 1.9732 | 1.9617 | 3.0481 | 2.5373 | |
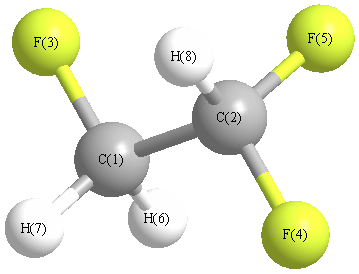 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.360 |
|
C1 |
C2 |
F5 |
110.473 |
| C1 |
C2 |
H8 |
112.781 |
|
C2 |
C1 |
F3 |
109.000 |
| C2 |
C1 |
H6 |
109.779 |
|
C2 |
C1 |
H7 |
109.552 |
| F3 |
C1 |
H6 |
109.296 |
|
F3 |
C1 |
H7 |
109.029 |
| F4 |
C2 |
F5 |
107.630 |
|
F4 |
C2 |
H8 |
109.062 |
| F5 |
C2 |
H8 |
108.400 |
|
H6 |
C1 |
H7 |
110.162 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.217 |
|
|
|
| 2 |
C |
0.481 |
|
|
|
| 3 |
F |
-0.290 |
|
|
|
| 4 |
F |
-0.273 |
|
|
|
| 5 |
F |
-0.265 |
|
|
|
| 6 |
H |
0.056 |
|
|
|
| 7 |
H |
0.047 |
|
|
|
| 8 |
H |
0.027 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.318 |
-1.571 |
0.416 |
1.656 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.592 |
1.836 |
1.651 |
| y |
1.836 |
-27.879 |
0.554 |
| z |
1.651 |
0.554 |
-24.488 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.408 |
1.836 |
1.651 |
| y |
1.836 |
0.661 |
0.554 |
| z |
1.651 |
0.554 |
5.748 |
|
| Polar |
|---|
| 3z2-r2 | 11.495 |
|---|
| x2-y2 | -4.713 |
|---|
| xy | 1.836 |
|---|
| xz | 1.651 |
|---|
| yz | 0.554 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.737 |
-0.038 |
-0.011 |
| y |
-0.038 |
3.683 |
0.037 |
| z |
-0.011 |
0.037 |
3.537 |
<r2> (average value of r
2) Å
2
| <r2> |
109.096 |
| (<r2>)1/2 |
10.445 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -375.956241 |
| Energy at 298.15K | |
| HF Energy | -375.956241 |
| Nuclear repulsion energy | 196.060896 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3239 |
2936 |
63.28 |
107.48 |
0.08 |
0.15 |
| 2 |
A' |
3209 |
2909 |
10.45 |
78.41 |
0.19 |
0.32 |
| 3 |
A' |
1625 |
1473 |
10.05 |
3.01 |
0.74 |
0.85 |
| 4 |
A' |
1582 |
1434 |
24.37 |
2.61 |
0.27 |
0.42 |
| 5 |
A' |
1551 |
1406 |
27.75 |
1.52 |
0.69 |
0.81 |
| 6 |
A' |
1305 |
1183 |
156.39 |
3.55 |
0.41 |
0.59 |
| 7 |
A' |
1216 |
1103 |
33.34 |
3.51 |
0.75 |
0.86 |
| 8 |
A' |
942 |
854 |
43.53 |
6.74 |
0.14 |
0.24 |
| 9 |
A' |
827 |
750 |
63.10 |
2.31 |
0.46 |
0.63 |
| 10 |
A' |
570 |
516 |
16.23 |
0.96 |
0.75 |
0.86 |
| 11 |
A' |
257 |
233 |
2.24 |
0.16 |
0.46 |
0.63 |
| 12 |
A" |
3262 |
2957 |
26.27 |
46.55 |
0.75 |
0.86 |
| 13 |
A" |
1532 |
1389 |
34.93 |
1.01 |
0.75 |
0.86 |
| 14 |
A" |
1402 |
1270 |
28.29 |
7.85 |
0.75 |
0.86 |
| 15 |
A" |
1267 |
1149 |
130.01 |
1.51 |
0.75 |
0.86 |
| 16 |
A" |
1057 |
958 |
66.18 |
2.35 |
0.75 |
0.86 |
| 17 |
A" |
399 |
361 |
0.24 |
0.15 |
0.75 |
0.86 |
| 18 |
A" |
127 |
116 |
2.56 |
0.04 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12684.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 11497.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.726 |
-0.835 |
0.000 |
| C2 |
0.348 |
0.625 |
0.000 |
| F3 |
-0.381 |
-1.609 |
0.000 |
| F4 |
-0.381 |
0.922 |
1.075 |
| F5 |
-0.381 |
0.922 |
-1.075 |
| H6 |
1.308 |
-1.052 |
-0.885 |
| H7 |
1.308 |
-1.052 |
0.885 |
| H8 |
1.230 |
1.252 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5075 | 1.3509 | 2.3379 | 2.3379 | 1.0816 | 1.0816 | 2.1465 |
C2 | 1.5075 | | 2.3504 | 1.3327 | 1.3327 | 2.1252 | 2.1252 | 1.0819 | F3 | 1.3509 | 2.3504 | | 2.7501 | 2.7501 | 1.9865 | 1.9865 | 3.2837 | F4 | 2.3379 | 1.3327 | 2.7501 | | 2.1501 | 3.2545 | 2.6047 | 1.9651 | F5 | 2.3379 | 1.3327 | 2.7501 | 2.1501 | | 2.6047 | 3.2545 | 1.9651 | H6 | 1.0816 | 2.1252 | 1.9865 | 3.2545 | 2.6047 | | 1.7706 | 2.4692 | H7 | 1.0816 | 2.1252 | 1.9865 | 2.6047 | 3.2545 | 1.7706 | | 2.4692 | H8 | 2.1465 | 1.0819 | 3.2837 | 1.9651 | 1.9651 | 2.4692 | 2.4692 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.649 |
|
C1 |
C2 |
F5 |
110.649 |
| C1 |
C2 |
H8 |
110.911 |
|
C2 |
C1 |
F3 |
110.509 |
| C2 |
C1 |
H6 |
109.224 |
|
C2 |
C1 |
H7 |
109.224 |
| F3 |
C1 |
H6 |
109.000 |
|
F3 |
C1 |
H7 |
109.000 |
| F4 |
C2 |
F5 |
107.540 |
|
F4 |
C2 |
H8 |
108.493 |
| F5 |
C2 |
H8 |
108.493 |
|
H6 |
C1 |
H7 |
109.871 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.218 |
|
|
|
| 2 |
C |
0.497 |
|
|
|
| 3 |
F |
-0.281 |
|
|
|
| 4 |
F |
-0.265 |
|
|
|
| 5 |
F |
-0.265 |
|
|
|
| 6 |
H |
0.043 |
|
|
|
| 7 |
H |
0.043 |
|
|
|
| 8 |
H |
0.011 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.382 |
0.090 |
0.000 |
3.383 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.095 |
-0.368 |
0.000 |
| y |
-0.368 |
-28.772 |
0.000 |
| z |
0.000 |
0.000 |
-28.547 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.565 |
-0.368 |
0.000 |
| y |
-0.368 |
-1.951 |
0.000 |
| z |
0.000 |
0.000 |
-1.613 |
|
| Polar |
|---|
| 3z2-r2 | -3.227 |
|---|
| x2-y2 | 3.677 |
|---|
| xy | -0.368 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.675 |
-0.003 |
0.000 |
| y |
-0.003 |
3.668 |
0.000 |
| z |
0.000 |
0.000 |
3.641 |
<r2> (average value of r
2) Å
2
| <r2> |
101.183 |
| (<r2>)1/2 |
10.059 |
