Jump to
S1C2
Energy calculated at QCISD(T)=FULL/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -377.186460 |
| Energy at 298.15K | |
| HF Energy | -375.956938 |
| Nuclear repulsion energy | 191.152787 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/Def2TZVPP
Geometric Data calculated at QCISD(T)=FULL/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.772 |
-0.592 |
-0.282 |
| C2 |
0.467 |
0.020 |
0.329 |
| F3 |
-1.877 |
0.110 |
0.150 |
| F4 |
1.531 |
-0.757 |
-0.008 |
| F5 |
0.678 |
1.257 |
-0.182 |
| H6 |
-0.707 |
-0.532 |
-1.367 |
| H7 |
-0.861 |
-1.629 |
0.036 |
| H8 |
0.415 |
0.101 |
1.412 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5109 | 1.3780 | 2.3250 | 2.3517 | 1.0882 | 1.0880 | 2.1820 |
C2 | 1.5109 | | 2.3518 | 1.3597 | 1.3546 | 2.1346 | 2.1368 | 1.0876 | F3 | 1.3780 | 2.3518 | | 3.5193 | 2.8195 | 2.0198 | 2.0171 | 2.6162 | F4 | 2.3250 | 1.3597 | 3.5193 | | 2.1937 | 2.6276 | 2.5455 | 1.9994 | F5 | 2.3517 | 1.3546 | 2.8195 | 2.1937 | | 2.5535 | 3.2772 | 1.9861 | H6 | 1.0882 | 2.1346 | 2.0198 | 2.6276 | 2.5535 | | 1.7871 | 3.0630 | H7 | 1.0880 | 2.1368 | 2.0171 | 2.5455 | 3.2772 | 1.7871 | | 2.5525 | H8 | 2.1820 | 1.0876 | 2.6162 | 1.9994 | 1.9861 | 3.0630 | 2.5525 | |
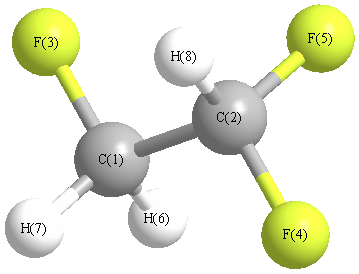 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.068 |
|
C1 |
C2 |
F5 |
110.192 |
| C1 |
C2 |
H8 |
113.210 |
|
C2 |
C1 |
F3 |
108.907 |
| C2 |
C1 |
H6 |
109.355 |
|
C2 |
C1 |
H7 |
109.535 |
| F3 |
C1 |
H6 |
109.406 |
|
F3 |
C1 |
H7 |
109.203 |
| F4 |
C2 |
F5 |
107.844 |
|
F4 |
C2 |
H8 |
109.060 |
| F5 |
C2 |
H8 |
108.330 |
|
H6 |
C1 |
H7 |
110.411 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)=FULL/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -377.184396 |
| Energy at 298.15K | |
| HF Energy | -375.954696 |
| Nuclear repulsion energy | 193.875445 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3087 |
3087 |
|
|
|
|
| 2 |
A' |
3067 |
3067 |
|
|
|
|
| 3 |
A' |
1501 |
1501 |
|
|
|
|
| 4 |
A' |
1455 |
1455 |
|
|
|
|
| 5 |
A' |
1433 |
1433 |
|
|
|
|
| 6 |
A' |
1207 |
1207 |
|
|
|
|
| 7 |
A' |
1123 |
1123 |
|
|
|
|
| 8 |
A' |
885 |
885 |
|
|
|
|
| 9 |
A' |
767 |
767 |
|
|
|
|
| 10 |
A' |
523 |
523 |
|
|
|
|
| 11 |
A' |
236 |
236 |
|
|
|
|
| 12 |
A" |
3126 |
3126 |
|
|
|
|
| 13 |
A" |
1409 |
1409 |
|
|
|
|
| 14 |
A" |
1299 |
1299 |
|
|
|
|
| 15 |
A" |
1163 |
1163 |
|
|
|
|
| 16 |
A" |
970 |
970 |
|
|
|
|
| 17 |
A" |
371 |
371 |
|
|
|
|
| 18 |
A" |
117 |
117 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11869.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11869.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)=FULL/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.738 |
-0.832 |
0.000 |
| C2 |
0.356 |
0.629 |
0.000 |
| F3 |
-0.387 |
-1.619 |
0.000 |
| F4 |
-0.387 |
0.924 |
1.094 |
| F5 |
-0.387 |
0.924 |
-1.094 |
| H6 |
1.324 |
-1.047 |
-0.893 |
| H7 |
1.324 |
-1.047 |
0.893 |
| H8 |
1.240 |
1.268 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5100 | 1.3731 | 2.3553 | 2.3553 | 1.0891 | 1.0891 | 2.1591 |
C2 | 1.5100 | | 2.3678 | 1.3557 | 1.3557 | 2.1311 | 2.1311 | 1.0901 | F3 | 1.3731 | 2.3678 | | 2.7685 | 2.7685 | 2.0130 | 2.0130 | 3.3139 | F4 | 2.3553 | 1.3557 | 2.7685 | | 2.1890 | 3.2804 | 2.6179 | 1.9907 | F5 | 2.3553 | 1.3557 | 2.7685 | 2.1890 | | 2.6179 | 3.2804 | 1.9907 | H6 | 1.0891 | 2.1311 | 2.0130 | 3.2804 | 2.6179 | | 1.7851 | 2.4826 | H7 | 1.0891 | 2.1311 | 2.0130 | 2.6179 | 3.2804 | 1.7851 | | 2.4826 | H8 | 2.1591 | 1.0901 | 3.3139 | 1.9907 | 1.9907 | 2.4826 | 2.4826 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.436 |
|
C1 |
C2 |
F5 |
110.436 |
| C1 |
C2 |
H8 |
111.254 |
|
C2 |
C1 |
F3 |
110.333 |
| C2 |
C1 |
H6 |
109.082 |
|
C2 |
C1 |
H7 |
109.082 |
| F3 |
C1 |
H6 |
109.131 |
|
F3 |
C1 |
H7 |
109.131 |
| F4 |
C2 |
F5 |
107.670 |
|
F4 |
C2 |
H8 |
108.470 |
| F5 |
C2 |
H8 |
108.470 |
|
H6 |
C1 |
H7 |
110.072 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
