Jump to
S1C2
S1C3
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -304.813911 |
| Energy at 298.15K | -304.820355 |
| HF Energy | -304.813911 |
| Nuclear repulsion energy | 228.226685 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3285 |
2978 |
0.00 |
|
|
|
| 2 |
Ag |
3187 |
2888 |
0.00 |
|
|
|
| 3 |
Ag |
2007 |
1819 |
0.00 |
|
|
|
| 4 |
Ag |
1577 |
1430 |
0.00 |
|
|
|
| 5 |
Ag |
1534 |
1391 |
0.00 |
|
|
|
| 6 |
Ag |
1406 |
1275 |
0.00 |
|
|
|
| 7 |
Ag |
1107 |
1004 |
0.00 |
|
|
|
| 8 |
Ag |
742 |
673 |
0.00 |
|
|
|
| 9 |
Ag |
571 |
518 |
0.00 |
|
|
|
| 10 |
Ag |
386 |
350 |
0.00 |
|
|
|
| 11 |
Au |
3247 |
2943 |
16.10 |
|
|
|
| 12 |
Au |
1584 |
1436 |
17.80 |
|
|
|
| 13 |
Au |
1062 |
962 |
4.22 |
|
|
|
| 14 |
Au |
395 |
358 |
12.54 |
|
|
|
| 15 |
Au |
142 |
129 |
0.12 |
|
|
|
| 16 |
Au |
50 |
45 |
13.02 |
|
|
|
| 17 |
Bg |
3247 |
2943 |
0.00 |
|
|
|
| 18 |
Bg |
1588 |
1439 |
0.00 |
|
|
|
| 19 |
Bg |
1184 |
1073 |
0.00 |
|
|
|
| 20 |
Bg |
682 |
618 |
0.00 |
|
|
|
| 21 |
Bg |
144 |
130 |
0.00 |
|
|
|
| 22 |
Bu |
3286 |
2978 |
23.08 |
|
|
|
| 23 |
Bu |
3186 |
2888 |
4.42 |
|
|
|
| 24 |
Bu |
1983 |
1798 |
379.87 |
|
|
|
| 25 |
Bu |
1577 |
1430 |
27.22 |
|
|
|
| 26 |
Bu |
1523 |
1381 |
37.61 |
|
|
|
| 27 |
Bu |
1228 |
1113 |
82.68 |
|
|
|
| 28 |
Bu |
981 |
889 |
21.04 |
|
|
|
| 29 |
Bu |
589 |
534 |
58.73 |
|
|
|
| 30 |
Bu |
255 |
231 |
18.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21866.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 19820.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.055 |
0.744 |
0.000 |
| C2 |
0.055 |
-0.744 |
0.000 |
| C3 |
1.162 |
1.595 |
0.000 |
| C4 |
-1.162 |
-1.595 |
0.000 |
| O5 |
-1.162 |
1.211 |
0.000 |
| O6 |
1.162 |
-1.211 |
0.000 |
| H7 |
0.867 |
2.632 |
0.000 |
| H8 |
-0.867 |
-2.632 |
0.000 |
| H9 |
1.762 |
1.380 |
0.875 |
| H10 |
1.762 |
1.380 |
-0.875 |
| H11 |
-1.762 |
-1.380 |
0.875 |
| H12 |
-1.762 |
-1.380 |
-0.875 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
O5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4911 | 1.4848 | 2.5874 | 1.2018 | 2.3025 | 2.1018 | 3.4723 | 2.1147 | 2.1147 | 2.8622 | 2.8622 |
C2 | 1.4911 | | 2.5874 | 1.4848 | 2.3025 | 1.2018 | 3.4723 | 2.1018 | 2.8622 | 2.8622 | 2.1147 | 2.1147 | C3 | 1.4848 | 2.5874 | | 3.9466 | 2.3547 | 2.8066 | 1.0782 | 4.6891 | 1.0827 | 1.0827 | 4.2624 | 4.2624 | C4 | 2.5874 | 1.4848 | 3.9466 | | 2.8066 | 2.3547 | 4.6891 | 1.0782 | 4.2624 | 4.2624 | 1.0827 | 1.0827 | O5 | 1.2018 | 2.3025 | 2.3547 | 2.8066 | | 3.3568 | 2.4768 | 3.8551 | 3.0565 | 3.0565 | 2.8007 | 2.8007 | O6 | 2.3025 | 1.2018 | 2.8066 | 2.3547 | 3.3568 | | 3.8551 | 2.4768 | 2.8007 | 2.8007 | 3.0565 | 3.0565 | H7 | 2.1018 | 3.4723 | 1.0782 | 4.6891 | 2.4768 | 3.8551 | | 5.5430 | 1.7703 | 1.7703 | 4.8766 | 4.8766 | H8 | 3.4723 | 2.1018 | 4.6891 | 1.0782 | 3.8551 | 2.4768 | 5.5430 | | 4.8766 | 4.8766 | 1.7703 | 1.7703 | H9 | 2.1147 | 2.8622 | 1.0827 | 4.2624 | 3.0565 | 2.8007 | 1.7703 | 4.8766 | | 1.7500 | 4.4769 | 4.8067 | H10 | 2.1147 | 2.8622 | 1.0827 | 4.2624 | 3.0565 | 2.8007 | 1.7703 | 4.8766 | 1.7500 | | 4.8067 | 4.4769 | H11 | 2.8622 | 2.1147 | 4.2624 | 1.0827 | 2.8007 | 3.0565 | 4.8766 | 1.7703 | 4.4769 | 4.8067 | | 1.7500 | H12 | 2.8622 | 2.1147 | 4.2624 | 1.0827 | 2.8007 | 3.0565 | 4.8766 | 1.7703 | 4.8067 | 4.4769 | 1.7500 | |
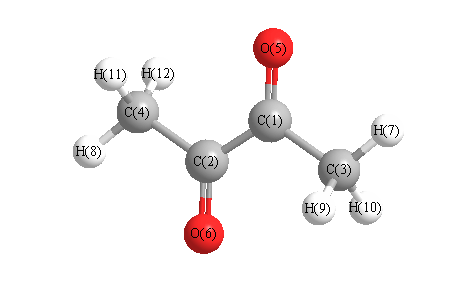 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
120.791 |
|
C1 |
C2 |
O6 |
117.122 |
| C1 |
C3 |
H7 |
109.144 |
|
C1 |
C3 |
H9 |
109.911 |
| C1 |
C3 |
H10 |
109.911 |
|
C2 |
C1 |
C3 |
120.791 |
| C2 |
C1 |
O5 |
117.122 |
|
C2 |
C4 |
H8 |
109.144 |
| C2 |
C4 |
H11 |
109.911 |
|
C2 |
C4 |
H12 |
109.911 |
| C3 |
C1 |
O5 |
122.087 |
|
C4 |
C2 |
O6 |
122.087 |
| H7 |
C3 |
H9 |
110.014 |
|
H7 |
C3 |
H10 |
110.014 |
| H8 |
C4 |
H11 |
110.014 |
|
H8 |
C4 |
H12 |
110.014 |
| H9 |
C3 |
H10 |
107.835 |
|
H11 |
C4 |
H12 |
107.835 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.287 |
|
|
|
| 2 |
C |
0.287 |
|
|
|
| 3 |
C |
-0.161 |
|
|
|
| 4 |
C |
-0.161 |
|
|
|
| 5 |
O |
-0.391 |
|
|
|
| 6 |
O |
-0.391 |
|
|
|
| 7 |
H |
0.108 |
|
|
|
| 8 |
H |
0.108 |
|
|
|
| 9 |
H |
0.079 |
|
|
|
| 10 |
H |
0.079 |
|
|
|
| 11 |
H |
0.079 |
|
|
|
| 12 |
H |
0.079 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.376 |
8.038 |
0.000 |
| y |
8.038 |
-36.876 |
0.000 |
| z |
0.000 |
0.000 |
-33.632 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.123 |
8.038 |
0.000 |
| y |
8.038 |
-0.372 |
0.000 |
| z |
0.000 |
0.000 |
4.495 |
|
| Polar |
|---|
| 3z2-r2 | 8.989 |
|---|
| x2-y2 | -2.500 |
|---|
| xy | 8.038 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.314 |
-0.020 |
0.000 |
| y |
-0.020 |
7.613 |
0.000 |
| z |
0.000 |
0.000 |
5.420 |
<r2> (average value of r
2) Å
2
| <r2> |
161.359 |
| (<r2>)1/2 |
12.703 |
Jump to
S1C1
S1C3
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -304.813911 |
| Energy at 298.15K | -304.820358 |
| HF Energy | -304.813911 |
| Nuclear repulsion energy | 228.241181 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3286 |
2978 |
0.00 |
|
|
|
| 2 |
A |
3246 |
2942 |
16.05 |
|
|
|
| 3 |
A |
3186 |
2888 |
0.00 |
|
|
|
| 4 |
A |
2005 |
1818 |
0.00 |
|
|
|
| 5 |
A |
1584 |
1436 |
17.82 |
|
|
|
| 6 |
A |
1577 |
1430 |
0.00 |
|
|
|
| 7 |
A |
1535 |
1391 |
0.00 |
|
|
|
| 8 |
A |
1407 |
1275 |
0.00 |
|
|
|
| 9 |
A |
1107 |
1004 |
0.00 |
|
|
|
| 10 |
A |
1062 |
962 |
4.23 |
|
|
|
| 11 |
A |
743 |
673 |
0.00 |
|
|
|
| 12 |
A |
572 |
518 |
0.00 |
|
|
|
| 13 |
A |
395 |
358 |
12.58 |
|
|
|
| 14 |
A |
386 |
350 |
0.00 |
|
|
|
| 15 |
A |
142 |
129 |
0.11 |
|
|
|
| 16 |
A |
51 |
46 |
12.99 |
|
|
|
| 17 |
B |
3286 |
2979 |
23.16 |
|
|
|
| 18 |
B |
3246 |
2942 |
0.00 |
|
|
|
| 19 |
B |
3186 |
2888 |
4.30 |
|
|
|
| 20 |
B |
1982 |
1797 |
380.00 |
|
|
|
| 21 |
B |
1587 |
1439 |
0.00 |
|
|
|
| 22 |
B |
1577 |
1430 |
27.34 |
|
|
|
| 23 |
B |
1524 |
1381 |
37.80 |
|
|
|
| 24 |
B |
1229 |
1114 |
82.83 |
|
|
|
| 25 |
B |
1184 |
1073 |
0.00 |
|
|
|
| 26 |
B |
982 |
890 |
20.73 |
|
|
|
| 27 |
B |
682 |
618 |
0.00 |
|
|
|
| 28 |
B |
589 |
534 |
58.83 |
|
|
|
| 29 |
B |
255 |
232 |
18.52 |
|
|
|
| 30 |
B |
144 |
130 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21868.1 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 19821.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.075 |
0.769 |
-0.023 |
| C2 |
0.075 |
-0.769 |
-0.023 |
| C3 |
1.167 |
1.604 |
0.059 |
| C4 |
-1.167 |
-1.604 |
0.059 |
| O5 |
-1.167 |
1.224 |
-0.067 |
| O6 |
1.167 |
-1.224 |
-0.067 |
| H7 |
1.027 |
2.528 |
-0.484 |
| H8 |
-1.027 |
-2.528 |
-0.484 |
| H9 |
1.338 |
1.850 |
1.103 |
| H10 |
2.020 |
1.056 |
-0.301 |
| H11 |
-1.338 |
-1.850 |
1.103 |
| H12 |
-2.020 |
-1.056 |
-0.301 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
O5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5458 | 1.4990 | 2.6137 | 1.1828 | 2.3487 | 2.1265 | 3.4628 | 2.1063 | 2.1333 | 3.1186 | 2.6816 |
C2 | 1.5458 | | 2.6137 | 1.4990 | 2.3487 | 1.1828 | 3.4628 | 2.1265 | 3.1186 | 2.6816 | 2.1063 | 2.1333 | C3 | 1.4990 | 2.6137 | | 3.9674 | 2.3673 | 2.8310 | 1.0806 | 4.7102 | 1.0865 | 1.0762 | 4.3933 | 4.1669 | C4 | 2.6137 | 1.4990 | 3.9674 | | 2.8310 | 2.3673 | 4.7102 | 1.0806 | 4.3933 | 4.1669 | 1.0865 | 1.0762 | O5 | 1.1828 | 2.3487 | 2.3673 | 2.8310 | | 3.3814 | 2.5860 | 3.7778 | 2.8348 | 3.1996 | 3.2937 | 2.4458 | O6 | 2.3487 | 1.1828 | 2.8310 | 2.3673 | 3.3814 | | 3.7778 | 2.5860 | 3.2937 | 2.4458 | 2.8348 | 3.1996 | H7 | 2.1265 | 3.4628 | 1.0806 | 4.7102 | 2.5860 | 3.7778 | | 5.4578 | 1.7536 | 1.7852 | 5.2237 | 4.7082 | H8 | 3.4628 | 2.1265 | 4.7102 | 1.0806 | 3.7778 | 2.5860 | 5.4578 | | 5.2237 | 4.7082 | 1.7536 | 1.7852 | H9 | 2.1063 | 3.1186 | 1.0865 | 4.3933 | 2.8348 | 3.2937 | 1.7536 | 5.2237 | | 1.7510 | 4.5675 | 4.6583 | H10 | 2.1333 | 2.6816 | 1.0762 | 4.1669 | 3.1996 | 2.4458 | 1.7852 | 4.7082 | 1.7510 | | 4.6583 | 4.5591 | H11 | 3.1186 | 2.1063 | 4.3933 | 1.0865 | 3.2937 | 2.8348 | 5.2237 | 1.7536 | 4.5675 | 4.6583 | | 1.7510 | H12 | 2.6816 | 2.1333 | 4.1669 | 1.0762 | 2.4458 | 3.1996 | 4.7082 | 1.7852 | 4.6583 | 4.5591 | 1.7510 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
118.268 |
|
C1 |
C2 |
O6 |
118.202 |
| C1 |
C3 |
H7 |
109.989 |
|
C1 |
C3 |
H9 |
108.046 |
| C1 |
C3 |
H10 |
110.797 |
|
C2 |
C1 |
C3 |
118.268 |
| C2 |
C1 |
O5 |
118.202 |
|
C2 |
C4 |
H8 |
109.989 |
| C2 |
C4 |
H11 |
108.046 |
|
C2 |
C4 |
H12 |
110.797 |
| C3 |
C1 |
O5 |
123.518 |
|
C4 |
C2 |
O6 |
123.518 |
| H7 |
C3 |
H9 |
108.035 |
|
H7 |
C3 |
H10 |
111.722 |
| H8 |
C4 |
H11 |
108.035 |
|
H8 |
C4 |
H12 |
111.722 |
| H9 |
C3 |
H10 |
108.123 |
|
H11 |
C4 |
H12 |
108.123 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.287 |
|
|
|
| 2 |
C |
0.287 |
|
|
|
| 3 |
C |
-0.161 |
|
|
|
| 4 |
C |
-0.161 |
|
|
|
| 5 |
O |
-0.392 |
|
|
|
| 6 |
O |
-0.392 |
|
|
|
| 7 |
H |
0.108 |
|
|
|
| 8 |
H |
0.108 |
|
|
|
| 9 |
H |
0.079 |
|
|
|
| 10 |
H |
0.079 |
|
|
|
| 11 |
H |
0.079 |
|
|
|
| 12 |
H |
0.079 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.370 |
8.047 |
0.000 |
| y |
8.047 |
-36.878 |
0.000 |
| z |
0.000 |
0.000 |
-33.632 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.115 |
8.047 |
0.000 |
| y |
8.047 |
-0.377 |
0.000 |
| z |
0.000 |
0.000 |
4.491 |
|
| Polar |
|---|
| 3z2-r2 | 8.983 |
|---|
| x2-y2 | -2.492 |
|---|
| xy | 8.047 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.315 |
-0.022 |
0.000 |
| y |
-0.022 |
7.610 |
0.000 |
| z |
0.000 |
0.000 |
5.420 |
<r2> (average value of r
2) Å
2
| <r2> |
161.323 |
| (<r2>)1/2 |
12.701 |
Jump to
S1C1
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -304.813911 |
| Energy at 298.15K | -304.820359 |
| HF Energy | -304.813911 |
| Nuclear repulsion energy | 228.239252 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3286 |
2978 |
0.00 |
|
|
|
| 2 |
Ag |
3246 |
2942 |
0.00 |
|
|
|
| 3 |
Ag |
3186 |
2888 |
0.00 |
|
|
|
| 4 |
Ag |
2005 |
1818 |
0.00 |
|
|
|
| 5 |
Ag |
1588 |
1439 |
0.00 |
|
|
|
| 6 |
Ag |
1577 |
1430 |
0.00 |
|
|
|
| 7 |
Ag |
1535 |
1391 |
0.00 |
|
|
|
| 8 |
Ag |
1407 |
1275 |
0.00 |
|
|
|
| 9 |
Ag |
1184 |
1073 |
0.00 |
|
|
|
| 10 |
Ag |
1107 |
1004 |
0.00 |
|
|
|
| 11 |
Ag |
743 |
673 |
0.00 |
|
|
|
| 12 |
Ag |
682 |
618 |
0.00 |
|
|
|
| 13 |
Ag |
572 |
518 |
0.00 |
|
|
|
| 14 |
Ag |
386 |
350 |
0.00 |
|
|
|
| 15 |
Ag |
144 |
130 |
0.00 |
|
|
|
| 16 |
Au |
3286 |
2979 |
23.16 |
|
|
|
| 17 |
Au |
3246 |
2942 |
16.05 |
|
|
|
| 18 |
Au |
3186 |
2888 |
4.32 |
|
|
|
| 19 |
Au |
1982 |
1797 |
379.85 |
|
|
|
| 20 |
Au |
1584 |
1436 |
17.81 |
|
|
|
| 21 |
Au |
1577 |
1430 |
27.34 |
|
|
|
| 22 |
Au |
1524 |
1381 |
37.75 |
|
|
|
| 23 |
Au |
1228 |
1114 |
82.81 |
|
|
|
| 24 |
Au |
1062 |
962 |
4.22 |
|
|
|
| 25 |
Au |
982 |
890 |
20.77 |
|
|
|
| 26 |
Au |
589 |
534 |
58.82 |
|
|
|
| 27 |
Au |
395 |
358 |
12.58 |
|
|
|
| 28 |
Au |
255 |
232 |
18.52 |
|
|
|
| 29 |
Au |
142 |
129 |
0.11 |
|
|
|
| 30 |
Au |
51 |
46 |
13.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21868.2 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 19821.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Ci
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.021 |
0.000 |
-0.770 |
| C2 |
-0.021 |
-0.000 |
0.770 |
| C3 |
1.364 |
-0.000 |
-1.429 |
| C4 |
-1.364 |
0.000 |
1.429 |
| O5 |
-1.005 |
-0.000 |
-1.363 |
| O6 |
1.005 |
0.000 |
1.363 |
| H7 |
1.930 |
0.865 |
-1.108 |
| H8 |
-1.930 |
-0.865 |
1.108 |
| H9 |
1.930 |
-0.865 |
-1.107 |
| H10 |
1.253 |
-0.000 |
-2.501 |
| H11 |
-1.930 |
0.865 |
1.107 |
| H12 |
-1.253 |
0.000 |
2.501 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
O5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5415 | 1.4955 | 2.5992 | 1.1854 | 2.3495 | 2.1225 | 2.8429 | 2.1225 | 2.1241 | 2.8427 | 3.5107 |
C2 | 1.5415 | | 2.5992 | 1.4955 | 2.3495 | 1.1854 | 2.8429 | 2.1225 | 2.8427 | 3.5107 | 2.1225 | 2.1241 | C3 | 1.4955 | 2.5992 | | 3.9507 | 2.3703 | 2.8149 | 1.0824 | 4.2464 | 1.0824 | 1.0776 | 4.2463 | 4.7214 | C4 | 2.5992 | 1.4955 | 3.9507 | | 2.8149 | 2.3703 | 4.2464 | 1.0824 | 4.2463 | 4.7214 | 1.0824 | 1.0776 | O5 | 1.1854 | 2.3495 | 2.3703 | 2.8149 | | 3.3875 | 3.0709 | 2.7761 | 3.0710 | 2.5290 | 2.7757 | 3.8718 | O6 | 2.3495 | 1.1854 | 2.8149 | 2.3703 | 3.3875 | | 2.7761 | 3.0709 | 2.7757 | 3.8718 | 3.0710 | 2.5290 | H7 | 2.1225 | 2.8429 | 1.0824 | 4.2464 | 3.0709 | 2.7761 | | 4.7748 | 1.7293 | 1.7741 | 4.4505 | 4.8888 | H8 | 2.8429 | 2.1225 | 4.2464 | 1.0824 | 2.7761 | 3.0709 | 4.7748 | | 4.4505 | 4.8888 | 1.7293 | 1.7741 | H9 | 2.1225 | 2.8427 | 1.0824 | 4.2463 | 3.0710 | 2.7757 | 1.7293 | 4.4505 | | 1.7741 | 4.7745 | 4.8887 | H10 | 2.1241 | 3.5107 | 1.0776 | 4.7214 | 2.5290 | 3.8718 | 1.7741 | 4.8888 | 1.7741 | | 4.8887 | 5.5944 | H11 | 2.8427 | 2.1225 | 4.2463 | 1.0824 | 2.7757 | 3.0710 | 4.4505 | 1.7293 | 4.7745 | 4.8887 | | 1.7741 | H12 | 3.5107 | 2.1241 | 4.7214 | 1.0776 | 3.8718 | 2.5290 | 4.8888 | 1.7741 | 4.8887 | 5.5944 | 1.7741 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
117.703 |
|
C1 |
C2 |
O6 |
118.415 |
| C1 |
C3 |
H7 |
109.808 |
|
C1 |
C3 |
H9 |
109.807 |
| C1 |
C3 |
H10 |
110.225 |
|
C2 |
C1 |
C3 |
117.703 |
| C2 |
C1 |
O5 |
118.415 |
|
C2 |
C4 |
H8 |
109.808 |
| C2 |
C4 |
H11 |
109.807 |
|
C2 |
C4 |
H12 |
110.225 |
| C3 |
C1 |
O5 |
123.882 |
|
C4 |
C2 |
O6 |
123.882 |
| H7 |
C3 |
H9 |
106.033 |
|
H7 |
C3 |
H10 |
110.442 |
| H8 |
C4 |
H11 |
106.033 |
|
H8 |
C4 |
H12 |
110.442 |
| H9 |
C3 |
H10 |
110.442 |
|
H11 |
C4 |
H12 |
110.442 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.287 |
|
|
|
| 2 |
C |
0.287 |
|
|
|
| 3 |
C |
-0.161 |
|
|
|
| 4 |
C |
-0.161 |
|
|
|
| 5 |
O |
-0.392 |
|
|
|
| 6 |
O |
-0.392 |
|
|
|
| 7 |
H |
0.079 |
|
|
|
| 8 |
H |
0.079 |
|
|
|
| 9 |
H |
0.079 |
|
|
|
| 10 |
H |
0.108 |
|
|
|
| 11 |
H |
0.079 |
|
|
|
| 12 |
H |
0.108 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.283 |
-0.001 |
-8.099 |
| y |
-0.001 |
-33.632 |
-0.000 |
| z |
-8.099 |
-0.000 |
-38.966 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.984 |
-0.001 |
-8.099 |
| y |
-0.001 |
4.493 |
-0.000 |
| z |
-8.099 |
-0.000 |
-3.509 |
|
| Polar |
|---|
| 3z2-r2 | -7.018 |
|---|
| x2-y2 | -3.651 |
|---|
| xy | -0.001 |
|---|
| xz | -8.099 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.298 |
0.000 |
0.111 |
| y |
0.000 |
5.420 |
0.000 |
| z |
0.111 |
0.000 |
7.628 |
<r2> (average value of r
2) Å
2
| <r2> |
161.328 |
| (<r2>)1/2 |
12.702 |
