Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -153.515359 |
| Energy at 298.15K | -153.520066 |
| HF Energy | -153.515359 |
| Nuclear repulsion energy | 75.399126 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
4170 |
3780 |
47.93 |
|
|
|
| 2 |
A' |
3272 |
2966 |
11.27 |
|
|
|
| 3 |
A' |
3145 |
2851 |
72.71 |
|
|
|
| 4 |
A' |
1648 |
1494 |
0.61 |
|
|
|
| 5 |
A' |
1601 |
1451 |
8.07 |
|
|
|
| 6 |
A' |
1550 |
1405 |
6.78 |
|
|
|
| 7 |
A' |
1325 |
1201 |
87.37 |
|
|
|
| 8 |
A' |
1155 |
1046 |
128.50 |
|
|
|
| 9 |
A' |
1059 |
960 |
2.87 |
|
|
|
| 10 |
A' |
642 |
582 |
12.71 |
|
|
|
| 11 |
A' |
401 |
364 |
34.38 |
|
|
|
| 12 |
A" |
3376 |
3060 |
17.75 |
|
|
|
| 13 |
A" |
3171 |
2875 |
64.30 |
|
|
|
| 14 |
A" |
1403 |
1272 |
1.25 |
|
|
|
| 15 |
A" |
1270 |
1151 |
1.54 |
|
|
|
| 16 |
A" |
861 |
780 |
0.62 |
|
|
|
| 17 |
A" |
277 |
251 |
135.56 |
|
|
|
| 18 |
A" |
72 |
65 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15198.3 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 13775.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
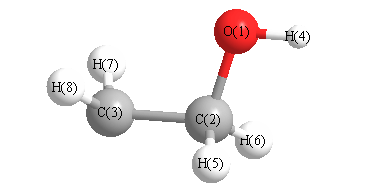
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.106 |
-0.342 |
0.000 |
| C2 |
0.000 |
0.533 |
0.000 |
| C3 |
1.258 |
-0.268 |
0.000 |
| H4 |
-1.905 |
0.153 |
0.000 |
| H5 |
-0.031 |
1.172 |
0.878 |
| H6 |
-0.031 |
1.172 |
-0.878 |
| H7 |
1.632 |
-0.676 |
-0.919 |
| H8 |
1.632 |
-0.676 |
0.919 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4103 | 2.3657 | 0.9392 | 2.0537 | 2.0537 | 2.9077 | 2.9077 |
C2 | 1.4103 | | 1.4916 | 1.9423 | 1.0859 | 1.0859 | 2.2297 | 2.2297 | C3 | 2.3657 | 1.4916 | | 3.1908 | 2.1222 | 2.1222 | 1.0732 | 1.0732 | H4 | 0.9392 | 1.9423 | 3.1908 | | 2.3067 | 2.3067 | 3.7471 | 3.7471 | H5 | 2.0537 | 1.0859 | 2.1222 | 2.3067 | | 1.7554 | 3.0674 | 2.4863 | H6 | 2.0537 | 1.0859 | 2.1222 | 2.3067 | 1.7554 | | 2.4863 | 3.0674 | H7 | 2.9077 | 2.2297 | 1.0732 | 3.7471 | 3.0674 | 2.4863 | | 1.8385 | H8 | 2.9077 | 2.2297 | 1.0732 | 3.7471 | 2.4863 | 3.0674 | 1.8385 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
109.187 |
|
O1 |
C2 |
H5 |
110.041 |
| O1 |
C2 |
H6 |
110.041 |
|
C2 |
O1 |
H4 |
109.893 |
| C2 |
C3 |
H7 |
119.883 |
|
C2 |
C3 |
H8 |
119.883 |
| C3 |
C2 |
H5 |
109.848 |
|
C3 |
C2 |
H6 |
109.848 |
| H5 |
C2 |
H6 |
107.861 |
|
H7 |
C3 |
H8 |
117.868 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.410 |
|
|
|
| 2 |
C |
0.171 |
|
|
|
| 3 |
C |
-0.147 |
|
|
|
| 4 |
H |
0.199 |
|
|
|
| 5 |
H |
0.031 |
|
|
|
| 6 |
H |
0.031 |
|
|
|
| 7 |
H |
0.062 |
|
|
|
| 8 |
H |
0.062 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.245 |
1.543 |
0.000 |
1.562 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.139 |
-2.907 |
0.000 |
| y |
-2.907 |
-20.087 |
0.000 |
| z |
0.000 |
0.000 |
-18.861 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.335 |
-2.907 |
0.000 |
| y |
-2.907 |
-2.087 |
0.000 |
| z |
0.000 |
0.000 |
-0.248 |
|
| Polar |
|---|
| 3z2-r2 | -0.495 |
|---|
| x2-y2 | 2.948 |
|---|
| xy | -2.907 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.524 |
-0.272 |
0.000 |
| y |
-0.272 |
3.823 |
0.000 |
| z |
0.000 |
0.000 |
3.783 |
<r2> (average value of r
2) Å
2
| <r2> |
49.916 |
| (<r2>)1/2 |
7.065 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -153.516360 |
| Energy at 298.15K | -153.521156 |
| HF Energy | -153.516360 |
| Nuclear repulsion energy | 75.611363 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4184 |
3793 |
56.89 |
|
|
|
| 2 |
A |
3391 |
3074 |
10.69 |
|
|
|
| 3 |
A |
3281 |
2974 |
8.88 |
|
|
|
| 4 |
A |
3130 |
2837 |
70.39 |
|
|
|
| 5 |
A |
3081 |
2793 |
78.87 |
|
|
|
| 6 |
A |
1634 |
1481 |
3.08 |
|
|
|
| 7 |
A |
1595 |
1446 |
11.16 |
|
|
|
| 8 |
A |
1543 |
1399 |
1.77 |
|
|
|
| 9 |
A |
1376 |
1247 |
63.05 |
|
|
|
| 10 |
A |
1346 |
1220 |
37.27 |
|
|
|
| 11 |
A |
1211 |
1098 |
40.66 |
|
|
|
| 12 |
A |
1142 |
1035 |
47.84 |
|
|
|
| 13 |
A |
1043 |
945 |
5.67 |
|
|
|
| 14 |
A |
940 |
852 |
15.34 |
|
|
|
| 15 |
A |
482 |
437 |
18.42 |
|
|
|
| 16 |
A |
421 |
381 |
39.90 |
|
|
|
| 17 |
A |
283 |
256 |
115.06 |
|
|
|
| 18 |
A |
163 |
148 |
16.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15122.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 13707.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.100 |
-0.376 |
-0.056 |
| C2 |
-0.019 |
0.518 |
0.038 |
| C3 |
1.253 |
-0.249 |
-0.028 |
| H4 |
-1.905 |
0.084 |
0.091 |
| H5 |
-0.087 |
1.074 |
0.973 |
| H6 |
-0.065 |
1.246 |
-0.767 |
| H7 |
2.180 |
0.261 |
-0.200 |
| H8 |
1.269 |
-1.271 |
0.292 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4058 | 2.3572 | 0.9378 | 2.0463 | 2.0515 | 3.3450 | 2.5567 |
C2 | 1.4058 | | 1.4867 | 1.9358 | 1.0905 | 1.0870 | 2.2268 | 2.2185 | C3 | 2.3572 | 1.4867 | | 3.1777 | 2.1329 | 2.1261 | 1.0719 | 1.0710 | H4 | 0.9378 | 1.9358 | 3.1777 | | 2.2502 | 2.3394 | 4.0992 | 3.4566 | H5 | 2.0463 | 1.0905 | 2.1329 | 2.2502 | | 1.7494 | 2.6793 | 2.7929 | H6 | 2.0515 | 1.0870 | 2.1261 | 2.3394 | 1.7494 | | 2.5167 | 3.0395 | H7 | 3.3450 | 2.2268 | 1.0719 | 4.0992 | 2.6793 | 2.5167 | | 1.8492 | H8 | 2.5567 | 2.2185 | 1.0710 | 3.4566 | 2.7929 | 3.0395 | 1.8492 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
109.133 |
|
O1 |
C2 |
H5 |
109.468 |
| O1 |
C2 |
H6 |
110.111 |
|
C2 |
O1 |
H4 |
109.767 |
| C2 |
C3 |
H7 |
120.119 |
|
C2 |
C3 |
H8 |
119.422 |
| C3 |
C2 |
H5 |
110.769 |
|
C3 |
C2 |
H6 |
110.431 |
| H5 |
C2 |
H6 |
106.907 |
|
H7 |
C3 |
H8 |
119.290 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.410 |
|
|
|
| 2 |
C |
0.176 |
|
|
|
| 3 |
C |
-0.167 |
|
|
|
| 4 |
H |
0.198 |
|
|
|
| 5 |
H |
0.033 |
|
|
|
| 6 |
H |
0.034 |
|
|
|
| 7 |
H |
0.058 |
|
|
|
| 8 |
H |
0.078 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.239 |
1.729 |
0.418 |
1.795 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.097 |
-1.829 |
-0.298 |
| y |
-1.829 |
-19.472 |
-0.477 |
| z |
-0.298 |
-0.477 |
-20.319 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.799 |
-1.829 |
-0.298 |
| y |
-1.829 |
-1.264 |
-0.477 |
| z |
-0.298 |
-0.477 |
-2.535 |
|
| Polar |
|---|
| 3z2-r2 | -5.069 |
|---|
| x2-y2 | 3.375 |
|---|
| xy | -1.829 |
|---|
| xz | -0.298 |
|---|
| yz | -0.477 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.575 |
-0.115 |
-0.064 |
| y |
-0.115 |
4.046 |
-0.038 |
| z |
-0.064 |
-0.038 |
3.478 |
<r2> (average value of r
2) Å
2
| <r2> |
49.914 |
| (<r2>)1/2 |
7.065 |
