Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -217.207440 |
| Energy at 298.15K | -217.215474 |
| HF Energy | -217.207440 |
| Nuclear repulsion energy | 128.989708 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3228 |
2926 |
48.59 |
|
|
|
| 2 |
A' |
3187 |
2889 |
59.93 |
|
|
|
| 3 |
A' |
3172 |
2875 |
15.83 |
|
|
|
| 4 |
A' |
3158 |
2863 |
26.87 |
|
|
|
| 5 |
A' |
1646 |
1492 |
2.14 |
|
|
|
| 6 |
A' |
1629 |
1477 |
5.15 |
|
|
|
| 7 |
A' |
1611 |
1460 |
0.86 |
|
|
|
| 8 |
A' |
1565 |
1419 |
14.82 |
|
|
|
| 9 |
A' |
1539 |
1395 |
2.04 |
|
|
|
| 10 |
A' |
1454 |
1318 |
2.29 |
|
|
|
| 11 |
A' |
1224 |
1109 |
5.01 |
|
|
|
| 12 |
A' |
1162 |
1053 |
137.27 |
|
|
|
| 13 |
A' |
1096 |
994 |
10.68 |
|
|
|
| 14 |
A' |
961 |
871 |
6.93 |
|
|
|
| 15 |
A' |
482 |
437 |
7.10 |
|
|
|
| 16 |
A' |
292 |
264 |
4.00 |
|
|
|
| 17 |
A" |
3239 |
2936 |
113.90 |
|
|
|
| 18 |
A" |
3213 |
2912 |
11.89 |
|
|
|
| 19 |
A" |
3192 |
2894 |
0.82 |
|
|
|
| 20 |
A" |
1616 |
1465 |
6.11 |
|
|
|
| 21 |
A" |
1428 |
1294 |
0.01 |
|
|
|
| 22 |
A" |
1382 |
1252 |
1.20 |
|
|
|
| 23 |
A" |
1299 |
1178 |
2.48 |
|
|
|
| 24 |
A" |
963 |
873 |
1.33 |
|
|
|
| 25 |
A" |
816 |
740 |
0.98 |
|
|
|
| 26 |
A" |
243 |
220 |
0.01 |
|
|
|
| 27 |
A" |
139 |
126 |
3.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22467.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 20364.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.132 |
-0.784 |
0.000 |
| C2 |
0.000 |
0.722 |
0.000 |
| C3 |
-1.458 |
1.166 |
0.000 |
| F4 |
1.453 |
-1.140 |
0.000 |
| H5 |
-0.321 |
-1.223 |
0.880 |
| H6 |
-0.321 |
-1.223 |
-0.880 |
| H7 |
0.511 |
1.115 |
-0.872 |
| H8 |
0.511 |
1.115 |
0.872 |
| H9 |
-1.529 |
2.246 |
0.000 |
| H10 |
-1.982 |
0.801 |
-0.877 |
| H11 |
-1.982 |
0.801 |
0.877 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5117 | 2.5164 | 1.3679 | 1.0827 | 1.0827 | 2.1235 | 2.1235 | 3.4559 | 2.7840 | 2.7840 |
C2 | 1.5117 | | 1.5245 | 2.3615 | 2.1585 | 2.1585 | 1.0842 | 1.0842 | 2.1595 | 2.1689 | 2.1689 | C3 | 2.5164 | 1.5245 | | 3.7137 | 2.7885 | 2.7885 | 2.1544 | 2.1544 | 1.0824 | 1.0845 | 1.0845 | F4 | 1.3679 | 2.3615 | 3.7137 | | 1.9817 | 1.9817 | 2.5942 | 2.5942 | 4.5122 | 4.0417 | 4.0417 | H5 | 1.0827 | 2.1585 | 2.7885 | 1.9817 | | 1.7608 | 3.0374 | 2.4811 | 3.7776 | 3.1533 | 2.6185 | H6 | 1.0827 | 2.1585 | 2.7885 | 1.9817 | 1.7608 | | 2.4811 | 3.0374 | 3.7776 | 2.6185 | 3.1533 | H7 | 2.1235 | 1.0842 | 2.1544 | 2.5942 | 3.0374 | 2.4811 | | 1.7433 | 2.4910 | 2.5132 | 3.0614 | H8 | 2.1235 | 1.0842 | 2.1544 | 2.5942 | 2.4811 | 3.0374 | 1.7433 | | 2.4910 | 3.0614 | 2.5132 | H9 | 3.4559 | 2.1595 | 1.0824 | 4.5122 | 3.7776 | 3.7776 | 2.4910 | 2.4910 | | 1.7497 | 1.7497 | H10 | 2.7840 | 2.1689 | 1.0845 | 4.0417 | 3.1533 | 2.6185 | 2.5132 | 3.0614 | 1.7497 | | 1.7530 | H11 | 2.7840 | 2.1689 | 1.0845 | 4.0417 | 2.6185 | 3.1533 | 3.0614 | 2.5132 | 1.7497 | 1.7530 | |
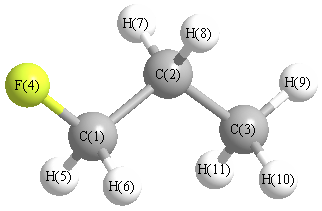 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.949 |
|
C1 |
C2 |
H7 |
108.655 |
| C1 |
C2 |
H8 |
108.655 |
|
C2 |
C1 |
F4 |
110.080 |
| C2 |
C1 |
H5 |
111.537 |
|
C2 |
C1 |
H6 |
111.537 |
| C2 |
C3 |
H9 |
110.726 |
|
C2 |
C3 |
H10 |
111.352 |
| C2 |
C3 |
H11 |
111.352 |
|
C3 |
C2 |
H7 |
110.209 |
| C3 |
C2 |
H8 |
110.209 |
|
F4 |
C1 |
H5 |
107.352 |
| F4 |
C1 |
H6 |
107.352 |
|
H5 |
C1 |
H6 |
108.804 |
| H7 |
C2 |
H8 |
107.018 |
|
H9 |
C3 |
H10 |
107.702 |
| H9 |
C3 |
H11 |
107.702 |
|
H10 |
C3 |
H11 |
107.839 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.233 |
|
|
|
| 2 |
C |
-0.026 |
|
|
|
| 3 |
C |
-0.193 |
|
|
|
| 4 |
F |
-0.312 |
|
|
|
| 5 |
H |
0.029 |
|
|
|
| 6 |
H |
0.029 |
|
|
|
| 7 |
H |
0.035 |
|
|
|
| 8 |
H |
0.035 |
|
|
|
| 9 |
H |
0.066 |
|
|
|
| 10 |
H |
0.052 |
|
|
|
| 11 |
H |
0.052 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.881 |
0.812 |
0.000 |
2.049 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.879 |
2.428 |
0.000 |
| y |
2.428 |
-25.986 |
0.000 |
| z |
0.000 |
0.000 |
-24.610 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.581 |
2.428 |
0.000 |
| y |
2.428 |
0.758 |
0.000 |
| z |
0.000 |
0.000 |
2.823 |
|
| Polar |
|---|
| 3z2-r2 | 5.646 |
|---|
| x2-y2 | -2.892 |
|---|
| xy | 2.428 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.627 |
-0.296 |
0.000 |
| y |
-0.296 |
5.523 |
0.000 |
| z |
0.000 |
0.000 |
5.048 |
<r2> (average value of r
2) Å
2
| <r2> |
102.440 |
| (<r2>)1/2 |
10.121 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -217.207479 |
| Energy at 298.15K | |
| HF Energy | -217.207479 |
| Nuclear repulsion energy | 131.168102 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3238 |
2935 |
52.62 |
37.71 |
0.70 |
0.82 |
| 2 |
A |
3235 |
2932 |
66.96 |
61.17 |
0.56 |
0.72 |
| 3 |
A |
3218 |
2916 |
60.30 |
45.09 |
0.68 |
0.81 |
| 4 |
A |
3196 |
2897 |
41.28 |
118.35 |
0.11 |
0.20 |
| 5 |
A |
3183 |
2885 |
11.73 |
164.19 |
0.33 |
0.50 |
| 6 |
A |
3162 |
2866 |
17.95 |
188.51 |
0.03 |
0.06 |
| 7 |
A |
3157 |
2861 |
35.94 |
30.97 |
0.73 |
0.84 |
| 8 |
A |
1644 |
1490 |
0.97 |
3.02 |
0.74 |
0.85 |
| 9 |
A |
1623 |
1471 |
8.20 |
2.39 |
0.75 |
0.85 |
| 10 |
A |
1613 |
1462 |
6.27 |
8.19 |
0.75 |
0.86 |
| 11 |
A |
1597 |
1448 |
1.28 |
9.63 |
0.73 |
0.84 |
| 12 |
A |
1556 |
1411 |
13.92 |
1.23 |
0.07 |
0.13 |
| 13 |
A |
1541 |
1397 |
2.87 |
0.45 |
0.75 |
0.86 |
| 14 |
A |
1501 |
1360 |
0.99 |
0.78 |
0.75 |
0.85 |
| 15 |
A |
1410 |
1278 |
1.91 |
8.04 |
0.67 |
0.80 |
| 16 |
A |
1385 |
1256 |
2.20 |
3.21 |
0.75 |
0.86 |
| 17 |
A |
1280 |
1160 |
2.27 |
0.90 |
0.04 |
0.08 |
| 18 |
A |
1222 |
1107 |
32.13 |
2.30 |
0.67 |
0.80 |
| 19 |
A |
1166 |
1057 |
62.19 |
4.44 |
0.68 |
0.81 |
| 20 |
A |
1057 |
958 |
42.30 |
4.60 |
0.73 |
0.84 |
| 21 |
A |
993 |
900 |
3.93 |
0.51 |
0.74 |
0.85 |
| 22 |
A |
921 |
835 |
3.91 |
12.51 |
0.13 |
0.23 |
| 23 |
A |
829 |
751 |
0.98 |
0.91 |
0.28 |
0.44 |
| 24 |
A |
514 |
466 |
5.45 |
0.40 |
0.73 |
0.84 |
| 25 |
A |
337 |
306 |
1.24 |
0.27 |
0.20 |
0.34 |
| 26 |
A |
235 |
213 |
1.64 |
0.05 |
0.36 |
0.53 |
| 27 |
A |
150 |
136 |
2.75 |
0.10 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 22480.1 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 20376.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.802 |
0.495 |
0.300 |
| C2 |
-0.585 |
0.649 |
-0.282 |
| C3 |
-1.539 |
-0.472 |
0.117 |
| F4 |
1.394 |
-0.646 |
-0.173 |
| H5 |
1.438 |
1.326 |
0.023 |
| H6 |
0.773 |
0.419 |
1.380 |
| H7 |
-0.503 |
0.702 |
-1.362 |
| H8 |
-0.974 |
1.606 |
0.052 |
| H9 |
-2.517 |
-0.319 |
-0.323 |
| H10 |
-1.169 |
-1.433 |
-0.215 |
| H11 |
-1.662 |
-0.515 |
1.194 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5119 | 2.5393 | 1.3696 | 1.0820 | 1.0832 | 2.1234 | 2.1101 | 3.4735 | 2.8049 | 2.8093 |
C2 | 1.5119 | | 1.5248 | 2.3675 | 2.1547 | 2.1582 | 1.0848 | 1.0862 | 2.1610 | 2.1635 | 2.1661 | C3 | 2.5393 | 1.5248 | | 2.9522 | 3.4786 | 2.7810 | 2.1540 | 2.1546 | 1.0832 | 1.0822 | 1.0849 | F4 | 1.3696 | 2.3675 | 2.9522 | | 1.9823 | 1.9829 | 2.6140 | 3.2762 | 3.9273 | 2.6813 | 3.3506 | H5 | 1.0820 | 2.1547 | 3.4786 | 1.9823 | | 1.7616 | 2.4649 | 2.4285 | 4.2969 | 3.8031 | 3.7904 | H6 | 1.0832 | 2.1582 | 2.7810 | 1.9829 | 1.7616 | | 3.0377 | 2.4950 | 3.7772 | 3.1220 | 2.6149 | H7 | 2.1234 | 1.0848 | 2.1540 | 2.6140 | 2.4649 | 3.0377 | | 1.7433 | 2.4855 | 2.5141 | 3.0592 | H8 | 2.1101 | 1.0862 | 2.1546 | 3.2762 | 2.4285 | 2.4950 | 1.7433 | | 2.4953 | 3.0575 | 2.5052 | H9 | 3.4735 | 2.1610 | 1.0832 | 3.9273 | 4.2969 | 3.7772 | 2.4855 | 2.4953 | | 1.7523 | 1.7520 | H10 | 2.8049 | 2.1635 | 1.0822 | 2.6813 | 3.8031 | 3.1220 | 2.5141 | 3.0575 | 1.7523 | | 1.7528 | H11 | 2.8093 | 2.1661 | 1.0849 | 3.3506 | 3.7904 | 2.6149 | 3.0592 | 2.5052 | 1.7520 | 1.7528 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.484 |
|
C1 |
C2 |
H7 |
108.600 |
| C1 |
C2 |
H8 |
107.482 |
|
C2 |
C1 |
F4 |
110.403 |
| C2 |
C1 |
H5 |
111.257 |
|
C2 |
C1 |
H6 |
111.468 |
| C2 |
C3 |
H9 |
110.774 |
|
C2 |
C3 |
H10 |
111.032 |
| C2 |
C3 |
H11 |
111.083 |
|
C3 |
C2 |
H7 |
110.117 |
| C3 |
C2 |
H8 |
110.080 |
|
F4 |
C1 |
H5 |
107.333 |
| F4 |
C1 |
H6 |
107.311 |
|
H5 |
C1 |
H6 |
108.899 |
| H7 |
C2 |
H8 |
106.827 |
|
H9 |
C3 |
H10 |
108.035 |
| H9 |
C3 |
H11 |
107.816 |
|
H10 |
C3 |
H11 |
107.967 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.227 |
|
|
|
| 2 |
C |
-0.040 |
|
|
|
| 3 |
C |
-0.178 |
|
|
|
| 4 |
F |
-0.316 |
|
|
|
| 5 |
H |
0.042 |
|
|
|
| 6 |
H |
0.027 |
|
|
|
| 7 |
H |
0.036 |
|
|
|
| 8 |
H |
0.025 |
|
|
|
| 9 |
H |
0.059 |
|
|
|
| 10 |
H |
0.074 |
|
|
|
| 11 |
H |
0.044 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.045 |
1.559 |
0.524 |
1.949 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.250 |
1.895 |
0.778 |
| y |
1.895 |
-25.640 |
-0.033 |
| z |
0.778 |
-0.033 |
-24.812 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.024 |
1.895 |
0.778 |
| y |
1.895 |
0.391 |
-0.033 |
| z |
0.778 |
-0.033 |
1.633 |
|
| Polar |
|---|
| 3z2-r2 | 3.266 |
|---|
| x2-y2 | -1.610 |
|---|
| xy | 1.895 |
|---|
| xz | 0.778 |
|---|
| yz | -0.033 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.707 |
0.235 |
0.040 |
| y |
0.235 |
5.351 |
-0.030 |
| z |
0.040 |
-0.030 |
5.153 |
<r2> (average value of r
2) Å
2
| <r2> |
91.342 |
| (<r2>)1/2 |
9.557 |
