Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP/Def2TZVPP
Geometric Data calculated at B2PLYP/Def2TZVPP
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -218.294112 |
| Energy at 298.15K | |
| HF Energy | -218.042967 |
| Nuclear repulsion energy | 130.434176 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3141 |
3141 |
21.99 |
|
|
|
| 2 |
A |
3126 |
3126 |
48.61 |
|
|
|
| 3 |
A |
3116 |
3116 |
35.04 |
|
|
|
| 4 |
A |
3091 |
3091 |
16.10 |
|
|
|
| 5 |
A |
3062 |
3062 |
31.79 |
|
|
|
| 6 |
A |
3057 |
3057 |
24.66 |
|
|
|
| 7 |
A |
3051 |
3051 |
15.36 |
|
|
|
| 8 |
A |
1528 |
1528 |
1.79 |
|
|
|
| 9 |
A |
1522 |
1522 |
7.75 |
|
|
|
| 10 |
A |
1510 |
1510 |
6.03 |
|
|
|
| 11 |
A |
1487 |
1487 |
2.52 |
|
|
|
| 12 |
A |
1438 |
1438 |
9.18 |
|
|
|
| 13 |
A |
1421 |
1421 |
4.70 |
|
|
|
| 14 |
A |
1392 |
1392 |
0.31 |
|
|
|
| 15 |
A |
1313 |
1313 |
1.21 |
|
|
|
| 16 |
A |
1286 |
1286 |
1.12 |
|
|
|
| 17 |
A |
1186 |
1186 |
1.52 |
|
|
|
| 18 |
A |
1131 |
1131 |
8.39 |
|
|
|
| 19 |
A |
1090 |
1090 |
45.50 |
|
|
|
| 20 |
A |
982 |
982 |
53.98 |
|
|
|
| 21 |
A |
929 |
929 |
4.20 |
|
|
|
| 22 |
A |
881 |
881 |
4.29 |
|
|
|
| 23 |
A |
774 |
774 |
0.70 |
|
|
|
| 24 |
A |
480 |
480 |
4.24 |
|
|
|
| 25 |
A |
317 |
317 |
1.13 |
|
|
|
| 26 |
A |
216 |
216 |
1.54 |
|
|
|
| 27 |
A |
141 |
141 |
2.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21333.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 21333.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.791 |
0.513 |
0.301 |
| C2 |
-0.594 |
0.651 |
-0.287 |
| C3 |
-1.531 |
-0.482 |
0.118 |
| F4 |
1.402 |
-0.652 |
-0.172 |
| H5 |
1.432 |
1.348 |
0.020 |
| H6 |
0.757 |
0.437 |
1.389 |
| H7 |
-0.507 |
0.702 |
-1.373 |
| H8 |
-0.995 |
1.612 |
0.043 |
| H9 |
-2.516 |
-0.353 |
-0.326 |
| H10 |
-1.138 |
-1.444 |
-0.205 |
| H11 |
-1.655 |
-0.517 |
1.201 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5111 | 2.5330 | 1.3979 | 1.0894 | 1.0908 | 2.1274 | 2.1134 | 3.4760 | 2.7937 | 2.8025 |
C2 | 1.5111 | | 1.5250 | 2.3864 | 2.1647 | 2.1632 | 1.0909 | 1.0926 | 2.1690 | 2.1652 | 2.1689 | C3 | 2.5330 | 1.5250 | | 2.9523 | 3.4840 | 2.7738 | 2.1621 | 2.1626 | 1.0886 | 1.0880 | 1.0904 | F4 | 1.3979 | 2.3864 | 2.9523 | | 2.0095 | 2.0093 | 2.6312 | 3.3045 | 3.9328 | 2.6605 | 3.3538 | H5 | 1.0894 | 2.1647 | 3.4840 | 2.0095 | | 1.7769 | 2.4742 | 2.4420 | 4.3132 | 3.8009 | 3.7950 | H6 | 1.0908 | 2.1632 | 2.7738 | 2.0093 | 1.7769 | | 3.0493 | 2.5027 | 3.7790 | 3.1092 | 2.6006 | H7 | 2.1274 | 1.0909 | 2.1621 | 2.6312 | 2.4742 | 3.0493 | | 1.7527 | 2.4991 | 2.5229 | 3.0710 | H8 | 2.1134 | 1.0926 | 2.1626 | 3.3045 | 2.4420 | 2.5027 | 1.7527 | | 2.5120 | 3.0687 | 2.5117 | H9 | 3.4760 | 2.1690 | 1.0886 | 3.9328 | 4.3132 | 3.7790 | 2.4991 | 2.5120 | | 1.7618 | 1.7612 | H10 | 2.7937 | 2.1652 | 1.0880 | 2.6605 | 3.8009 | 3.1092 | 2.5229 | 3.0687 | 1.7618 | | 1.7616 | H11 | 2.8025 | 2.1689 | 1.0904 | 3.3538 | 3.7950 | 2.6006 | 3.0710 | 2.5117 | 1.7612 | 1.7616 | |
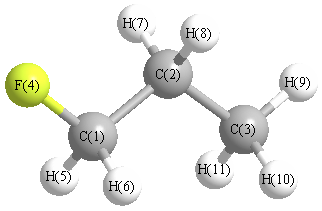 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.079 |
|
C1 |
C2 |
H7 |
108.611 |
| C1 |
C2 |
H8 |
107.426 |
|
C2 |
C1 |
F4 |
110.173 |
| C2 |
C1 |
H5 |
111.663 |
|
C2 |
C1 |
H6 |
111.464 |
| C2 |
C3 |
H9 |
111.075 |
|
C2 |
C3 |
H10 |
110.808 |
| C2 |
C3 |
H11 |
110.959 |
|
C3 |
C2 |
H7 |
110.388 |
| C3 |
C2 |
H8 |
110.321 |
|
F4 |
C1 |
H5 |
107.127 |
| F4 |
C1 |
H6 |
107.030 |
|
H5 |
C1 |
H6 |
109.184 |
| H7 |
C2 |
H8 |
106.774 |
|
H9 |
C3 |
H10 |
108.084 |
| H9 |
C3 |
H11 |
107.851 |
|
H10 |
C3 |
H11 |
107.932 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.093 |
|
|
|
| 2 |
C |
-0.100 |
|
|
|
| 3 |
C |
-0.249 |
|
|
|
| 4 |
F |
-0.264 |
|
|
|
| 5 |
H |
0.080 |
|
|
|
| 6 |
H |
0.065 |
|
|
|
| 7 |
H |
0.068 |
|
|
|
| 8 |
H |
0.054 |
|
|
|
| 9 |
H |
0.084 |
|
|
|
| 10 |
H |
0.098 |
|
|
|
| 11 |
H |
0.070 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.031 |
1.485 |
0.473 |
1.868 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.410 |
1.747 |
0.701 |
| y |
1.747 |
-25.803 |
0.001 |
| z |
0.701 |
0.001 |
-25.104 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.957 |
1.747 |
0.701 |
| y |
1.747 |
0.454 |
0.001 |
| z |
0.701 |
0.001 |
1.503 |
|
| Polar |
|---|
| 3z2-r2 | 3.005 |
|---|
| x2-y2 | -1.607 |
|---|
| xy | 1.747 |
|---|
| xz | 0.701 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.124 |
0.246 |
0.051 |
| y |
0.246 |
5.710 |
-0.028 |
| z |
0.051 |
-0.028 |
5.447 |
<r2> (average value of r
2) Å
2
| <r2> |
91.882 |
| (<r2>)1/2 |
9.586 |
