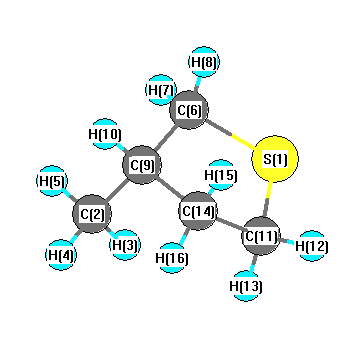
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3253 |
2949 |
26.53 |
|
|
|
| 2 |
A |
3247 |
2943 |
19.35 |
|
|
|
| 3 |
A |
3229 |
2927 |
58.34 |
|
|
|
| 4 |
A |
3216 |
2915 |
90.50 |
|
|
|
| 5 |
A |
3213 |
2912 |
9.82 |
|
|
|
| 6 |
A |
3204 |
2904 |
46.38 |
|
|
|
| 7 |
A |
3199 |
2900 |
13.34 |
|
|
|
| 8 |
A |
3176 |
2879 |
44.70 |
|
|
|
| 9 |
A |
3161 |
2865 |
33.92 |
|
|
|
| 10 |
A |
3158 |
2863 |
10.28 |
|
|
|
| 11 |
A |
1635 |
1482 |
2.09 |
|
|
|
| 12 |
A |
1629 |
1477 |
3.51 |
|
|
|
| 13 |
A |
1615 |
1463 |
3.58 |
|
|
|
| 14 |
A |
1612 |
1461 |
6.71 |
|
|
|
| 15 |
A |
1607 |
1456 |
1.88 |
|
|
|
| 16 |
A |
1542 |
1398 |
7.05 |
|
|
|
| 17 |
A |
1493 |
1353 |
4.78 |
|
|
|
| 18 |
A |
1472 |
1335 |
1.14 |
|
|
|
| 19 |
A |
1446 |
1311 |
4.10 |
|
|
|
| 20 |
A |
1416 |
1283 |
5.38 |
|
|
|
| 21 |
A |
1378 |
1249 |
13.89 |
|
|
|
| 22 |
A |
1339 |
1214 |
2.17 |
|
|
|
| 23 |
A |
1291 |
1171 |
2.46 |
|
|
|
| 24 |
A |
1243 |
1127 |
0.11 |
|
|
|
| 25 |
A |
1167 |
1057 |
0.94 |
|
|
|
| 26 |
A |
1155 |
1047 |
0.24 |
|
|
|
| 27 |
A |
1117 |
1013 |
0.49 |
|
|
|
| 28 |
A |
1062 |
962 |
0.82 |
|
|
|
| 29 |
A |
1034 |
937 |
0.12 |
|
|
|
| 30 |
A |
995 |
902 |
0.81 |
|
|
|
| 31 |
A |
972 |
881 |
2.09 |
|
|
|
| 32 |
A |
917 |
831 |
0.88 |
|
|
|
| 33 |
A |
848 |
768 |
1.75 |
|
|
|
| 34 |
A |
779 |
706 |
4.47 |
|
|
|
| 35 |
A |
748 |
678 |
3.78 |
|
|
|
| 36 |
A |
658 |
596 |
0.49 |
|
|
|
| 37 |
A |
533 |
483 |
0.08 |
|
|
|
| 38 |
A |
401 |
364 |
0.07 |
|
|
|
| 39 |
A |
377 |
342 |
0.18 |
|
|
|
| 40 |
A |
243 |
221 |
0.06 |
|
|
|
| 41 |
A |
204 |
185 |
0.24 |
|
|
|
| 42 |
A |
118 |
107 |
2.77 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33050.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 29957.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.127 |
|
|
|
| 2 |
C |
-0.246 |
|
|
|
| 3 |
H |
0.068 |
|
|
|
| 4 |
H |
0.059 |
|
|
|
| 5 |
H |
0.060 |
|
|
|
| 6 |
C |
-0.131 |
|
|
|
| 7 |
H |
0.072 |
|
|
|
| 8 |
H |
0.075 |
|
|
|
| 9 |
C |
0.119 |
|
|
|
| 10 |
H |
0.026 |
|
|
|
| 11 |
C |
-0.102 |
|
|
|
| 12 |
H |
0.078 |
|
|
|
| 13 |
H |
0.063 |
|
|
|
| 14 |
C |
-0.127 |
|
|
|
| 15 |
H |
0.062 |
|
|
|
| 16 |
H |
0.051 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.933 |
0.842 |
0.733 |
2.232 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.591 |
-2.113 |
-0.125 |
| y |
-2.113 |
-43.829 |
-1.278 |
| z |
-0.125 |
-1.278 |
-46.382 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.486 |
-2.113 |
-0.125 |
| y |
-2.113 |
4.157 |
-1.278 |
| z |
-0.125 |
-1.278 |
0.328 |
|
| Polar |
|---|
| 3z2-r2 | 0.657 |
|---|
| x2-y2 | -5.762 |
|---|
| xy | -2.113 |
|---|
| xz | -0.125 |
|---|
| yz | -1.278 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.303 |
0.018 |
0.341 |
| y |
0.018 |
10.960 |
-0.226 |
| z |
0.341 |
-0.226 |
9.378 |
<r2> (average value of r
2) Å
2
| <r2> |
194.424 |
| (<r2>)1/2 |
13.944 |