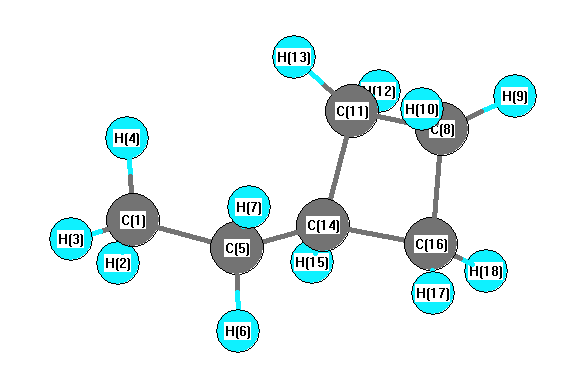
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3242 |
2938 |
125.11 |
|
|
|
| 2 |
A |
3222 |
2920 |
36.82 |
|
|
|
| 3 |
A |
3215 |
2914 |
52.81 |
|
|
|
| 4 |
A |
3212 |
2911 |
77.13 |
|
|
|
| 5 |
A |
3209 |
2909 |
5.32 |
|
|
|
| 6 |
A |
3197 |
2898 |
7.88 |
|
|
|
| 7 |
A |
3183 |
2885 |
89.36 |
|
|
|
| 8 |
A |
3178 |
2881 |
106.41 |
|
|
|
| 9 |
A |
3171 |
2874 |
15.67 |
|
|
|
| 10 |
A |
3157 |
2861 |
8.08 |
|
|
|
| 11 |
A |
3156 |
2860 |
26.59 |
|
|
|
| 12 |
A |
3140 |
2846 |
31.11 |
|
|
|
| 13 |
A |
1637 |
1484 |
1.66 |
|
|
|
| 14 |
A |
1623 |
1471 |
4.45 |
|
|
|
| 15 |
A |
1614 |
1463 |
5.98 |
|
|
|
| 16 |
A |
1609 |
1458 |
0.45 |
|
|
|
| 17 |
A |
1604 |
1454 |
1.38 |
|
|
|
| 18 |
A |
1599 |
1449 |
1.46 |
|
|
|
| 19 |
A |
1537 |
1393 |
2.98 |
|
|
|
| 20 |
A |
1521 |
1378 |
0.10 |
|
|
|
| 21 |
A |
1461 |
1324 |
5.09 |
|
|
|
| 22 |
A |
1418 |
1285 |
0.62 |
|
|
|
| 23 |
A |
1411 |
1279 |
1.30 |
|
|
|
| 24 |
A |
1397 |
1266 |
0.35 |
|
|
|
| 25 |
A |
1380 |
1251 |
0.03 |
|
|
|
| 26 |
A |
1366 |
1238 |
0.04 |
|
|
|
| 27 |
A |
1345 |
1219 |
0.10 |
|
|
|
| 28 |
A |
1300 |
1178 |
0.58 |
|
|
|
| 29 |
A |
1289 |
1168 |
0.05 |
|
|
|
| 30 |
A |
1228 |
1113 |
0.53 |
|
|
|
| 31 |
A |
1147 |
1040 |
0.07 |
|
|
|
| 32 |
A |
1101 |
998 |
0.15 |
|
|
|
| 33 |
A |
1074 |
973 |
0.33 |
|
|
|
| 34 |
A |
1053 |
954 |
1.44 |
|
|
|
| 35 |
A |
1015 |
920 |
0.21 |
|
|
|
| 36 |
A |
993 |
900 |
1.76 |
|
|
|
| 37 |
A |
986 |
893 |
1.06 |
|
|
|
| 38 |
A |
927 |
840 |
0.38 |
|
|
|
| 39 |
A |
869 |
788 |
0.33 |
|
|
|
| 40 |
A |
822 |
745 |
0.12 |
|
|
|
| 41 |
A |
806 |
731 |
1.16 |
|
|
|
| 42 |
A |
727 |
659 |
1.75 |
|
|
|
| 43 |
A |
446 |
404 |
0.05 |
|
|
|
| 44 |
A |
386 |
350 |
0.05 |
|
|
|
| 45 |
A |
261 |
237 |
0.03 |
|
|
|
| 46 |
A |
225 |
204 |
0.02 |
|
|
|
| 47 |
A |
139 |
126 |
0.01 |
|
|
|
| 48 |
A |
113 |
102 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38852.8 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 35216.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.178 |
|
|
|
| 2 |
H |
0.047 |
|
|
|
| 3 |
H |
0.060 |
|
|
|
| 4 |
H |
0.046 |
|
|
|
| 5 |
C |
-0.103 |
|
|
|
| 6 |
H |
0.035 |
|
|
|
| 7 |
H |
0.035 |
|
|
|
| 8 |
C |
-0.099 |
|
|
|
| 9 |
H |
0.045 |
|
|
|
| 10 |
H |
0.045 |
|
|
|
| 11 |
C |
-0.133 |
|
|
|
| 12 |
H |
0.051 |
|
|
|
| 13 |
H |
0.040 |
|
|
|
| 14 |
C |
0.116 |
|
|
|
| 15 |
H |
0.027 |
|
|
|
| 16 |
C |
-0.126 |
|
|
|
| 17 |
H |
0.040 |
|
|
|
| 18 |
H |
0.051 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.019 |
-0.049 |
0.046 |
0.070 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.061 |
-0.366 |
-0.556 |
| y |
-0.366 |
-40.853 |
-0.045 |
| z |
-0.556 |
-0.045 |
-39.707 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.781 |
-0.366 |
-0.556 |
| y |
-0.366 |
-0.469 |
-0.045 |
| z |
-0.556 |
-0.045 |
1.250 |
|
| Polar |
|---|
| 3z2-r2 | 2.500 |
|---|
| x2-y2 | -0.208 |
|---|
| xy | -0.366 |
|---|
| xz | -0.556 |
|---|
| yz | -0.045 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.008 |
0.055 |
0.111 |
| y |
0.055 |
9.618 |
0.015 |
| z |
0.111 |
0.015 |
8.954 |
<r2> (average value of r
2) Å
2
| <r2> |
194.047 |
| (<r2>)1/2 |
13.930 |