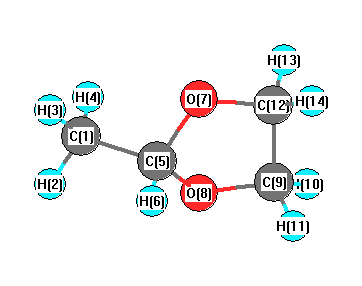
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3259 |
2954 |
52.36 |
|
|
|
| 2 |
A |
3254 |
2950 |
35.27 |
|
|
|
| 3 |
A |
3246 |
2942 |
26.23 |
|
|
|
| 4 |
A |
3221 |
2920 |
55.52 |
|
|
|
| 5 |
A |
3187 |
2888 |
64.50 |
|
|
|
| 6 |
A |
3179 |
2881 |
20.67 |
|
|
|
| 7 |
A |
3153 |
2858 |
64.18 |
|
|
|
| 8 |
A |
3109 |
2818 |
91.87 |
|
|
|
| 9 |
A |
1666 |
1510 |
0.51 |
|
|
|
| 10 |
A |
1648 |
1494 |
3.48 |
|
|
|
| 11 |
A |
1605 |
1455 |
20.86 |
|
|
|
| 12 |
A |
1604 |
1454 |
5.59 |
|
|
|
| 13 |
A |
1581 |
1433 |
72.30 |
|
|
|
| 14 |
A |
1530 |
1387 |
17.29 |
|
|
|
| 15 |
A |
1522 |
1380 |
1.58 |
|
|
|
| 16 |
A |
1517 |
1375 |
1.95 |
|
|
|
| 17 |
A |
1471 |
1334 |
0.63 |
|
|
|
| 18 |
A |
1370 |
1241 |
29.59 |
|
|
|
| 19 |
A |
1343 |
1217 |
3.91 |
|
|
|
| 20 |
A |
1297 |
1175 |
227.00 |
|
|
|
| 21 |
A |
1276 |
1156 |
48.98 |
|
|
|
| 22 |
A |
1240 |
1124 |
46.30 |
|
|
|
| 23 |
A |
1224 |
1109 |
41.31 |
|
|
|
| 24 |
A |
1189 |
1078 |
7.12 |
|
|
|
| 25 |
A |
1153 |
1045 |
45.76 |
|
|
|
| 26 |
A |
1025 |
929 |
13.38 |
|
|
|
| 27 |
A |
984 |
892 |
50.44 |
|
|
|
| 28 |
A |
955 |
866 |
4.59 |
|
|
|
| 29 |
A |
922 |
835 |
4.48 |
|
|
|
| 30 |
A |
767 |
695 |
2.41 |
|
|
|
| 31 |
A |
746 |
676 |
4.58 |
|
|
|
| 32 |
A |
540 |
490 |
5.85 |
|
|
|
| 33 |
A |
359 |
325 |
5.54 |
|
|
|
| 34 |
A |
249 |
226 |
0.07 |
|
|
|
| 35 |
A |
203 |
184 |
4.56 |
|
|
|
| 36 |
A |
66 |
60 |
5.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28329.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 25677.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.162 |
|
|
|
| 2 |
H |
0.059 |
|
|
|
| 3 |
H |
0.058 |
|
|
|
| 4 |
H |
0.052 |
|
|
|
| 5 |
C |
0.405 |
|
|
|
| 6 |
H |
0.016 |
|
|
|
| 7 |
O |
-0.427 |
|
|
|
| 8 |
O |
-0.415 |
|
|
|
| 9 |
C |
0.099 |
|
|
|
| 10 |
H |
0.056 |
|
|
|
| 11 |
H |
0.039 |
|
|
|
| 12 |
C |
0.150 |
|
|
|
| 13 |
H |
0.036 |
|
|
|
| 14 |
H |
0.033 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.171 |
-0.187 |
0.417 |
1.257 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.419 |
-0.088 |
-0.758 |
| y |
-0.088 |
-41.062 |
-0.704 |
| z |
-0.758 |
-0.704 |
-35.708 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.966 |
-0.088 |
-0.758 |
| y |
-0.088 |
-7.499 |
-0.704 |
| z |
-0.758 |
-0.704 |
0.533 |
|
| Polar |
|---|
| 3z2-r2 | 1.066 |
|---|
| x2-y2 | 9.643 |
|---|
| xy | -0.088 |
|---|
| xz | -0.758 |
|---|
| yz | -0.704 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.614 |
0.016 |
-0.126 |
| y |
0.016 |
6.883 |
-0.102 |
| z |
-0.126 |
-0.102 |
6.765 |
<r2> (average value of r
2) Å
2
| <r2> |
147.840 |
| (<r2>)1/2 |
12.159 |