Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -151.148084 |
| Energy at 298.15K | |
| HF Energy | -151.148084 |
| Nuclear repulsion energy | 53.402442 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3601 |
3264 |
123.16 |
55.89 |
0.25 |
0.40 |
| 2 |
Σ |
2098 |
1902 |
192.48 |
110.92 |
0.44 |
0.61 |
| 3 |
Σ |
1397 |
1266 |
16.54 |
80.81 |
0.22 |
0.36 |
| 4 |
Π |
639 |
579 |
0.40 |
0.75 |
0.75 |
0.86 |
| 4 |
Π |
577 |
523 |
10.11 |
0.11 |
0.75 |
0.86 |
| 5 |
Π |
396 |
359 |
32.44 |
0.65 |
0.75 |
0.86 |
| 5 |
Π |
137i |
124i |
157.81 |
9.11 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4285.3 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 3884.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
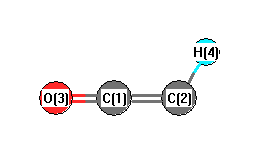
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.024 |
| C2 |
0.000 |
0.000 |
-1.222 |
| O3 |
0.000 |
0.000 |
1.183 |
| H4 |
0.000 |
0.000 |
-2.274 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2455 | 1.1588 | 2.2980 |
C2 | 1.2455 | | 2.4043 | 1.0526 | O3 | 1.1588 | 2.4043 | | 3.4569 | H4 | 2.2980 | 1.0526 | 3.4569 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.433 |
|
|
|
| 2 |
C |
-0.374 |
|
|
|
| 3 |
O |
-0.290 |
|
|
|
| 4 |
H |
0.232 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.201 |
2.201 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.265 |
0.000 |
0.000 |
| y |
0.000 |
-18.039 |
0.000 |
| z |
0.000 |
0.000 |
-15.092 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.301 |
0.000 |
0.000 |
| y |
0.000 |
-2.361 |
0.000 |
| z |
0.000 |
0.000 |
2.060 |
|
| Polar |
|---|
| 3z2-r2 | 4.120 |
|---|
| x2-y2 | 1.774 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.095 |
0.000 |
0.000 |
| y |
0.000 |
2.334 |
0.000 |
| z |
0.000 |
0.000 |
6.032 |
<r2> (average value of r
2) Å
2
| <r2> |
35.603 |
| (<r2>)1/2 |
5.967 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -151.148326 |
| Energy at 298.15K | |
| HF Energy | -151.148326 |
| Nuclear repulsion energy | 53.357464 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3531 |
3201 |
81.21 |
65.76 |
0.32 |
0.49 |
| 2 |
A' |
2151 |
1950 |
403.23 |
80.26 |
0.33 |
0.49 |
| 3 |
A' |
1366 |
1238 |
3.89 |
52.80 |
0.24 |
0.39 |
| 4 |
A' |
637 |
578 |
8.92 |
21.93 |
0.51 |
0.67 |
| 5 |
A' |
251 |
227 |
328.40 |
5.16 |
0.52 |
0.68 |
| 6 |
A" |
561 |
509 |
16.24 |
28.61 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4248.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 3851.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.040 |
0.000 |
| C2 |
0.793 |
-0.952 |
0.000 |
| O3 |
-0.821 |
0.841 |
0.000 |
| H4 |
1.807 |
-1.255 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2703 | 1.1470 | 2.2231 |
C2 | 1.2703 | | 2.4127 | 1.0582 | O3 | 1.1470 | 2.4127 | | 3.3615 | H4 | 2.2231 | 1.0582 | 3.3615 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
145.251 |
|
C2 |
C1 |
O3 |
172.947 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.375 |
|
|
|
| 2 |
C |
-0.276 |
|
|
|
| 3 |
O |
-0.262 |
|
|
|
| 4 |
H |
0.163 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.824 |
-0.407 |
0.000 |
1.869 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.269 |
-0.709 |
0.000 |
| y |
-0.709 |
-19.104 |
0.000 |
| z |
0.000 |
0.000 |
-16.344 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.454 |
-0.709 |
0.000 |
| y |
-0.709 |
-3.297 |
0.000 |
| z |
0.000 |
0.000 |
0.843 |
|
| Polar |
|---|
| 3z2-r2 | 1.686 |
|---|
| x2-y2 | 3.834 |
|---|
| xy | -0.709 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.877 |
-1.752 |
0.000 |
| y |
-1.752 |
4.634 |
0.000 |
| z |
0.000 |
0.000 |
2.916 |
<r2> (average value of r
2) Å
2
| <r2> |
35.675 |
| (<r2>)1/2 |
5.973 |
