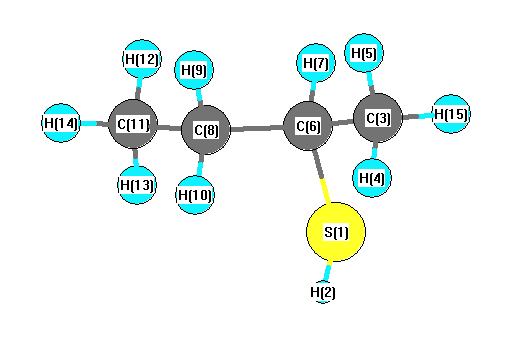
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3244 |
2940 |
42.80 |
|
|
|
| 2 |
A |
3242 |
2939 |
26.32 |
|
|
|
| 3 |
A |
3215 |
2914 |
86.23 |
|
|
|
| 4 |
A |
3211 |
2910 |
29.59 |
|
|
|
| 5 |
A |
3184 |
2886 |
14.20 |
|
|
|
| 6 |
A |
3168 |
2872 |
13.04 |
|
|
|
| 7 |
A |
3160 |
2864 |
14.61 |
|
|
|
| 8 |
A |
3158 |
2862 |
60.56 |
|
|
|
| 9 |
A |
3144 |
2850 |
19.68 |
|
|
|
| 10 |
A |
2842 |
2576 |
4.05 |
|
|
|
| 11 |
A |
1625 |
1473 |
4.78 |
|
|
|
| 12 |
A |
1614 |
1463 |
4.82 |
|
|
|
| 13 |
A |
1613 |
1462 |
7.87 |
|
|
|
| 14 |
A |
1611 |
1460 |
3.75 |
|
|
|
| 15 |
A |
1604 |
1454 |
1.21 |
|
|
|
| 16 |
A |
1540 |
1396 |
6.21 |
|
|
|
| 17 |
A |
1538 |
1394 |
0.65 |
|
|
|
| 18 |
A |
1506 |
1365 |
1.66 |
|
|
|
| 19 |
A |
1447 |
1311 |
14.98 |
|
|
|
| 20 |
A |
1425 |
1291 |
1.07 |
|
|
|
| 21 |
A |
1383 |
1254 |
4.56 |
|
|
|
| 22 |
A |
1280 |
1161 |
1.29 |
|
|
|
| 23 |
A |
1215 |
1102 |
10.31 |
|
|
|
| 24 |
A |
1199 |
1087 |
1.70 |
|
|
|
| 25 |
A |
1095 |
993 |
0.14 |
|
|
|
| 26 |
A |
1089 |
987 |
4.07 |
|
|
|
| 27 |
A |
1039 |
942 |
2.33 |
|
|
|
| 28 |
A |
950 |
861 |
5.81 |
|
|
|
| 29 |
A |
911 |
826 |
3.93 |
|
|
|
| 30 |
A |
851 |
771 |
5.02 |
|
|
|
| 31 |
A |
657 |
595 |
5.47 |
|
|
|
| 32 |
A |
485 |
440 |
0.41 |
|
|
|
| 33 |
A |
403 |
365 |
0.75 |
|
|
|
| 34 |
A |
357 |
324 |
0.90 |
|
|
|
| 35 |
A |
269 |
244 |
0.13 |
|
|
|
| 36 |
A |
248 |
225 |
0.15 |
|
|
|
| 37 |
A |
233 |
211 |
1.43 |
|
|
|
| 38 |
A |
211 |
191 |
15.37 |
|
|
|
| 39 |
A |
111 |
101 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30537.6 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 27679.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.176 |
|
|
|
| 2 |
H |
0.050 |
|
|
|
| 3 |
C |
-0.190 |
|
|
|
| 4 |
H |
0.049 |
|
|
|
| 5 |
H |
0.054 |
|
|
|
| 6 |
C |
0.099 |
|
|
|
| 7 |
H |
0.032 |
|
|
|
| 8 |
C |
-0.055 |
|
|
|
| 9 |
H |
0.039 |
|
|
|
| 10 |
H |
0.030 |
|
|
|
| 11 |
C |
-0.181 |
|
|
|
| 12 |
H |
0.042 |
|
|
|
| 13 |
H |
0.068 |
|
|
|
| 14 |
H |
0.060 |
|
|
|
| 15 |
H |
0.078 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.653 |
1.596 |
0.452 |
1.782 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.008 |
0.675 |
1.285 |
| y |
0.675 |
-43.242 |
-1.364 |
| z |
1.285 |
-1.364 |
-39.006 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.884 |
0.675 |
1.285 |
| y |
0.675 |
-2.235 |
-1.364 |
| z |
1.285 |
-1.364 |
4.119 |
|
| Polar |
|---|
| 3z2-r2 | 8.238 |
|---|
| x2-y2 | 0.234 |
|---|
| xy | 0.675 |
|---|
| xz | 1.285 |
|---|
| yz | -1.364 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.165 |
-0.328 |
0.181 |
| y |
-0.328 |
10.276 |
-0.127 |
| z |
0.181 |
-0.127 |
8.568 |
<r2> (average value of r
2) Å
2
| <r2> |
188.344 |
| (<r2>)1/2 |
13.724 |