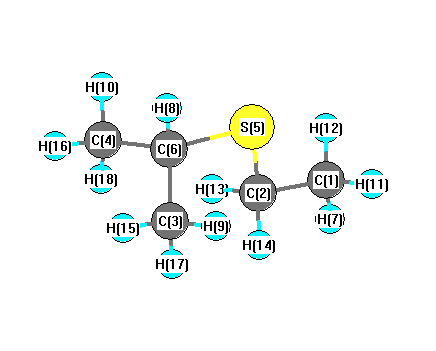
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3243 |
2939 |
51.92 |
|
|
|
| 2 |
A' |
3226 |
2924 |
48.19 |
|
|
|
| 3 |
A' |
3220 |
2918 |
73.04 |
|
|
|
| 4 |
A' |
3194 |
2895 |
15.04 |
|
|
|
| 5 |
A' |
3178 |
2881 |
3.97 |
|
|
|
| 6 |
A' |
3165 |
2869 |
42.16 |
|
|
|
| 7 |
A' |
3163 |
2867 |
43.23 |
|
|
|
| 8 |
A' |
1626 |
1474 |
5.54 |
|
|
|
| 9 |
A' |
1620 |
1468 |
0.61 |
|
|
|
| 10 |
A' |
1619 |
1467 |
13.53 |
|
|
|
| 11 |
A' |
1612 |
1461 |
0.95 |
|
|
|
| 12 |
A' |
1545 |
1400 |
2.40 |
|
|
|
| 13 |
A' |
1536 |
1392 |
1.06 |
|
|
|
| 14 |
A' |
1421 |
1288 |
32.97 |
|
|
|
| 15 |
A' |
1419 |
1286 |
10.43 |
|
|
|
| 16 |
A' |
1286 |
1166 |
6.83 |
|
|
|
| 17 |
A' |
1188 |
1076 |
16.99 |
|
|
|
| 18 |
A' |
1133 |
1027 |
0.12 |
|
|
|
| 19 |
A' |
1052 |
954 |
2.22 |
|
|
|
| 20 |
A' |
957 |
867 |
1.14 |
|
|
|
| 21 |
A' |
756 |
685 |
1.58 |
|
|
|
| 22 |
A' |
664 |
601 |
2.58 |
|
|
|
| 23 |
A' |
500 |
453 |
1.09 |
|
|
|
| 24 |
A' |
396 |
359 |
0.09 |
|
|
|
| 25 |
A' |
309 |
280 |
0.71 |
|
|
|
| 26 |
A' |
270 |
244 |
0.02 |
|
|
|
| 27 |
A' |
170 |
154 |
0.23 |
|
|
|
| 28 |
A" |
3250 |
2946 |
32.13 |
|
|
|
| 29 |
A" |
3238 |
2935 |
41.70 |
|
|
|
| 30 |
A" |
3223 |
2922 |
0.21 |
|
|
|
| 31 |
A" |
3209 |
2908 |
0.79 |
|
|
|
| 32 |
A" |
3157 |
2861 |
21.09 |
|
|
|
| 33 |
A" |
1611 |
1461 |
0.40 |
|
|
|
| 34 |
A" |
1609 |
1458 |
8.47 |
|
|
|
| 35 |
A" |
1603 |
1453 |
0.80 |
|
|
|
| 36 |
A" |
1527 |
1384 |
5.18 |
|
|
|
| 37 |
A" |
1453 |
1317 |
0.31 |
|
|
|
| 38 |
A" |
1377 |
1248 |
0.00 |
|
|
|
| 39 |
A" |
1204 |
1092 |
1.23 |
|
|
|
| 40 |
A" |
1131 |
1025 |
0.11 |
|
|
|
| 41 |
A" |
1029 |
932 |
0.14 |
|
|
|
| 42 |
A" |
1004 |
910 |
0.50 |
|
|
|
| 43 |
A" |
850 |
771 |
1.97 |
|
|
|
| 44 |
A" |
347 |
314 |
1.55 |
|
|
|
| 45 |
A" |
256 |
232 |
0.10 |
|
|
|
| 46 |
A" |
246 |
223 |
0.00 |
|
|
|
| 47 |
A" |
76 |
68 |
1.41 |
|
|
|
| 48 |
A" |
45 |
41 |
0.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37455.0 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 33949.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.180 |
|
|
|
| 2 |
C |
-0.079 |
|
|
|
| 3 |
C |
-0.220 |
|
|
|
| 4 |
C |
-0.220 |
|
|
|
| 5 |
S |
-0.129 |
|
|
|
| 6 |
C |
0.128 |
|
|
|
| 7 |
H |
0.056 |
|
|
|
| 8 |
H |
0.038 |
|
|
|
| 9 |
H |
0.076 |
|
|
|
| 10 |
H |
0.076 |
|
|
|
| 11 |
H |
0.066 |
|
|
|
| 12 |
H |
0.066 |
|
|
|
| 13 |
H |
0.052 |
|
|
|
| 14 |
H |
0.052 |
|
|
|
| 15 |
H |
0.060 |
|
|
|
| 16 |
H |
0.060 |
|
|
|
| 17 |
H |
0.048 |
|
|
|
| 18 |
H |
0.048 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.661 |
-0.515 |
0.000 |
1.739 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.422 |
1.977 |
0.000 |
| y |
1.977 |
-44.015 |
0.000 |
| z |
0.000 |
0.000 |
-48.659 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.085 |
1.977 |
0.000 |
| y |
1.977 |
5.025 |
0.000 |
| z |
0.000 |
0.000 |
-1.940 |
|
| Polar |
|---|
| 3z2-r2 | -3.880 |
|---|
| x2-y2 | -5.407 |
|---|
| xy | 1.977 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.478 |
-0.099 |
0.000 |
| y |
-0.099 |
13.864 |
0.000 |
| z |
0.000 |
0.000 |
10.193 |
<r2> (average value of r
2) Å
2
| <r2> |
266.843 |
| (<r2>)1/2 |
16.335 |