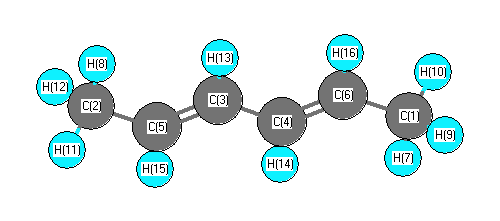
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3278 |
2971 |
0.00 |
|
|
|
| 2 |
Ag |
3274 |
2968 |
0.00 |
|
|
|
| 3 |
Ag |
3227 |
2925 |
0.00 |
|
|
|
| 4 |
Ag |
3151 |
2856 |
0.00 |
|
|
|
| 5 |
Ag |
1889 |
1713 |
0.00 |
|
|
|
| 6 |
Ag |
1610 |
1460 |
0.00 |
|
|
|
| 7 |
Ag |
1541 |
1397 |
0.00 |
|
|
|
| 8 |
Ag |
1470 |
1333 |
0.00 |
|
|
|
| 9 |
Ag |
1434 |
1299 |
0.00 |
|
|
|
| 10 |
Ag |
1274 |
1155 |
0.00 |
|
|
|
| 11 |
Ag |
1163 |
1054 |
0.00 |
|
|
|
| 12 |
Ag |
1000 |
907 |
0.00 |
|
|
|
| 13 |
Ag |
459 |
416 |
0.00 |
|
|
|
| 14 |
Ag |
358 |
325 |
0.00 |
|
|
|
| 15 |
Au |
3191 |
2892 |
65.28 |
|
|
|
| 16 |
Au |
1596 |
1447 |
11.92 |
|
|
|
| 17 |
Au |
1171 |
1061 |
7.01 |
|
|
|
| 18 |
Au |
1123 |
1018 |
80.83 |
|
|
|
| 19 |
Au |
852 |
772 |
2.19 |
|
|
|
| 20 |
Au |
257 |
233 |
1.18 |
|
|
|
| 21 |
Au |
179 |
162 |
0.33 |
|
|
|
| 22 |
Au |
108 |
98 |
1.77 |
|
|
|
| 23 |
Bg |
3191 |
2892 |
0.00 |
|
|
|
| 24 |
Bg |
1596 |
1447 |
0.00 |
|
|
|
| 25 |
Bg |
1173 |
1063 |
0.00 |
|
|
|
| 26 |
Bg |
1074 |
974 |
0.00 |
|
|
|
| 27 |
Bg |
964 |
874 |
0.00 |
|
|
|
| 28 |
Bg |
367 |
333 |
0.00 |
|
|
|
| 29 |
Bg |
211 |
191 |
0.00 |
|
|
|
| 30 |
Bu |
3288 |
2980 |
74.35 |
|
|
|
| 31 |
Bu |
3271 |
2965 |
22.93 |
|
|
|
| 32 |
Bu |
3227 |
2925 |
38.97 |
|
|
|
| 33 |
Bu |
3150 |
2856 |
111.68 |
|
|
|
| 34 |
Bu |
1830 |
1659 |
3.67 |
|
|
|
| 35 |
Bu |
1606 |
1456 |
17.87 |
|
|
|
| 36 |
Bu |
1535 |
1391 |
14.50 |
|
|
|
| 37 |
Bu |
1445 |
1310 |
3.03 |
|
|
|
| 38 |
Bu |
1370 |
1242 |
3.52 |
|
|
|
| 39 |
Bu |
1169 |
1060 |
1.40 |
|
|
|
| 40 |
Bu |
994 |
901 |
20.32 |
|
|
|
| 41 |
Bu |
583 |
528 |
0.26 |
|
|
|
| 42 |
Bu |
152 |
138 |
1.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32900.1 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 29820.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.141 |
|
|
|
| 2 |
C |
-0.141 |
|
|
|
| 3 |
C |
-0.091 |
|
|
|
| 4 |
C |
-0.091 |
|
|
|
| 5 |
C |
-0.032 |
|
|
|
| 6 |
C |
-0.032 |
|
|
|
| 7 |
H |
0.058 |
|
|
|
| 8 |
H |
0.058 |
|
|
|
| 9 |
H |
0.061 |
|
|
|
| 10 |
H |
0.061 |
|
|
|
| 11 |
H |
0.061 |
|
|
|
| 12 |
H |
0.061 |
|
|
|
| 13 |
H |
0.039 |
|
|
|
| 14 |
H |
0.039 |
|
|
|
| 15 |
H |
0.046 |
|
|
|
| 16 |
H |
0.046 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.389 |
-0.601 |
0.000 |
| y |
-0.601 |
-35.287 |
0.000 |
| z |
0.000 |
0.000 |
-41.849 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.179 |
-0.601 |
0.000 |
| y |
-0.601 |
3.832 |
0.000 |
| z |
0.000 |
0.000 |
-6.011 |
|
| Polar |
|---|
| 3z2-r2 | -12.021 |
|---|
| x2-y2 | -1.102 |
|---|
| xy | -0.601 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.263 |
-1.446 |
0.000 |
| y |
-1.446 |
18.180 |
0.000 |
| z |
0.000 |
0.000 |
7.414 |
<r2> (average value of r
2) Å
2
| <r2> |
288.274 |
| (<r2>)1/2 |
16.979 |